Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -554.205332 |
| Energy at 298.15K | -554.210383 |
| Nuclear repulsion energy | 199.922681 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3263 |
3123 |
7.68 |
|
|
|
| 2 |
A |
3187 |
3050 |
4.01 |
|
|
|
| 3 |
A |
3170 |
3034 |
4.59 |
|
|
|
| 4 |
A |
1684 |
1612 |
4.05 |
|
|
|
| 5 |
A |
1426 |
1365 |
8.66 |
|
|
|
| 6 |
A |
1322 |
1265 |
0.41 |
|
|
|
| 7 |
A |
1065 |
1020 |
0.23 |
|
|
|
| 8 |
A |
985 |
943 |
0.00 |
|
|
|
| 9 |
A |
908 |
869 |
0.00 |
|
|
|
| 10 |
A |
735 |
704 |
0.46 |
|
|
|
| 11 |
A |
624 |
597 |
0.00 |
|
|
|
| 12 |
A |
409 |
392 |
0.55 |
|
|
|
| 13 |
A |
178 |
170 |
0.19 |
|
|
|
| 14 |
A |
45 |
43 |
0.00 |
|
|
|
| 15 |
A |
3263 |
3123 |
0.39 |
|
|
|
| 16 |
A |
3185 |
3049 |
1.33 |
|
|
|
| 17 |
A |
3170 |
3034 |
0.04 |
|
|
|
| 18 |
A |
1660 |
1589 |
166.88 |
|
|
|
| 19 |
A |
1420 |
1359 |
7.43 |
|
|
|
| 20 |
A |
1281 |
1226 |
12.85 |
|
|
|
| 21 |
A |
1031 |
986 |
27.52 |
|
|
|
| 22 |
A |
994 |
951 |
40.42 |
|
|
|
| 23 |
A |
909 |
870 |
81.90 |
|
|
|
| 24 |
A |
753 |
721 |
43.35 |
|
|
|
| 25 |
A |
562 |
538 |
22.83 |
|
|
|
| 26 |
A |
383 |
367 |
4.77 |
|
|
|
| 27 |
A |
18 |
17 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18814.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 18007.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.623 |
| C2 |
0.000 |
1.351 |
-0.484 |
| C3 |
0.000 |
-1.351 |
-0.484 |
| C4 |
0.000 |
2.620 |
-0.105 |
| C5 |
-0.000 |
-2.620 |
-0.105 |
| H6 |
-0.000 |
1.079 |
-1.531 |
| H7 |
0.000 |
-1.079 |
-1.531 |
| H8 |
0.000 |
3.405 |
-0.845 |
| H9 |
0.000 |
2.914 |
0.934 |
| H10 |
-0.000 |
-3.405 |
-0.845 |
| H11 |
-0.000 |
-2.914 |
0.934 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7470 | 1.7470 | 2.7195 | 2.7195 | 2.4091 | 2.4091 | 3.7082 | 2.9308 | 3.7082 | 2.9308 |
C2 | 1.7470 | | 2.7028 | 1.3242 | 3.9896 | 1.0817 | 2.6463 | 2.0853 | 2.1103 | 4.7703 | 4.4952 | C3 | 1.7470 | 2.7028 | | 3.9896 | 1.3242 | 2.6463 | 1.0817 | 4.7703 | 4.4952 | 2.0853 | 2.1103 | C4 | 2.7195 | 1.3242 | 3.9896 | | 5.2404 | 2.0995 | 3.9645 | 1.0787 | 1.0798 | 6.0707 | 5.6312 | C5 | 2.7195 | 3.9896 | 1.3242 | 5.2404 | | 3.9645 | 2.0995 | 6.0707 | 5.6312 | 1.0787 | 1.0798 | H6 | 2.4091 | 1.0817 | 2.6463 | 2.0995 | 3.9645 | | 2.1581 | 2.4253 | 3.0730 | 4.5365 | 4.6928 | H7 | 2.4091 | 2.6463 | 1.0817 | 3.9645 | 2.0995 | 2.1581 | | 4.5365 | 4.6928 | 2.4253 | 3.0730 | H8 | 3.7082 | 2.0853 | 4.7703 | 1.0787 | 6.0707 | 2.4253 | 4.5365 | | 1.8453 | 6.8105 | 6.5651 | H9 | 2.9308 | 2.1103 | 4.4952 | 1.0798 | 5.6312 | 3.0730 | 4.6928 | 1.8453 | | 6.5651 | 5.8286 | H10 | 3.7082 | 4.7703 | 2.0853 | 6.0707 | 1.0787 | 4.5365 | 2.4253 | 6.8105 | 6.5651 | | 1.8453 | H11 | 2.9308 | 4.4952 | 2.1103 | 5.6312 | 1.0798 | 4.6928 | 3.0730 | 6.5651 | 5.8286 | 1.8453 | |
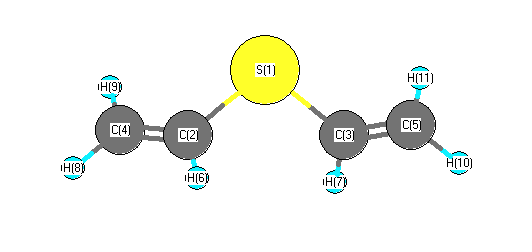 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
124.044 |
|
S1 |
C2 |
H6 |
114.744 |
| S1 |
C3 |
C5 |
124.044 |
|
S1 |
C3 |
H7 |
114.744 |
| C2 |
S1 |
C3 |
101.348 |
|
C2 |
C4 |
H8 |
120.073 |
| C2 |
C4 |
H9 |
122.433 |
|
C3 |
C5 |
H10 |
120.073 |
| C3 |
C5 |
H11 |
122.433 |
|
C4 |
C2 |
H6 |
121.212 |
| C5 |
C3 |
H7 |
121.212 |
|
H8 |
C4 |
H9 |
117.494 |
| H10 |
C5 |
H11 |
117.494 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.075 |
|
|
|
| 2 |
C |
-0.114 |
|
|
|
| 3 |
C |
-0.114 |
|
|
|
| 4 |
C |
-0.333 |
|
|
|
| 5 |
C |
-0.333 |
|
|
|
| 6 |
H |
0.135 |
|
|
|
| 7 |
H |
0.135 |
|
|
|
| 8 |
H |
0.142 |
|
|
|
| 9 |
H |
0.133 |
|
|
|
| 10 |
H |
0.142 |
|
|
|
| 11 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.087 |
1.087 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.032 |
0.000 |
0.000 |
| y |
0.000 |
-33.727 |
0.000 |
| z |
0.000 |
0.000 |
-35.072 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.632 |
0.000 |
0.000 |
| y |
0.000 |
4.825 |
0.000 |
| z |
0.000 |
0.000 |
2.807 |
|
| Polar |
|---|
| 3z2-r2 | 5.614 |
|---|
| x2-y2 | -8.305 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.340 |
0.000 |
0.000 |
| y |
0.000 |
17.577 |
0.000 |
| z |
0.000 |
0.000 |
8.334 |
<r2> (average value of r
2) Å
2
| <r2> |
186.896 |
| (<r2>)1/2 |
13.671 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -554.206053 |
| Energy at 298.15K | -554.211605 |
| HF Energy | -554.206053 |
| Nuclear repulsion energy | 203.273877 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3261 |
3121 |
6.74 |
|
|
|
| 2 |
A |
3259 |
3119 |
3.85 |
|
|
|
| 3 |
A |
3195 |
3058 |
2.55 |
|
|
|
| 4 |
A |
3182 |
3045 |
4.47 |
|
|
|
| 5 |
A |
3170 |
3034 |
0.85 |
|
|
|
| 6 |
A |
3165 |
3029 |
1.90 |
|
|
|
| 7 |
A |
1675 |
1603 |
50.30 |
|
|
|
| 8 |
A |
1665 |
1594 |
35.63 |
|
|
|
| 9 |
A |
1426 |
1365 |
5.91 |
|
|
|
| 10 |
A |
1420 |
1359 |
5.34 |
|
|
|
| 11 |
A |
1305 |
1249 |
1.85 |
|
|
|
| 12 |
A |
1288 |
1233 |
7.19 |
|
|
|
| 13 |
A |
1061 |
1016 |
8.37 |
|
|
|
| 14 |
A |
1031 |
987 |
10.31 |
|
|
|
| 15 |
A |
1016 |
972 |
28.96 |
|
|
|
| 16 |
A |
997 |
954 |
22.63 |
|
|
|
| 17 |
A |
943 |
903 |
36.95 |
|
|
|
| 18 |
A |
909 |
870 |
44.53 |
|
|
|
| 19 |
A |
757 |
724 |
16.05 |
|
|
|
| 20 |
A |
711 |
681 |
4.92 |
|
|
|
| 21 |
A |
633 |
606 |
13.72 |
|
|
|
| 22 |
A |
613 |
587 |
13.84 |
|
|
|
| 23 |
A |
467 |
447 |
0.57 |
|
|
|
| 24 |
A |
365 |
350 |
0.53 |
|
|
|
| 25 |
A |
239 |
229 |
0.85 |
|
|
|
| 26 |
A |
157 |
150 |
4.17 |
|
|
|
| 27 |
A |
92 |
88 |
0.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19000.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 18185.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.025 |
-0.957 |
0.011 |
| C2 |
1.067 |
0.413 |
0.317 |
| C3 |
-1.557 |
-0.216 |
-0.016 |
| C4 |
2.284 |
0.526 |
-0.192 |
| C5 |
-1.854 |
1.070 |
-0.129 |
| H6 |
0.676 |
1.130 |
1.028 |
| H7 |
-2.337 |
-0.960 |
0.077 |
| H8 |
2.932 |
1.334 |
0.114 |
| H9 |
2.671 |
-0.182 |
-0.910 |
| H10 |
-1.099 |
1.832 |
-0.255 |
| H11 |
-2.884 |
1.390 |
-0.106 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7480 | 1.7472 | 2.7096 | 2.7673 | 2.4108 | 2.3634 | 3.7023 | 2.9067 | 3.0182 | 3.7397 |
C2 | 1.7480 | | 2.7193 | 1.3240 | 3.0268 | 1.0824 | 3.6786 | 2.0898 | 2.1053 | 2.6521 | 4.0922 | C3 | 1.7472 | 2.7193 | | 3.9162 | 1.3253 | 2.8092 | 1.0814 | 4.7511 | 4.3220 | 2.1123 | 2.0856 | C4 | 2.7096 | 1.3240 | 3.9162 | | 4.1735 | 2.1064 | 4.8617 | 1.0798 | 1.0799 | 3.6267 | 5.2404 | C5 | 2.7673 | 3.0268 | 1.3253 | 4.1735 | | 2.7826 | 2.0970 | 4.7988 | 4.7592 | 1.0792 | 1.0793 | H6 | 2.4108 | 1.0824 | 2.8092 | 2.1064 | 2.7826 | | 3.7885 | 2.4423 | 3.0747 | 2.3005 | 3.7458 | H7 | 2.3634 | 3.6786 | 1.0814 | 4.8617 | 2.0970 | 3.7885 | | 5.7469 | 5.1637 | 3.0717 | 2.4195 | H8 | 3.7023 | 2.0898 | 4.7511 | 1.0798 | 4.7988 | 2.4423 | 5.7469 | | 1.8474 | 4.0783 | 5.8204 | H9 | 2.9067 | 2.1053 | 4.3220 | 1.0799 | 4.7592 | 3.0747 | 5.1637 | 1.8474 | | 4.3241 | 5.8291 | H10 | 3.0182 | 2.6521 | 2.1123 | 3.6267 | 1.0792 | 2.3005 | 3.0717 | 4.0783 | 4.3241 | | 1.8448 | H11 | 3.7397 | 4.0922 | 2.0856 | 5.2404 | 1.0793 | 3.7458 | 2.4195 | 5.8204 | 5.8291 | 1.8448 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.186 |
|
S1 |
C2 |
H6 |
114.768 |
| S1 |
C3 |
C5 |
127.961 |
|
S1 |
C3 |
H7 |
111.151 |
| C2 |
S1 |
C3 |
102.159 |
|
C2 |
C4 |
H8 |
120.431 |
| C2 |
C4 |
H9 |
121.954 |
|
C3 |
C5 |
H10 |
122.594 |
| C3 |
C5 |
H11 |
119.957 |
|
C4 |
C2 |
H6 |
121.851 |
| C5 |
C3 |
H7 |
120.888 |
|
H8 |
C4 |
H9 |
117.610 |
| H10 |
C5 |
H11 |
117.447 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.038 |
|
|
|
| 2 |
C |
-0.138 |
|
|
|
| 3 |
C |
-0.119 |
|
|
|
| 4 |
C |
-0.310 |
|
|
|
| 5 |
C |
-0.340 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
H |
0.145 |
|
|
|
| 8 |
H |
0.143 |
|
|
|
| 9 |
H |
0.141 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.079 |
0.867 |
0.170 |
0.887 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.106 |
1.540 |
-0.687 |
| y |
1.540 |
-37.023 |
1.102 |
| z |
-0.687 |
1.102 |
-40.502 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.657 |
1.540 |
-0.687 |
| y |
1.540 |
-0.220 |
1.102 |
| z |
-0.687 |
1.102 |
-5.438 |
|
| Polar |
|---|
| 3z2-r2 | -10.876 |
|---|
| x2-y2 | 3.918 |
|---|
| xy | 1.540 |
|---|
| xz | -0.687 |
|---|
| yz | 1.102 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.100 |
-0.189 |
-0.551 |
| y |
-0.189 |
9.935 |
0.177 |
| z |
-0.551 |
0.177 |
6.047 |
<r2> (average value of r
2) Å
2
| <r2> |
164.203 |
| (<r2>)1/2 |
12.814 |
