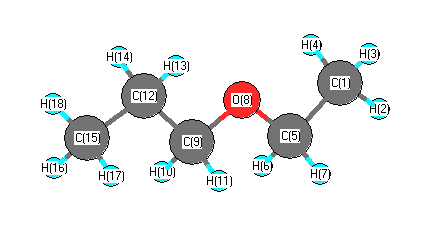
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3143 |
3008 |
25.37 |
|
|
|
| 2 |
A' |
3131 |
2997 |
31.47 |
|
|
|
| 3 |
A' |
3065 |
2933 |
23.72 |
|
|
|
| 4 |
A' |
3064 |
2932 |
22.98 |
|
|
|
| 5 |
A' |
3052 |
2921 |
26.81 |
|
|
|
| 6 |
A' |
2978 |
2851 |
88.14 |
|
|
|
| 7 |
A' |
2963 |
2836 |
27.24 |
|
|
|
| 8 |
A' |
1536 |
1470 |
2.50 |
|
|
|
| 9 |
A' |
1519 |
1453 |
6.41 |
|
|
|
| 10 |
A' |
1507 |
1442 |
4.05 |
|
|
|
| 11 |
A' |
1500 |
1436 |
1.87 |
|
|
|
| 12 |
A' |
1493 |
1429 |
1.12 |
|
|
|
| 13 |
A' |
1451 |
1389 |
3.63 |
|
|
|
| 14 |
A' |
1412 |
1351 |
0.96 |
|
|
|
| 15 |
A' |
1408 |
1347 |
34.39 |
|
|
|
| 16 |
A' |
1388 |
1328 |
16.96 |
|
|
|
| 17 |
A' |
1328 |
1271 |
8.66 |
|
|
|
| 18 |
A' |
1195 |
1144 |
175.36 |
|
|
|
| 19 |
A' |
1181 |
1131 |
75.75 |
|
|
|
| 20 |
A' |
1122 |
1074 |
6.13 |
|
|
|
| 21 |
A' |
1080 |
1034 |
7.22 |
|
|
|
| 22 |
A' |
1052 |
1007 |
7.96 |
|
|
|
| 23 |
A' |
916 |
877 |
3.44 |
|
|
|
| 24 |
A' |
906 |
867 |
7.06 |
|
|
|
| 25 |
A' |
487 |
466 |
3.50 |
|
|
|
| 26 |
A' |
404 |
387 |
0.73 |
|
|
|
| 27 |
A' |
298 |
285 |
0.59 |
|
|
|
| 28 |
A' |
135 |
129 |
0.33 |
|
|
|
| 29 |
A" |
3144 |
3009 |
26.92 |
|
|
|
| 30 |
A" |
3124 |
2990 |
57.84 |
|
|
|
| 31 |
A" |
3097 |
2964 |
2.31 |
|
|
|
| 32 |
A" |
3002 |
2873 |
34.38 |
|
|
|
| 33 |
A" |
2990 |
2862 |
77.46 |
|
|
|
| 34 |
A" |
1501 |
1436 |
7.40 |
|
|
|
| 35 |
A" |
1482 |
1418 |
6.16 |
|
|
|
| 36 |
A" |
1315 |
1259 |
0.48 |
|
|
|
| 37 |
A" |
1301 |
1245 |
2.58 |
|
|
|
| 38 |
A" |
1268 |
1214 |
1.81 |
|
|
|
| 39 |
A" |
1201 |
1149 |
7.10 |
|
|
|
| 40 |
A" |
1175 |
1125 |
0.55 |
|
|
|
| 41 |
A" |
902 |
863 |
1.29 |
|
|
|
| 42 |
A" |
819 |
784 |
0.65 |
|
|
|
| 43 |
A" |
762 |
730 |
1.79 |
|
|
|
| 44 |
A" |
249 |
238 |
0.73 |
|
|
|
| 45 |
A" |
234 |
224 |
0.02 |
|
|
|
| 46 |
A" |
152 |
145 |
2.99 |
|
|
|
| 47 |
A" |
97 |
93 |
1.23 |
|
|
|
| 48 |
A" |
68 |
65 |
0.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36296.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34739.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.343 |
|
|
|
| 2 |
H |
0.105 |
|
|
|
| 3 |
H |
0.119 |
|
|
|
| 4 |
H |
0.119 |
|
|
|
| 5 |
C |
-0.014 |
|
|
|
| 6 |
H |
0.072 |
|
|
|
| 7 |
H |
0.072 |
|
|
|
| 8 |
O |
-0.266 |
|
|
|
| 9 |
C |
-0.008 |
|
|
|
| 10 |
H |
0.070 |
|
|
|
| 11 |
H |
0.070 |
|
|
|
| 12 |
C |
-0.206 |
|
|
|
| 13 |
H |
0.118 |
|
|
|
| 14 |
H |
0.118 |
|
|
|
| 15 |
C |
-0.371 |
|
|
|
| 16 |
H |
0.111 |
|
|
|
| 17 |
H |
0.111 |
|
|
|
| 18 |
H |
0.123 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.812 |
-0.485 |
0.000 |
0.946 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.826 |
1.911 |
0.000 |
| y |
1.911 |
-39.033 |
0.000 |
| z |
0.000 |
0.000 |
-39.383 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.382 |
1.911 |
0.000 |
| y |
1.911 |
0.071 |
0.000 |
| z |
0.000 |
0.000 |
-0.453 |
|
| Polar |
|---|
| 3z2-r2 | -0.907 |
|---|
| x2-y2 | 0.207 |
|---|
| xy | 1.911 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.692 |
1.165 |
0.000 |
| y |
1.165 |
11.703 |
0.000 |
| z |
0.000 |
0.000 |
8.675 |
<r2> (average value of r
2) Å
2
| <r2> |
283.142 |
| (<r2>)1/2 |
16.827 |