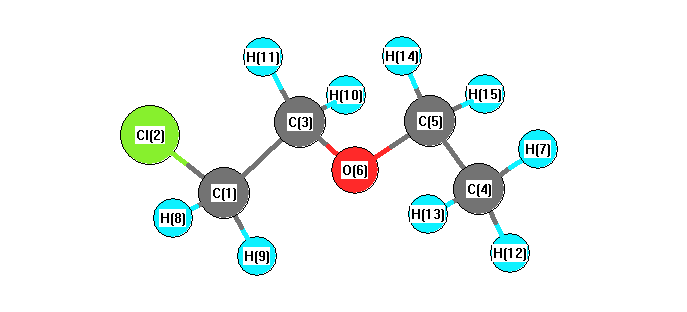
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3145 |
3011 |
22.59 |
|
|
|
| 2 |
A' |
3118 |
2984 |
14.23 |
|
|
|
| 3 |
A' |
3067 |
2935 |
16.86 |
|
|
|
| 4 |
A' |
3001 |
2872 |
54.31 |
|
|
|
| 5 |
A' |
2986 |
2858 |
33.35 |
|
|
|
| 6 |
A' |
1534 |
1468 |
3.02 |
|
|
|
| 7 |
A' |
1517 |
1452 |
4.39 |
|
|
|
| 8 |
A' |
1500 |
1436 |
3.77 |
|
|
|
| 9 |
A' |
1494 |
1430 |
2.80 |
|
|
|
| 10 |
A' |
1448 |
1386 |
10.12 |
|
|
|
| 11 |
A' |
1404 |
1344 |
14.85 |
|
|
|
| 12 |
A' |
1382 |
1323 |
20.01 |
|
|
|
| 13 |
A' |
1276 |
1221 |
14.44 |
|
|
|
| 14 |
A' |
1189 |
1138 |
247.92 |
|
|
|
| 15 |
A' |
1160 |
1111 |
28.74 |
|
|
|
| 16 |
A' |
1090 |
1043 |
3.43 |
|
|
|
| 17 |
A' |
1064 |
1019 |
15.51 |
|
|
|
| 18 |
A' |
910 |
871 |
11.28 |
|
|
|
| 19 |
A' |
780 |
747 |
41.70 |
|
|
|
| 20 |
A' |
474 |
453 |
0.58 |
|
|
|
| 21 |
A' |
374 |
358 |
2.34 |
|
|
|
| 22 |
A' |
263 |
251 |
2.91 |
|
|
|
| 23 |
A' |
119 |
114 |
1.29 |
|
|
|
| 24 |
A" |
3180 |
3044 |
6.02 |
|
|
|
| 25 |
A" |
3146 |
3011 |
24.84 |
|
|
|
| 26 |
A" |
3035 |
2905 |
27.21 |
|
|
|
| 27 |
A" |
3016 |
2886 |
60.48 |
|
|
|
| 28 |
A" |
1482 |
1419 |
6.52 |
|
|
|
| 29 |
A" |
1301 |
1245 |
3.24 |
|
|
|
| 30 |
A" |
1299 |
1243 |
0.25 |
|
|
|
| 31 |
A" |
1218 |
1166 |
5.21 |
|
|
|
| 32 |
A" |
1179 |
1129 |
3.19 |
|
|
|
| 33 |
A" |
1059 |
1014 |
1.86 |
|
|
|
| 34 |
A" |
819 |
784 |
0.81 |
|
|
|
| 35 |
A" |
799 |
765 |
0.29 |
|
|
|
| 36 |
A" |
246 |
236 |
0.84 |
|
|
|
| 37 |
A" |
146 |
140 |
6.86 |
|
|
|
| 38 |
A" |
86 |
82 |
2.08 |
|
|
|
| 39 |
A" |
54 |
52 |
0.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28181.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 26972.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.159 |
|
|
|
| 2 |
Cl |
-0.167 |
|
|
|
| 3 |
C |
-0.025 |
|
|
|
| 4 |
C |
-0.342 |
|
|
|
| 5 |
C |
-0.017 |
|
|
|
| 6 |
O |
-0.276 |
|
|
|
| 7 |
H |
0.109 |
|
|
|
| 8 |
H |
0.146 |
|
|
|
| 9 |
H |
0.146 |
|
|
|
| 10 |
H |
0.093 |
|
|
|
| 11 |
H |
0.093 |
|
|
|
| 12 |
H |
0.121 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.079 |
|
|
|
| 15 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.993 |
-0.310 |
0.000 |
2.017 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.896 |
3.094 |
0.000 |
| y |
3.094 |
-46.019 |
0.000 |
| z |
0.000 |
0.000 |
-44.058 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.857 |
3.094 |
0.000 |
| y |
3.094 |
-1.042 |
0.000 |
| z |
0.000 |
0.000 |
1.899 |
|
| Polar |
|---|
| 3z2-r2 | 3.798 |
|---|
| x2-y2 | 0.123 |
|---|
| xy | 3.094 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.132 |
-1.559 |
0.000 |
| y |
-1.559 |
9.290 |
0.000 |
| z |
0.000 |
0.000 |
7.921 |
<r2> (average value of r
2) Å
2
| <r2> |
361.185 |
| (<r2>)1/2 |
19.005 |