Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.503005 |
| Energy at 298.15K | -189.505726 |
| HF Energy | -188.840478 |
| Nuclear repulsion energy | 70.100241 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3758 |
3565 |
|
|
|
|
| 2 |
A' |
3086 |
2927 |
|
|
|
|
| 3 |
A' |
1819 |
1726 |
|
|
|
|
| 4 |
A' |
1414 |
1341 |
|
|
|
|
| 5 |
A' |
1324 |
1256 |
|
|
|
|
| 6 |
A' |
1133 |
1075 |
|
|
|
|
| 7 |
A' |
628 |
595 |
|
|
|
|
| 8 |
A" |
1060 |
1005 |
|
|
|
|
| 9 |
A" |
678 |
643 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7449.2 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 7066.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.422 |
0.000 |
| O2 |
-1.034 |
-0.442 |
0.000 |
| O3 |
1.162 |
0.110 |
0.000 |
| H4 |
-0.377 |
1.450 |
0.000 |
| H5 |
-0.643 |
-1.329 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3480 | 1.2029 | 1.0947 | 1.8652 |
O2 | 1.3480 | | 2.2644 | 2.0029 | 0.9690 | O3 | 1.2029 | 2.2644 | | 2.0404 | 2.3080 | H4 | 1.0947 | 2.0029 | 2.0404 | | 2.7912 | H5 | 1.8652 | 0.9690 | 2.3080 | 2.7912 | |
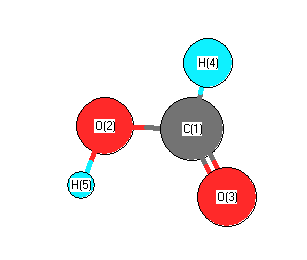 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.066 |
|
O2 |
C1 |
O3 |
125.076 |
| O2 |
C1 |
H4 |
109.724 |
|
O3 |
C1 |
H4 |
125.200 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.495899 |
| Energy at 298.15K | |
| HF Energy | -188.832546 |
| Nuclear repulsion energy | 69.872983 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3825 |
3629 |
|
|
|
|
| 2 |
A' |
2999 |
2845 |
|
|
|
|
| 3 |
A' |
1861 |
1765 |
|
|
|
|
| 4 |
A' |
1427 |
1354 |
|
|
|
|
| 5 |
A' |
1296 |
1229 |
|
|
|
|
| 6 |
A' |
1122 |
1065 |
|
|
|
|
| 7 |
A' |
658 |
624 |
|
|
|
|
| 8 |
A" |
1037 |
984 |
|
|
|
|
| 9 |
A" |
517 |
490 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7371.0 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 6992.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.385 |
0.000 |
| O2 |
-0.900 |
-0.627 |
0.000 |
| O3 |
1.181 |
0.196 |
0.000 |
| H4 |
-0.464 |
1.384 |
0.000 |
| H5 |
-1.785 |
-0.246 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3545 | 1.1965 | 1.1011 | 1.8936 |
O2 | 1.3545 | | 2.2384 | 2.0575 | 0.9636 | O3 | 1.1965 | 2.2384 | | 2.0293 | 2.9995 | H4 | 1.1011 | 2.0575 | 2.0293 | | 2.0981 | H5 | 1.8936 | 0.9636 | 2.9995 | 2.0981 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.365 |
|
O2 |
C1 |
O3 |
122.560 |
| O2 |
C1 |
H4 |
113.429 |
|
O3 |
C1 |
H4 |
124.012 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
