Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.512769 |
| Energy at 298.15K | -189.515506 |
| HF Energy | -188.841177 |
| Nuclear repulsion energy | 70.341383 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3780 |
3589 |
76.00 |
58.63 |
0.22 |
0.36 |
| 2 |
A' |
3141 |
2982 |
39.52 |
105.99 |
0.25 |
0.40 |
| 3 |
A' |
1829 |
1737 |
300.60 |
9.89 |
0.23 |
0.37 |
| 4 |
A' |
1432 |
1360 |
1.68 |
7.68 |
0.48 |
0.64 |
| 5 |
A' |
1322 |
1255 |
16.07 |
1.07 |
0.35 |
0.51 |
| 6 |
A' |
1143 |
1085 |
251.13 |
2.25 |
0.15 |
0.26 |
| 7 |
A' |
634 |
602 |
44.12 |
3.64 |
0.47 |
0.64 |
| 8 |
A" |
1084 |
1029 |
2.53 |
0.95 |
0.75 |
0.86 |
| 9 |
A" |
692 |
657 |
137.73 |
0.88 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7528.0 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 7147.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.420 |
0.000 |
| O2 |
-1.031 |
-0.438 |
0.000 |
| O3 |
1.159 |
0.108 |
0.000 |
| H4 |
-0.375 |
1.441 |
0.000 |
| H5 |
-0.644 |
-1.325 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3417 | 1.2001 | 1.0874 | 1.8601 |
O2 | 1.3417 | | 2.2573 | 1.9899 | 0.9679 | O3 | 1.2001 | 2.2573 | | 2.0320 | 2.3030 | H4 | 1.0874 | 1.9899 | 2.0320 | | 2.7787 | H5 | 1.8601 | 0.9679 | 2.3030 | 2.7787 | |
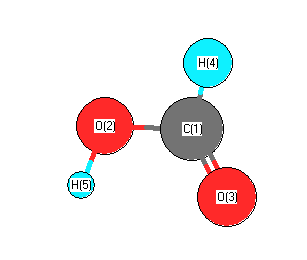 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.167 |
|
O2 |
C1 |
O3 |
125.174 |
| O2 |
C1 |
H4 |
109.572 |
|
O3 |
C1 |
H4 |
125.254 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.505575 |
| Energy at 298.15K | |
| HF Energy | -188.833284 |
| Nuclear repulsion energy | 70.117772 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3853 |
3658 |
78.85 |
77.97 |
0.23 |
0.38 |
| 2 |
A' |
3054 |
2899 |
74.67 |
102.42 |
0.24 |
0.39 |
| 3 |
A' |
1869 |
1774 |
246.43 |
10.32 |
0.22 |
0.36 |
| 4 |
A' |
1450 |
1377 |
0.04 |
7.41 |
0.50 |
0.67 |
| 5 |
A' |
1290 |
1225 |
313.06 |
2.37 |
0.70 |
0.83 |
| 6 |
A' |
1138 |
1080 |
41.37 |
6.66 |
0.34 |
0.50 |
| 7 |
A' |
667 |
634 |
10.87 |
1.09 |
0.51 |
0.68 |
| 8 |
A" |
1062 |
1008 |
0.00 |
1.16 |
0.75 |
0.86 |
| 9 |
A" |
537 |
510 |
85.59 |
1.08 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7459.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 7082.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.383 |
0.000 |
| O2 |
-0.898 |
-0.622 |
0.000 |
| O3 |
1.179 |
0.193 |
0.000 |
| H4 |
-0.460 |
1.375 |
0.000 |
| H5 |
-1.782 |
-0.242 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3478 | 1.1938 | 1.0937 | 1.8886 |
O2 | 1.3478 | | 2.2313 | 2.0446 | 0.9623 | O3 | 1.1938 | 2.2313 | | 2.0204 | 2.9928 | H4 | 1.0937 | 2.0446 | 2.0204 | | 2.0886 | H5 | 1.8886 | 0.9623 | 2.9928 | 2.0886 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.520 |
|
O2 |
C1 |
O3 |
122.670 |
| O2 |
C1 |
H4 |
113.326 |
|
O3 |
C1 |
H4 |
124.004 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
