Jump to
S1C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.550235 |
| Energy at 298.15K | -189.552959 |
| HF Energy | -188.840432 |
| Nuclear repulsion energy | 70.109277 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3759 |
3619 |
|
|
|
|
| 2 |
A' |
3095 |
2981 |
|
|
|
|
| 3 |
A' |
1795 |
1728 |
|
|
|
|
| 4 |
A' |
1416 |
1363 |
|
|
|
|
| 5 |
A' |
1317 |
1268 |
|
|
|
|
| 6 |
A' |
1125 |
1083 |
|
|
|
|
| 7 |
A' |
625 |
602 |
|
|
|
|
| 8 |
A" |
1066 |
1027 |
|
|
|
|
| 9 |
A" |
687 |
662 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7442.2 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 7166.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.422 |
0.000 |
| O2 |
-1.035 |
-0.440 |
0.000 |
| O3 |
1.163 |
0.109 |
0.000 |
| H4 |
-0.377 |
1.445 |
0.000 |
| H5 |
-0.645 |
-1.326 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3467 | 1.2043 | 1.0902 | 1.8634 |
O2 | 1.3467 | | 2.2656 | 1.9963 | 0.9687 | O3 | 1.2043 | 2.2656 | | 2.0383 | 2.3086 | H4 | 1.0902 | 1.9963 | 2.0383 | | 2.7842 | H5 | 1.8634 | 0.9687 | 2.3086 | 2.7842 | |
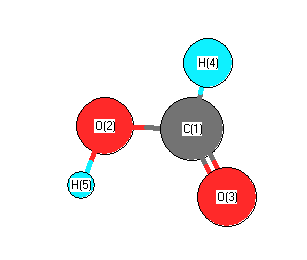 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.020 |
|
O2 |
C1 |
O3 |
125.181 |
| O2 |
C1 |
H4 |
109.556 |
|
O3 |
C1 |
H4 |
125.263 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.543175 |
| Energy at 298.15K | |
| HF Energy | -188.832534 |
| Nuclear repulsion energy | 69.891323 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3827 |
3685 |
|
|
|
|
| 2 |
A' |
3008 |
2896 |
|
|
|
|
| 3 |
A' |
1836 |
1768 |
|
|
|
|
| 4 |
A' |
1430 |
1377 |
|
|
|
|
| 5 |
A' |
1286 |
1239 |
|
|
|
|
| 6 |
A' |
1119 |
1078 |
|
|
|
|
| 7 |
A' |
657 |
633 |
|
|
|
|
| 8 |
A" |
1044 |
1005 |
|
|
|
|
| 9 |
A" |
530 |
511 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7368.7 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 7095.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.384 |
0.000 |
| O2 |
-0.902 |
-0.624 |
0.000 |
| O3 |
1.183 |
0.194 |
0.000 |
| H4 |
-0.462 |
1.379 |
0.000 |
| H5 |
-1.785 |
-0.240 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3530 | 1.1978 | 1.0966 | 1.8907 |
O2 | 1.3530 | | 2.2392 | 2.0507 | 0.9632 | O3 | 1.1978 | 2.2392 | | 2.0272 | 2.9989 | H4 | 1.0966 | 2.0507 | 2.0272 | | 2.0900 | H5 | 1.8907 | 0.9632 | 2.9989 | 2.0900 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.253 |
|
O2 |
C1 |
O3 |
122.646 |
| O2 |
C1 |
H4 |
113.272 |
|
O3 |
C1 |
H4 |
124.082 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
