Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.770897 |
| Energy at 298.15K | -189.773630 |
| HF Energy | -189.770897 |
| Nuclear repulsion energy | 70.644967 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3778 |
3616 |
62.15 |
68.48 |
0.24 |
0.39 |
| 2 |
A' |
3073 |
2941 |
47.57 |
114.02 |
0.25 |
0.41 |
| 3 |
A' |
1864 |
1784 |
353.40 |
7.02 |
0.22 |
0.36 |
| 4 |
A' |
1409 |
1348 |
3.60 |
6.88 |
0.57 |
0.72 |
| 5 |
A' |
1318 |
1262 |
18.94 |
1.07 |
0.30 |
0.46 |
| 6 |
A' |
1151 |
1101 |
240.57 |
1.62 |
0.21 |
0.35 |
| 7 |
A' |
633 |
606 |
48.31 |
3.49 |
0.47 |
0.64 |
| 8 |
A" |
1065 |
1020 |
1.27 |
1.16 |
0.75 |
0.86 |
| 9 |
A" |
691 |
662 |
140.28 |
1.04 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7490.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 7169.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| O2 |
-1.023 |
-0.439 |
0.000 |
| O3 |
1.152 |
0.110 |
0.000 |
| H4 |
-0.378 |
1.446 |
0.000 |
| H5 |
-0.650 |
-1.329 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3350 | 1.1924 | 1.0946 | 1.8649 |
O2 | 1.3350 | | 2.2434 | 1.9920 | 0.9660 | O3 | 1.1924 | 2.2434 | | 2.0309 | 2.3061 | H4 | 1.0946 | 1.9920 | 2.0309 | | 2.7886 | H5 | 1.8649 | 0.9660 | 2.3061 | 2.7886 | |
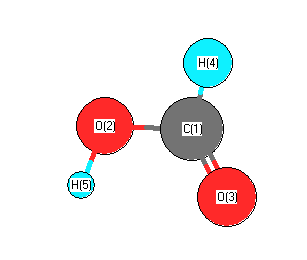 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
107.194 |
|
O2 |
C1 |
O3 |
125.053 |
| O2 |
C1 |
H4 |
109.753 |
|
O3 |
C1 |
H4 |
125.194 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.240 |
|
|
|
| 2 |
O |
-0.243 |
|
|
|
| 3 |
O |
-0.301 |
|
|
|
| 4 |
H |
0.075 |
|
|
|
| 5 |
H |
0.229 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.484 |
-0.333 |
0.000 |
1.521 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.683 |
-0.055 |
0.000 |
| y |
-0.055 |
-12.662 |
0.000 |
| z |
0.000 |
0.000 |
-16.767 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.968 |
-0.055 |
0.000 |
| y |
-0.055 |
6.563 |
0.000 |
| z |
0.000 |
0.000 |
0.406 |
|
| Polar |
|---|
| 3z2-r2 | 0.811 |
|---|
| x2-y2 | -9.021 |
|---|
| xy | -0.055 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.460 |
0.057 |
0.000 |
| y |
0.057 |
3.120 |
0.000 |
| z |
0.000 |
0.000 |
1.832 |
<r2> (average value of r
2) Å
2
| <r2> |
36.745 |
| (<r2>)1/2 |
6.062 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.763910 |
| Energy at 298.15K | |
| HF Energy | -189.763910 |
| Nuclear repulsion energy | 70.410030 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3851 |
3686 |
65.23 |
87.13 |
0.25 |
0.40 |
| 2 |
A' |
2985 |
2857 |
87.22 |
105.60 |
0.25 |
0.40 |
| 3 |
A' |
1908 |
1826 |
297.66 |
8.97 |
0.23 |
0.37 |
| 4 |
A' |
1426 |
1365 |
0.73 |
5.91 |
0.57 |
0.72 |
| 5 |
A' |
1287 |
1232 |
335.25 |
2.13 |
0.69 |
0.82 |
| 6 |
A' |
1136 |
1088 |
26.96 |
7.10 |
0.41 |
0.58 |
| 7 |
A' |
668 |
639 |
9.46 |
0.65 |
0.57 |
0.73 |
| 8 |
A" |
1046 |
1001 |
0.15 |
1.40 |
0.75 |
0.86 |
| 9 |
A" |
534 |
511 |
85.58 |
1.35 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7420.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 7101.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.381 |
0.000 |
| O2 |
-0.891 |
-0.622 |
0.000 |
| O3 |
1.172 |
0.197 |
0.000 |
| H4 |
-0.465 |
1.379 |
0.000 |
| H5 |
-1.782 |
-0.266 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3417 | 1.1859 | 1.1011 | 1.8962 |
O2 | 1.3417 | | 2.2192 | 2.0463 | 0.9604 | O3 | 1.1859 | 2.2192 | | 2.0187 | 2.9901 | H4 | 1.1011 | 2.0463 | 2.0187 | | 2.1074 | H5 | 1.8962 | 0.9604 | 2.9901 | 2.1074 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.796 |
|
O2 |
C1 |
O3 |
122.679 |
| O2 |
C1 |
H4 |
113.426 |
|
O3 |
C1 |
H4 |
123.895 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.230 |
|
|
|
| 2 |
O |
-0.199 |
|
|
|
| 3 |
O |
-0.271 |
|
|
|
| 4 |
H |
0.030 |
|
|
|
| 5 |
H |
0.211 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.591 |
1.473 |
0.000 |
3.881 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.526 |
-1.158 |
0.000 |
| y |
-1.158 |
-17.163 |
0.000 |
| z |
0.000 |
0.000 |
-16.763 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.437 |
-1.158 |
0.000 |
| y |
-1.158 |
-1.019 |
0.000 |
| z |
0.000 |
0.000 |
-0.418 |
|
| Polar |
|---|
| 3z2-r2 | -0.837 |
|---|
| x2-y2 | 1.637 |
|---|
| xy | -1.158 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.978 |
0.086 |
0.000 |
| y |
0.086 |
2.756 |
0.000 |
| z |
0.000 |
0.000 |
1.824 |
<r2> (average value of r
2) Å
2
| <r2> |
37.270 |
| (<r2>)1/2 |
6.105 |
