Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -268.331859 |
| Energy at 298.15K | -268.340160 |
| Nuclear repulsion energy | 195.810393 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3148 |
3013 |
23.33 |
|
|
|
| 2 |
A |
3111 |
2978 |
37.83 |
|
|
|
| 3 |
A |
3099 |
2966 |
43.72 |
|
|
|
| 4 |
A |
3055 |
2924 |
55.30 |
|
|
|
| 5 |
A |
3018 |
2889 |
50.61 |
|
|
|
| 6 |
A |
2966 |
2838 |
114.02 |
|
|
|
| 7 |
A |
1549 |
1482 |
2.24 |
|
|
|
| 8 |
A |
1528 |
1463 |
1.37 |
|
|
|
| 9 |
A |
1509 |
1444 |
2.55 |
|
|
|
| 10 |
A |
1432 |
1371 |
6.97 |
|
|
|
| 11 |
A |
1382 |
1323 |
0.68 |
|
|
|
| 12 |
A |
1353 |
1295 |
0.70 |
|
|
|
| 13 |
A |
1283 |
1227 |
7.61 |
|
|
|
| 14 |
A |
1245 |
1191 |
6.36 |
|
|
|
| 15 |
A |
1211 |
1159 |
20.66 |
|
|
|
| 16 |
A |
1199 |
1147 |
98.44 |
|
|
|
| 17 |
A |
1159 |
1109 |
32.39 |
|
|
|
| 18 |
A |
1136 |
1087 |
74.60 |
|
|
|
| 19 |
A |
1092 |
1045 |
36.28 |
|
|
|
| 20 |
A |
1015 |
972 |
30.84 |
|
|
|
| 21 |
A |
981 |
939 |
42.08 |
|
|
|
| 22 |
A |
964 |
922 |
31.38 |
|
|
|
| 23 |
A |
885 |
847 |
12.79 |
|
|
|
| 24 |
A |
742 |
710 |
0.38 |
|
|
|
| 25 |
A |
665 |
636 |
5.24 |
|
|
|
| 26 |
A |
272 |
260 |
9.12 |
|
|
|
| 27 |
A |
53 |
51 |
4.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20524.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 19643.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.166 |
-0.094 |
0.133 |
| C2 |
-0.829 |
0.843 |
0.171 |
| C3 |
-1.032 |
-0.645 |
-0.078 |
| O4 |
0.492 |
1.053 |
-0.282 |
| O5 |
0.282 |
-1.176 |
-0.032 |
| H6 |
2.044 |
-0.237 |
-0.493 |
| H7 |
1.457 |
-0.012 |
1.189 |
| H8 |
-0.911 |
1.082 |
1.236 |
| H9 |
-1.461 |
-0.837 |
-1.062 |
| H10 |
-1.655 |
-1.116 |
0.680 |
| H11 |
-1.498 |
1.483 |
-0.395 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 2.2040 | 2.2756 | 1.3935 | 1.4065 | 1.0877 | 1.0988 | 2.6292 | 2.9802 | 3.0500 | 3.1402 |
C2 | 2.2040 | | 1.5218 | 1.4116 | 2.3123 | 3.1398 | 2.6443 | 1.0944 | 2.1771 | 2.1864 | 1.0851 | C3 | 2.2756 | 1.5218 | | 2.2903 | 1.4175 | 3.1306 | 2.8639 | 2.1732 | 1.0902 | 1.0892 | 2.2010 | O4 | 1.3935 | 1.4116 | 2.2903 | | 2.2522 | 2.0291 | 2.0564 | 2.0663 | 2.8275 | 3.2000 | 2.0386 | O5 | 1.4065 | 2.3123 | 1.4175 | 2.2522 | | 2.0496 | 2.0553 | 2.8501 | 2.0530 | 2.0641 | 3.2196 | H6 | 1.0877 | 3.1398 | 3.1306 | 2.0291 | 2.0496 | | 1.7957 | 3.6686 | 3.6017 | 3.9792 | 3.9384 | H7 | 1.0988 | 2.6443 | 2.8639 | 2.0564 | 2.0553 | 1.7957 | | 2.6090 | 3.7768 | 3.3410 | 3.6708 | H8 | 2.6292 | 1.0944 | 2.1732 | 2.0663 | 2.8501 | 3.6686 | 2.6090 | | 3.0434 | 2.3863 | 1.7785 | H9 | 2.9802 | 2.1771 | 1.0902 | 2.8275 | 2.0530 | 3.6017 | 3.7768 | 3.0434 | | 1.7752 | 2.4140 | H10 | 3.0500 | 2.1864 | 1.0892 | 3.2000 | 2.0641 | 3.9792 | 3.3410 | 2.3863 | 1.7752 | | 2.8174 | H11 | 3.1402 | 1.0851 | 2.2010 | 2.0386 | 3.2196 | 3.9384 | 3.6708 | 1.7785 | 2.4140 | 2.8174 | |
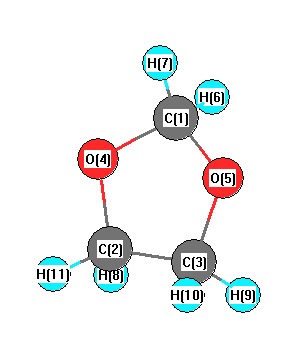 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
103.574 |
|
C1 |
O5 |
C3 |
107.380 |
| C2 |
C3 |
O5 |
103.694 |
|
C2 |
C3 |
H9 |
111.863 |
| C2 |
C3 |
H10 |
112.677 |
|
C3 |
C2 |
O4 |
102.598 |
| C3 |
C2 |
H8 |
111.283 |
|
C3 |
C2 |
H11 |
114.152 |
| O4 |
C1 |
O5 |
107.098 |
|
O4 |
C1 |
H6 |
109.109 |
| O4 |
C1 |
H7 |
110.640 |
|
O4 |
C2 |
H8 |
110.450 |
| O4 |
C2 |
H11 |
108.777 |
|
O5 |
C1 |
H6 |
109.865 |
| O5 |
C1 |
H7 |
109.642 |
|
O5 |
C3 |
H9 |
109.210 |
| O5 |
C3 |
H10 |
110.172 |
|
H6 |
C1 |
H7 |
110.426 |
| H8 |
C2 |
H11 |
109.372 |
|
H9 |
C3 |
H10 |
109.083 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.115 |
|
|
|
| 2 |
C |
-0.073 |
|
|
|
| 3 |
C |
-0.012 |
|
|
|
| 4 |
O |
-0.269 |
|
|
|
| 5 |
O |
-0.285 |
|
|
|
| 6 |
H |
0.100 |
|
|
|
| 7 |
H |
0.054 |
|
|
|
| 8 |
H |
0.093 |
|
|
|
| 9 |
H |
0.089 |
|
|
|
| 10 |
H |
0.083 |
|
|
|
| 11 |
H |
0.106 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.122 |
0.391 |
0.668 |
1.363 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.021 |
-0.834 |
0.392 |
| y |
-0.834 |
-33.941 |
0.625 |
| z |
0.392 |
0.625 |
-29.515 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.707 |
-0.834 |
0.392 |
| y |
-0.834 |
-6.173 |
0.625 |
| z |
0.392 |
0.625 |
0.465 |
|
| Polar |
|---|
| 3z2-r2 | 0.931 |
|---|
| x2-y2 | 7.920 |
|---|
| xy | -0.834 |
|---|
| xz | 0.392 |
|---|
| yz | 0.625 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.930 |
-0.082 |
0.077 |
| y |
-0.082 |
5.699 |
0.106 |
| z |
0.077 |
0.106 |
5.488 |
<r2> (average value of r
2) Å
2
| <r2> |
92.107 |
| (<r2>)1/2 |
9.597 |
