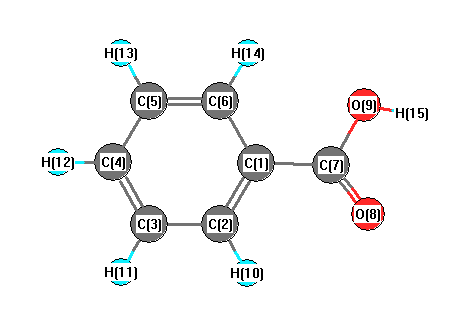
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3810 |
3647 |
98.05 |
|
|
|
| 2 |
A' |
3236 |
3098 |
3.07 |
|
|
|
| 3 |
A' |
3228 |
3090 |
7.23 |
|
|
|
| 4 |
A' |
3218 |
3080 |
10.72 |
|
|
|
| 5 |
A' |
3208 |
3071 |
9.70 |
|
|
|
| 6 |
A' |
3196 |
3059 |
0.33 |
|
|
|
| 7 |
A' |
1833 |
1755 |
338.11 |
|
|
|
| 8 |
A' |
1678 |
1606 |
17.31 |
|
|
|
| 9 |
A' |
1657 |
1586 |
4.31 |
|
|
|
| 10 |
A' |
1541 |
1475 |
2.99 |
|
|
|
| 11 |
A' |
1498 |
1434 |
18.88 |
|
|
|
| 12 |
A' |
1396 |
1336 |
155.64 |
|
|
|
| 13 |
A' |
1378 |
1319 |
4.75 |
|
|
|
| 14 |
A' |
1339 |
1282 |
3.13 |
|
|
|
| 15 |
A' |
1217 |
1165 |
113.87 |
|
|
|
| 16 |
A' |
1186 |
1135 |
108.34 |
|
|
|
| 17 |
A' |
1184 |
1133 |
0.68 |
|
|
|
| 18 |
A' |
1134 |
1085 |
74.92 |
|
|
|
| 19 |
A' |
1102 |
1055 |
34.84 |
|
|
|
| 20 |
A' |
1059 |
1014 |
11.63 |
|
|
|
| 21 |
A' |
1026 |
982 |
1.01 |
|
|
|
| 22 |
A' |
784 |
750 |
12.87 |
|
|
|
| 23 |
A' |
638 |
610 |
52.31 |
|
|
|
| 24 |
A' |
626 |
599 |
0.08 |
|
|
|
| 25 |
A' |
501 |
479 |
5.67 |
|
|
|
| 26 |
A' |
380 |
363 |
5.32 |
|
|
|
| 27 |
A' |
212 |
203 |
1.55 |
|
|
|
| 28 |
A" |
1030 |
986 |
0.06 |
|
|
|
| 29 |
A" |
1012 |
968 |
0.05 |
|
|
|
| 30 |
A" |
976 |
934 |
1.23 |
|
|
|
| 31 |
A" |
876 |
838 |
0.05 |
|
|
|
| 32 |
A" |
835 |
799 |
0.01 |
|
|
|
| 33 |
A" |
737 |
705 |
119.38 |
|
|
|
| 34 |
A" |
713 |
682 |
13.24 |
|
|
|
| 35 |
A" |
602 |
576 |
59.20 |
|
|
|
| 36 |
A" |
435 |
416 |
8.89 |
|
|
|
| 37 |
A" |
413 |
395 |
0.42 |
|
|
|
| 38 |
A" |
159 |
152 |
0.72 |
|
|
|
| 39 |
A" |
67 |
64 |
1.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25559.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 24462.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.013 |
|
|
|
| 2 |
C |
-0.111 |
|
|
|
| 3 |
C |
-0.137 |
|
|
|
| 4 |
C |
-0.114 |
|
|
|
| 5 |
C |
-0.130 |
|
|
|
| 6 |
C |
-0.134 |
|
|
|
| 7 |
C |
0.305 |
|
|
|
| 8 |
O |
-0.328 |
|
|
|
| 9 |
O |
-0.259 |
|
|
|
| 10 |
H |
0.146 |
|
|
|
| 11 |
H |
0.135 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
| 13 |
H |
0.133 |
|
|
|
| 14 |
H |
0.145 |
|
|
|
| 15 |
H |
0.225 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.253 |
-1.548 |
0.000 |
1.992 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.446 |
-5.839 |
0.000 |
| y |
-5.839 |
-45.142 |
0.000 |
| z |
0.000 |
0.000 |
-53.697 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.973 |
-5.839 |
0.000 |
| y |
-5.839 |
5.929 |
0.000 |
| z |
0.000 |
0.000 |
-6.902 |
|
| Polar |
|---|
| 3z2-r2 | -13.805 |
|---|
| x2-y2 | -3.304 |
|---|
| xy | -5.839 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.362 |
-0.224 |
0.000 |
| y |
-0.224 |
16.592 |
0.000 |
| z |
0.000 |
0.000 |
6.120 |
<r2> (average value of r
2) Å
2
| <r2> |
325.216 |
| (<r2>)1/2 |
18.034 |