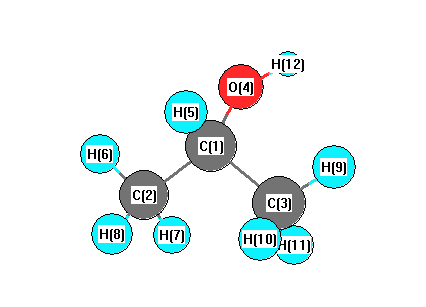
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3875 |
3709 |
19.79 |
|
|
|
| 2 |
A |
3148 |
3013 |
23.43 |
|
|
|
| 3 |
A |
3140 |
3006 |
46.93 |
|
|
|
| 4 |
A |
3133 |
2999 |
5.34 |
|
|
|
| 5 |
A |
3118 |
2984 |
32.66 |
|
|
|
| 6 |
A |
3062 |
2931 |
13.37 |
|
|
|
| 7 |
A |
3047 |
2916 |
21.97 |
|
|
|
| 8 |
A |
2991 |
2862 |
53.68 |
|
|
|
| 9 |
A |
1507 |
1442 |
6.24 |
|
|
|
| 10 |
A |
1495 |
1431 |
4.90 |
|
|
|
| 11 |
A |
1486 |
1422 |
2.77 |
|
|
|
| 12 |
A |
1480 |
1417 |
0.70 |
|
|
|
| 13 |
A |
1430 |
1369 |
24.11 |
|
|
|
| 14 |
A |
1408 |
1347 |
21.48 |
|
|
|
| 15 |
A |
1387 |
1328 |
2.30 |
|
|
|
| 16 |
A |
1375 |
1316 |
10.03 |
|
|
|
| 17 |
A |
1279 |
1224 |
50.46 |
|
|
|
| 18 |
A |
1193 |
1142 |
42.96 |
|
|
|
| 19 |
A |
1168 |
1118 |
18.74 |
|
|
|
| 20 |
A |
1095 |
1048 |
25.00 |
|
|
|
| 21 |
A |
981 |
939 |
38.98 |
|
|
|
| 22 |
A |
949 |
908 |
1.41 |
|
|
|
| 23 |
A |
924 |
884 |
0.35 |
|
|
|
| 24 |
A |
837 |
801 |
3.60 |
|
|
|
| 25 |
A |
472 |
452 |
6.36 |
|
|
|
| 26 |
A |
409 |
391 |
11.90 |
|
|
|
| 27 |
A |
351 |
336 |
2.88 |
|
|
|
| 28 |
A |
290 |
277 |
96.64 |
|
|
|
| 29 |
A |
258 |
247 |
4.70 |
|
|
|
| 30 |
A |
218 |
209 |
4.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23752.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22733.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.079 |
|
|
|
| 2 |
C |
-0.350 |
|
|
|
| 3 |
C |
-0.341 |
|
|
|
| 4 |
O |
-0.344 |
|
|
|
| 5 |
H |
0.082 |
|
|
|
| 6 |
H |
0.102 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.106 |
|
|
|
| 9 |
H |
0.123 |
|
|
|
| 10 |
H |
0.104 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
H |
0.203 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.110 |
-0.691 |
0.796 |
1.531 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.259 |
-2.569 |
-0.249 |
| y |
-2.569 |
-26.861 |
0.953 |
| z |
-0.249 |
0.953 |
-26.671 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.507 |
-2.569 |
-0.249 |
| y |
-2.569 |
-0.896 |
0.953 |
| z |
-0.249 |
0.953 |
-0.611 |
|
| Polar |
|---|
| 3z2-r2 | -1.222 |
|---|
| x2-y2 | 1.602 |
|---|
| xy | -2.569 |
|---|
| xz | -0.249 |
|---|
| yz | 0.953 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.783 |
-0.110 |
-0.040 |
| y |
-0.110 |
6.328 |
-0.001 |
| z |
-0.040 |
-0.001 |
5.762 |
<r2> (average value of r
2) Å
2
| <r2> |
88.179 |
| (<r2>)1/2 |
9.390 |