return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (dimethylformamide)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-248.110938
Energy at 298.15K-248.119376
HF Energy-247.082434
Nuclear repulsion energy182.642432
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3202 3040 0.31      
2 A' 3177 3016 10.78      
3 A' 3080 2924 45.92      
4 A' 3073 2917 49.41      
5 A' 3017 2864 81.02      
6 A' 1786 1696 435.10      
7 A' 1574 1494 23.21      
8 A' 1535 1457 13.33      
9 A' 1481 1406 4.94      
10 A' 1460 1386 59.51      
11 A' 1449 1376 28.04      
12 A' 1441 1368 23.39      
13 A' 1321 1255 30.75      
14 A' 1118 1061 105.77      
15 A' 1096 1040 2.99      
16 A' 898 852 2.00      
17 A' 670 636 6.15      
18 A' 390 370 1.44      
19 A' 321 305 10.98      
20 A" 3138 2980 11.42      
21 A" 3127 2968 39.12      
22 A" 1528 1451 14.24      
23 A" 1506 1430 4.50      
24 A" 1194 1133 3.10      
25 A" 1146 1088 0.16      
26 A" 1045 992 0.16      
27 A" 344 326 14.61      
28 A" 232 220 1.75      
29 A" 173 165 0.27      
30 A" 117 111 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 22818.0 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 21663.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.30252 0.14114 0.09982

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.691 -0.820 0.000
O2 0.209 -1.935 0.000
N3 0.000 0.344 0.000
C4 -1.442 0.344 0.000
C5 0.660 1.618 0.000
H6 1.774 -0.640 0.000
H7 -1.767 -0.687 0.000
H8 -1.824 0.845 0.885
H9 -1.824 0.845 -0.885
H10 1.733 1.463 0.000
H11 0.393 2.196 -0.882
H12 0.393 2.196 0.882

Atom - Atom Distances (Å)
  C1 O2 N3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.21381.35412.42932.43871.09812.46093.14373.14372.50993.15703.1570
O21.21382.28852.81343.58122.03092.33643.55603.55603.72364.22784.2278
N31.35412.28851.44171.43462.02862.04602.08872.08872.06232.08842.0884
C42.42932.81341.44172.45793.36251.08121.08691.08693.36582.75212.7521
C52.43873.58121.43462.45792.51783.34742.74842.74841.08371.08821.0882
H61.09812.03092.02863.36252.51783.54083.99203.99202.10343.27563.2756
H72.46092.33642.04601.08123.34743.54081.77081.77084.10733.70893.7089
H83.14373.55602.08871.08692.74843.99201.77081.77033.71723.14072.5961
H93.14373.55602.08871.08692.74843.99201.77081.77033.71722.59613.1407
H102.50993.72362.06233.36581.08372.10344.10733.71723.71721.76401.7640
H113.15704.22782.08842.75211.08823.27563.70893.14072.59611.76401.7646
H123.15704.22782.08842.75211.08823.27563.70892.59613.14071.76401.7646

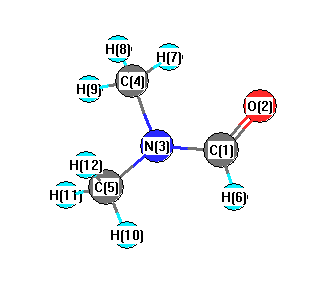
picture of dimethylformamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C4 120.638 C1 N3 C5 121.943
O2 C1 N3 125.959 O2 C1 H6 122.834
N3 C1 H6 111.207 N3 C4 H7 107.524
N3 C4 H8 110.607 N3 C4 H9 110.607
N3 C5 H10 109.168 N3 C5 H11 111.007
N3 C5 H12 111.007 C4 N3 C5 117.419
H7 C4 H8 109.518 H7 C4 H9 109.518
H8 C4 H9 109.044 H10 C5 H11 108.623
H10 C5 H12 108.623 H11 C5 H12 108.348
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability