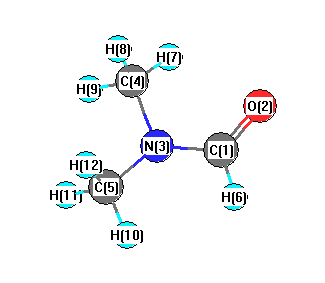
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3170 |
3041 |
0.87 |
|
|
|
| 2 |
A' |
3135 |
3008 |
14.28 |
|
|
|
| 3 |
A' |
3037 |
2913 |
54.28 |
|
|
|
| 4 |
A' |
3028 |
2904 |
46.48 |
|
|
|
| 5 |
A' |
2959 |
2838 |
88.06 |
|
|
|
| 6 |
A' |
1802 |
1728 |
461.18 |
|
|
|
| 7 |
A' |
1549 |
1486 |
32.92 |
|
|
|
| 8 |
A' |
1506 |
1445 |
13.21 |
|
|
|
| 9 |
A' |
1470 |
1410 |
4.92 |
|
|
|
| 10 |
A' |
1447 |
1388 |
37.24 |
|
|
|
| 11 |
A' |
1434 |
1376 |
13.11 |
|
|
|
| 12 |
A' |
1421 |
1363 |
49.64 |
|
|
|
| 13 |
A' |
1305 |
1252 |
31.04 |
|
|
|
| 14 |
A' |
1108 |
1063 |
109.63 |
|
|
|
| 15 |
A' |
1086 |
1042 |
2.76 |
|
|
|
| 16 |
A' |
891 |
855 |
1.97 |
|
|
|
| 17 |
A' |
670 |
643 |
7.11 |
|
|
|
| 18 |
A' |
390 |
374 |
1.66 |
|
|
|
| 19 |
A' |
321 |
308 |
12.06 |
|
|
|
| 20 |
A" |
3089 |
2963 |
18.53 |
|
|
|
| 21 |
A" |
3078 |
2953 |
43.56 |
|
|
|
| 22 |
A" |
1500 |
1439 |
15.02 |
|
|
|
| 23 |
A" |
1478 |
1418 |
4.28 |
|
|
|
| 24 |
A" |
1181 |
1132 |
2.00 |
|
|
|
| 25 |
A" |
1132 |
1086 |
0.13 |
|
|
|
| 26 |
A" |
1030 |
988 |
0.02 |
|
|
|
| 27 |
A" |
344 |
330 |
15.87 |
|
|
|
| 28 |
A" |
232 |
222 |
1.17 |
|
|
|
| 29 |
A" |
176 |
169 |
0.07 |
|
|
|
| 30 |
A" |
130 |
125 |
0.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22550.0 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 21629.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.145 |
|
|
|
| 2 |
O |
-0.353 |
|
|
|
| 3 |
N |
-0.036 |
|
|
|
| 4 |
C |
-0.177 |
|
|
|
| 5 |
C |
-0.150 |
|
|
|
| 6 |
H |
0.035 |
|
|
|
| 7 |
H |
0.117 |
|
|
|
| 8 |
H |
0.083 |
|
|
|
| 9 |
H |
0.083 |
|
|
|
| 10 |
H |
0.087 |
|
|
|
| 11 |
H |
0.083 |
|
|
|
| 12 |
H |
0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.186 |
3.946 |
0.000 |
3.950 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.517 |
0.546 |
0.000 |
| y |
0.546 |
-36.684 |
0.000 |
| z |
0.000 |
0.000 |
-30.837 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.244 |
0.546 |
0.000 |
| y |
0.546 |
-7.507 |
0.000 |
| z |
0.000 |
0.000 |
1.263 |
|
| Polar |
|---|
| 3z2-r2 | 2.527 |
|---|
| x2-y2 | 9.168 |
|---|
| xy | 0.546 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.420 |
-0.163 |
0.000 |
| y |
-0.163 |
8.615 |
0.000 |
| z |
0.000 |
0.000 |
5.261 |
<r2> (average value of r
2) Å
2
| <r2> |
122.793 |
| (<r2>)1/2 |
11.081 |