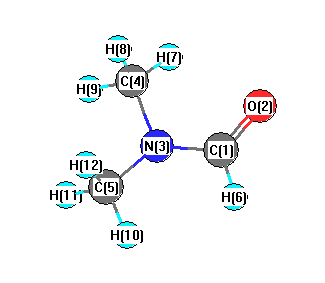
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3164 |
3028 |
0.91 |
|
|
|
| 2 |
A' |
3129 |
2995 |
14.81 |
|
|
|
| 3 |
A' |
3031 |
2901 |
56.91 |
|
|
|
| 4 |
A' |
3023 |
2893 |
47.47 |
|
|
|
| 5 |
A' |
2947 |
2820 |
91.26 |
|
|
|
| 6 |
A' |
1796 |
1719 |
457.81 |
|
|
|
| 7 |
A' |
1542 |
1476 |
30.84 |
|
|
|
| 8 |
A' |
1500 |
1435 |
13.88 |
|
|
|
| 9 |
A' |
1459 |
1397 |
3.57 |
|
|
|
| 10 |
A' |
1438 |
1376 |
44.17 |
|
|
|
| 11 |
A' |
1426 |
1364 |
10.38 |
|
|
|
| 12 |
A' |
1410 |
1350 |
48.38 |
|
|
|
| 13 |
A' |
1299 |
1243 |
30.71 |
|
|
|
| 14 |
A' |
1101 |
1054 |
107.96 |
|
|
|
| 15 |
A' |
1079 |
1032 |
2.79 |
|
|
|
| 16 |
A' |
886 |
848 |
1.84 |
|
|
|
| 17 |
A' |
665 |
637 |
7.18 |
|
|
|
| 18 |
A' |
380 |
363 |
1.61 |
|
|
|
| 19 |
A' |
320 |
306 |
11.88 |
|
|
|
| 20 |
A" |
3083 |
2951 |
15.98 |
|
|
|
| 21 |
A" |
3073 |
2941 |
48.76 |
|
|
|
| 22 |
A" |
1493 |
1429 |
14.12 |
|
|
|
| 23 |
A" |
1472 |
1409 |
4.31 |
|
|
|
| 24 |
A" |
1173 |
1123 |
2.31 |
|
|
|
| 25 |
A" |
1125 |
1077 |
0.13 |
|
|
|
| 26 |
A" |
1023 |
979 |
0.02 |
|
|
|
| 27 |
A" |
343 |
329 |
15.01 |
|
|
|
| 28 |
A" |
229 |
219 |
1.21 |
|
|
|
| 29 |
A" |
177 |
170 |
0.09 |
|
|
|
| 30 |
A" |
131 |
125 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22457.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 21494.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.112 |
|
|
|
| 2 |
O |
-0.352 |
|
|
|
| 3 |
N |
-0.005 |
|
|
|
| 4 |
C |
-0.265 |
|
|
|
| 5 |
C |
-0.237 |
|
|
|
| 6 |
H |
0.056 |
|
|
|
| 7 |
H |
0.141 |
|
|
|
| 8 |
H |
0.110 |
|
|
|
| 9 |
H |
0.110 |
|
|
|
| 10 |
H |
0.111 |
|
|
|
| 11 |
H |
0.109 |
|
|
|
| 12 |
H |
0.109 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.175 |
3.920 |
0.000 |
3.924 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.602 |
0.545 |
0.000 |
| y |
0.545 |
-36.676 |
0.000 |
| z |
0.000 |
0.000 |
-30.881 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.176 |
0.545 |
0.000 |
| y |
0.545 |
-7.435 |
0.000 |
| z |
0.000 |
0.000 |
1.259 |
|
| Polar |
|---|
| 3z2-r2 | 2.517 |
|---|
| x2-y2 | 9.074 |
|---|
| xy | 0.545 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.481 |
-0.168 |
0.000 |
| y |
-0.168 |
8.678 |
0.000 |
| z |
0.000 |
0.000 |
5.291 |
<r2> (average value of r
2) Å
2
| <r2> |
122.715 |
| (<r2>)1/2 |
11.078 |