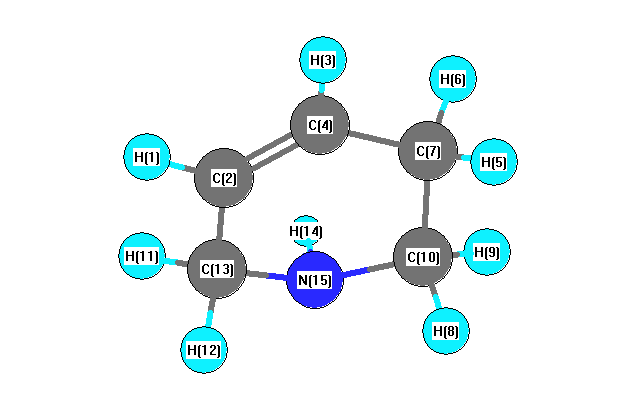
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3547 |
3394 |
0.19 |
|
|
|
| 2 |
A |
3184 |
3047 |
35.02 |
|
|
|
| 3 |
A |
3159 |
3024 |
10.03 |
|
|
|
| 4 |
A |
3101 |
2968 |
39.61 |
|
|
|
| 5 |
A |
3074 |
2942 |
24.84 |
|
|
|
| 6 |
A |
3065 |
2934 |
53.33 |
|
|
|
| 7 |
A |
3047 |
2916 |
21.28 |
|
|
|
| 8 |
A |
3022 |
2892 |
32.85 |
|
|
|
| 9 |
A |
3008 |
2879 |
44.72 |
|
|
|
| 10 |
A |
1742 |
1667 |
2.38 |
|
|
|
| 11 |
A |
1495 |
1431 |
8.47 |
|
|
|
| 12 |
A |
1488 |
1424 |
2.30 |
|
|
|
| 13 |
A |
1474 |
1411 |
8.61 |
|
|
|
| 14 |
A |
1470 |
1407 |
2.95 |
|
|
|
| 15 |
A |
1425 |
1364 |
0.58 |
|
|
|
| 16 |
A |
1396 |
1336 |
4.70 |
|
|
|
| 17 |
A |
1371 |
1312 |
5.16 |
|
|
|
| 18 |
A |
1353 |
1295 |
0.40 |
|
|
|
| 19 |
A |
1330 |
1273 |
0.18 |
|
|
|
| 20 |
A |
1262 |
1208 |
4.18 |
|
|
|
| 21 |
A |
1223 |
1170 |
16.31 |
|
|
|
| 22 |
A |
1196 |
1144 |
4.98 |
|
|
|
| 23 |
A |
1145 |
1096 |
14.26 |
|
|
|
| 24 |
A |
1109 |
1062 |
2.13 |
|
|
|
| 25 |
A |
1061 |
1016 |
5.22 |
|
|
|
| 26 |
A |
1013 |
970 |
3.02 |
|
|
|
| 27 |
A |
1011 |
968 |
1.54 |
|
|
|
| 28 |
A |
980 |
938 |
4.36 |
|
|
|
| 29 |
A |
932 |
892 |
5.44 |
|
|
|
| 30 |
A |
912 |
873 |
12.26 |
|
|
|
| 31 |
A |
867 |
829 |
2.71 |
|
|
|
| 32 |
A |
804 |
769 |
82.63 |
|
|
|
| 33 |
A |
753 |
721 |
44.96 |
|
|
|
| 34 |
A |
653 |
625 |
23.33 |
|
|
|
| 35 |
A |
524 |
501 |
1.23 |
|
|
|
| 36 |
A |
478 |
458 |
0.43 |
|
|
|
| 37 |
A |
400 |
383 |
0.85 |
|
|
|
| 38 |
A |
281 |
269 |
7.91 |
|
|
|
| 39 |
A |
167 |
160 |
2.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29759.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 28482.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.132 |
|
|
|
| 2 |
C |
-0.178 |
|
|
|
| 3 |
H |
0.133 |
|
|
|
| 4 |
C |
-0.155 |
|
|
|
| 5 |
H |
0.107 |
|
|
|
| 6 |
H |
0.114 |
|
|
|
| 7 |
C |
-0.204 |
|
|
|
| 8 |
H |
0.115 |
|
|
|
| 9 |
H |
0.110 |
|
|
|
| 10 |
C |
-0.159 |
|
|
|
| 11 |
H |
0.109 |
|
|
|
| 12 |
H |
0.116 |
|
|
|
| 13 |
C |
-0.128 |
|
|
|
| 14 |
H |
0.118 |
|
|
|
| 15 |
N |
-0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.475 |
0.627 |
-0.469 |
0.916 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.583 |
-1.858 |
0.221 |
| y |
-1.858 |
-39.871 |
0.883 |
| z |
0.221 |
0.883 |
-36.746 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.725 |
-1.858 |
0.221 |
| y |
-1.858 |
-3.706 |
0.883 |
| z |
0.221 |
0.883 |
0.981 |
|
| Polar |
|---|
| 3z2-r2 | 1.962 |
|---|
| x2-y2 | 4.287 |
|---|
| xy | -1.858 |
|---|
| xz | 0.221 |
|---|
| yz | 0.883 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.173 |
-0.074 |
-0.234 |
| y |
-0.074 |
9.238 |
0.022 |
| z |
-0.234 |
0.022 |
7.394 |
<r2> (average value of r
2) Å
2
| <r2> |
145.186 |
| (<r2>)1/2 |
12.049 |