Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -194.321978 |
| Energy at 298.15K | -194.330846 |
| Nuclear repulsion energy | 131.068908 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3871 |
3705 |
28.64 |
|
|
|
| 2 |
A' |
3123 |
2989 |
31.07 |
|
|
|
| 3 |
A' |
3054 |
2923 |
32.38 |
|
|
|
| 4 |
A' |
3043 |
2912 |
24.14 |
|
|
|
| 5 |
A' |
2979 |
2851 |
60.73 |
|
|
|
| 6 |
A' |
1522 |
1457 |
4.00 |
|
|
|
| 7 |
A' |
1503 |
1438 |
4.75 |
|
|
|
| 8 |
A' |
1488 |
1424 |
0.90 |
|
|
|
| 9 |
A' |
1451 |
1389 |
2.66 |
|
|
|
| 10 |
A' |
1405 |
1345 |
1.52 |
|
|
|
| 11 |
A' |
1342 |
1284 |
8.55 |
|
|
|
| 12 |
A' |
1254 |
1200 |
47.36 |
|
|
|
| 13 |
A' |
1107 |
1059 |
25.49 |
|
|
|
| 14 |
A' |
1075 |
1029 |
73.41 |
|
|
|
| 15 |
A' |
1054 |
1009 |
12.52 |
|
|
|
| 16 |
A' |
893 |
854 |
4.89 |
|
|
|
| 17 |
A' |
454 |
435 |
10.35 |
|
|
|
| 18 |
A' |
262 |
250 |
3.42 |
|
|
|
| 19 |
A" |
3114 |
2980 |
63.38 |
|
|
|
| 20 |
A" |
3088 |
2955 |
3.07 |
|
|
|
| 21 |
A" |
3008 |
2879 |
48.19 |
|
|
|
| 22 |
A" |
1494 |
1430 |
7.48 |
|
|
|
| 23 |
A" |
1315 |
1258 |
0.13 |
|
|
|
| 24 |
A" |
1261 |
1207 |
0.04 |
|
|
|
| 25 |
A" |
1180 |
1130 |
1.11 |
|
|
|
| 26 |
A" |
895 |
856 |
2.11 |
|
|
|
| 27 |
A" |
759 |
727 |
1.14 |
|
|
|
| 28 |
A" |
262 |
251 |
106.90 |
|
|
|
| 29 |
A" |
225 |
215 |
3.32 |
|
|
|
| 30 |
A" |
124 |
119 |
4.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23801.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22780.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.440 |
1.220 |
0.000 |
| C2 |
0.000 |
0.739 |
0.000 |
| C3 |
0.098 |
-0.768 |
0.000 |
| O4 |
1.467 |
-1.125 |
0.000 |
| H5 |
-1.493 |
2.307 |
0.000 |
| H6 |
-1.977 |
0.864 |
0.880 |
| H7 |
-1.977 |
0.864 |
-0.880 |
| H8 |
0.529 |
1.119 |
0.875 |
| H9 |
0.529 |
1.119 |
-0.875 |
| H10 |
-0.413 |
-1.169 |
0.884 |
| H11 |
-0.413 |
-1.169 |
-0.884 |
| H12 |
1.534 |
-2.081 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5185 | 2.5133 | 3.7353 | 1.0883 | 1.0903 | 1.0903 | 2.1571 | 2.1571 | 2.7465 | 2.7465 | 4.4432 |
C2 | 1.5185 | | 1.5098 | 2.3722 | 2.1650 | 2.1678 | 2.1678 | 1.0906 | 1.0906 | 2.1426 | 2.1426 | 3.2099 | C3 | 2.5133 | 1.5098 | | 1.4152 | 3.4617 | 2.7826 | 2.7826 | 2.1240 | 2.1240 | 1.0964 | 1.0964 | 1.9461 | O4 | 3.7353 | 2.3722 | 1.4152 | | 4.5322 | 4.0737 | 4.0737 | 2.5851 | 2.5851 | 2.0775 | 2.0775 | 0.9578 | H5 | 1.0883 | 2.1650 | 3.4617 | 4.5322 | | 1.7579 | 1.7579 | 2.5027 | 2.5027 | 3.7456 | 3.7456 | 5.3305 | H6 | 1.0903 | 2.1678 | 2.7826 | 4.0737 | 1.7579 | | 1.7594 | 2.5192 | 3.0699 | 2.5655 | 3.1130 | 4.6665 | H7 | 1.0903 | 2.1678 | 2.7826 | 4.0737 | 1.7579 | 1.7594 | | 3.0699 | 2.5192 | 3.1130 | 2.5655 | 4.6665 | H8 | 2.1571 | 1.0906 | 2.1240 | 2.5851 | 2.5027 | 2.5192 | 3.0699 | | 1.7494 | 2.4743 | 3.0354 | 3.4662 | H9 | 2.1571 | 1.0906 | 2.1240 | 2.5851 | 2.5027 | 3.0699 | 2.5192 | 1.7494 | | 3.0354 | 2.4743 | 3.4662 | H10 | 2.7465 | 2.1426 | 1.0964 | 2.0775 | 3.7456 | 2.5655 | 3.1130 | 2.4743 | 3.0354 | | 1.7671 | 2.3243 | H11 | 2.7465 | 2.1426 | 1.0964 | 2.0775 | 3.7456 | 3.1130 | 2.5655 | 3.0354 | 2.4743 | 1.7671 | | 2.3243 | H12 | 4.4432 | 3.2099 | 1.9461 | 0.9578 | 5.3305 | 4.6665 | 4.6665 | 3.4662 | 3.4662 | 2.3243 | 2.3243 | |
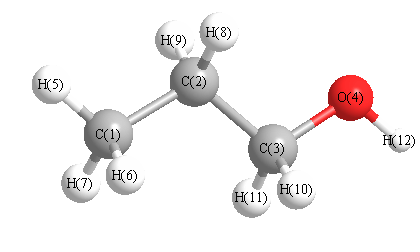 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.180 |
|
C1 |
C2 |
H8 |
110.460 |
| C1 |
C2 |
H9 |
110.460 |
|
C2 |
C1 |
H5 |
111.231 |
| C2 |
C1 |
H6 |
111.333 |
|
C2 |
C1 |
H7 |
111.333 |
| C2 |
C3 |
O4 |
108.347 |
|
C2 |
C3 |
H10 |
109.573 |
| C2 |
C3 |
H11 |
109.573 |
|
C3 |
C2 |
H8 |
108.453 |
| C3 |
C2 |
H9 |
108.453 |
|
C3 |
O4 |
H12 |
108.654 |
| O4 |
C3 |
H10 |
110.978 |
|
O4 |
C3 |
H11 |
110.978 |
| H5 |
C1 |
H6 |
107.586 |
|
H5 |
C1 |
H7 |
107.586 |
| H6 |
C1 |
H7 |
107.578 |
|
H8 |
C2 |
H9 |
106.647 |
| H10 |
C3 |
H11 |
107.380 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.368 |
|
|
|
| 2 |
C |
-0.201 |
|
|
|
| 3 |
C |
-0.012 |
|
|
|
| 4 |
O |
-0.333 |
|
|
|
| 5 |
H |
0.122 |
|
|
|
| 6 |
H |
0.109 |
|
|
|
| 7 |
H |
0.109 |
|
|
|
| 8 |
H |
0.117 |
|
|
|
| 9 |
H |
0.117 |
|
|
|
| 10 |
H |
0.068 |
|
|
|
| 11 |
H |
0.068 |
|
|
|
| 12 |
H |
0.203 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.059 |
-0.934 |
0.000 |
1.412 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.351 |
-0.984 |
0.000 |
| y |
-0.984 |
-21.979 |
0.000 |
| z |
0.000 |
0.000 |
-26.543 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.090 |
-0.984 |
0.000 |
| y |
-0.984 |
5.968 |
0.000 |
| z |
0.000 |
0.000 |
-0.878 |
|
| Polar |
|---|
| 3z2-r2 | -1.756 |
|---|
| x2-y2 | -7.372 |
|---|
| xy | -0.984 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.598 |
-0.600 |
0.000 |
| y |
-0.600 |
6.713 |
0.000 |
| z |
0.000 |
0.000 |
5.595 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -194.320879 |
| Energy at 298.15K | |
| HF Energy | -194.320879 |
| Nuclear repulsion energy | 133.482834 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3894 |
3727 |
33.01 |
106.25 |
0.27 |
0.42 |
| 2 |
A |
3145 |
3010 |
23.74 |
37.66 |
0.75 |
0.86 |
| 3 |
A |
3124 |
2990 |
46.19 |
57.23 |
0.56 |
0.72 |
| 4 |
A |
3094 |
2961 |
18.61 |
92.07 |
0.70 |
0.82 |
| 5 |
A |
3059 |
2928 |
49.47 |
123.53 |
0.06 |
0.12 |
| 6 |
A |
3055 |
2924 |
24.91 |
153.33 |
0.02 |
0.04 |
| 7 |
A |
3027 |
2898 |
42.90 |
94.43 |
0.75 |
0.86 |
| 8 |
A |
2994 |
2866 |
63.98 |
126.74 |
0.09 |
0.17 |
| 9 |
A |
1522 |
1457 |
2.72 |
4.93 |
0.67 |
0.80 |
| 10 |
A |
1509 |
1444 |
7.48 |
3.87 |
0.75 |
0.86 |
| 11 |
A |
1494 |
1430 |
7.16 |
10.06 |
0.75 |
0.86 |
| 12 |
A |
1476 |
1412 |
2.82 |
9.83 |
0.75 |
0.86 |
| 13 |
A |
1453 |
1391 |
3.86 |
1.59 |
0.14 |
0.25 |
| 14 |
A |
1409 |
1348 |
4.07 |
0.46 |
0.73 |
0.84 |
| 15 |
A |
1378 |
1318 |
0.70 |
0.97 |
0.75 |
0.86 |
| 16 |
A |
1325 |
1268 |
19.69 |
7.59 |
0.73 |
0.84 |
| 17 |
A |
1274 |
1219 |
1.22 |
5.08 |
0.73 |
0.85 |
| 18 |
A |
1246 |
1192 |
37.25 |
2.23 |
0.44 |
0.61 |
| 19 |
A |
1162 |
1113 |
7.43 |
0.84 |
0.09 |
0.16 |
| 20 |
A |
1135 |
1087 |
21.00 |
3.71 |
0.56 |
0.72 |
| 21 |
A |
1080 |
1033 |
36.09 |
2.56 |
0.71 |
0.83 |
| 22 |
A |
997 |
955 |
42.91 |
3.32 |
0.75 |
0.86 |
| 23 |
A |
925 |
885 |
1.97 |
0.21 |
0.54 |
0.70 |
| 24 |
A |
882 |
844 |
2.05 |
10.11 |
0.12 |
0.22 |
| 25 |
A |
772 |
739 |
0.62 |
0.34 |
0.55 |
0.71 |
| 26 |
A |
473 |
453 |
7.60 |
0.19 |
0.45 |
0.62 |
| 27 |
A |
319 |
306 |
6.57 |
0.25 |
0.43 |
0.60 |
| 28 |
A |
243 |
233 |
101.19 |
2.95 |
0.74 |
0.85 |
| 29 |
A |
222 |
212 |
1.89 |
0.05 |
0.69 |
0.82 |
| 30 |
A |
144 |
138 |
8.31 |
0.16 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 23914.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22889.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.526 |
-0.514 |
0.128 |
| C2 |
-0.630 |
0.639 |
-0.288 |
| C3 |
0.760 |
0.540 |
0.293 |
| O4 |
1.372 |
-0.628 |
-0.219 |
| H5 |
-2.510 |
-0.436 |
-0.330 |
| H6 |
-1.663 |
-0.531 |
1.209 |
| H7 |
-1.087 |
-1.463 |
-0.167 |
| H8 |
-0.544 |
0.673 |
-1.375 |
| H9 |
-1.064 |
1.589 |
0.024 |
| H10 |
0.699 |
0.499 |
1.386 |
| H11 |
1.338 |
1.430 |
0.025 |
| H12 |
2.230 |
-0.732 |
0.191 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5176 | 2.5229 | 2.9214 | 1.0880 | 1.0899 | 1.0869 | 2.1517 | 2.1557 | 2.7497 | 3.4630 | 3.7628 |
C2 | 1.5176 | | 1.5103 | 2.3706 | 2.1654 | 2.1622 | 2.1544 | 1.0904 | 1.0907 | 2.1426 | 2.1447 | 3.2077 | C3 | 2.5229 | 1.5103 | | 1.4148 | 3.4691 | 2.8032 | 2.7639 | 2.1213 | 2.1216 | 1.0957 | 1.0943 | 1.9465 | O4 | 2.9214 | 2.3706 | 1.4148 | | 3.8886 | 3.3561 | 2.5983 | 2.5886 | 3.3034 | 2.0736 | 2.0724 | 0.9562 | H5 | 1.0880 | 2.1654 | 3.4691 | 3.8886 | | 1.7593 | 1.7624 | 2.4869 | 2.5134 | 3.7575 | 4.2913 | 4.7775 | H6 | 1.0899 | 2.1622 | 2.8032 | 3.3561 | 1.7593 | | 1.7592 | 3.0620 | 2.5014 | 2.5830 | 3.7754 | 4.0288 | H7 | 1.0869 | 2.1544 | 2.7639 | 2.5983 | 1.7624 | 1.7592 | | 2.5135 | 3.0588 | 3.0748 | 3.7803 | 3.4156 | H8 | 2.1517 | 1.0904 | 2.1213 | 2.5886 | 2.4869 | 3.0620 | 2.5135 | | 1.7511 | 3.0329 | 2.4647 | 3.4816 | H9 | 2.1557 | 1.0907 | 2.1216 | 3.3034 | 2.5134 | 2.5014 | 3.0588 | 1.7511 | | 2.4807 | 2.4077 | 4.0333 | H10 | 2.7497 | 2.1426 | 1.0957 | 2.0736 | 3.7575 | 2.5830 | 3.0748 | 3.0329 | 2.4807 | | 1.7686 | 2.2991 | H11 | 3.4630 | 2.1447 | 1.0943 | 2.0724 | 4.2913 | 3.7754 | 3.7803 | 2.4647 | 2.4077 | 1.7686 | | 2.3444 | H12 | 3.7628 | 3.2077 | 1.9465 | 0.9562 | 4.7775 | 4.0288 | 3.4156 | 3.4816 | 4.0333 | 2.2991 | 2.3444 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.861 |
|
C1 |
C2 |
H8 |
110.109 |
| C1 |
C2 |
H9 |
110.405 |
|
C2 |
C1 |
H5 |
111.344 |
| C2 |
C1 |
H6 |
110.972 |
|
C2 |
C1 |
H7 |
110.536 |
| C2 |
C3 |
O4 |
108.237 |
|
C2 |
C3 |
H10 |
109.582 |
| C2 |
C3 |
H11 |
109.828 |
|
C3 |
C2 |
H8 |
108.217 |
| C3 |
C2 |
H9 |
108.228 |
|
C3 |
O4 |
H12 |
108.821 |
| O4 |
C3 |
H10 |
110.737 |
|
O4 |
C3 |
H11 |
110.728 |
| H5 |
C1 |
H6 |
107.763 |
|
H5 |
C1 |
H7 |
108.258 |
| H6 |
C1 |
H7 |
107.834 |
|
H8 |
C2 |
H9 |
106.806 |
| H10 |
C3 |
H11 |
107.724 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.356 |
|
|
|
| 2 |
C |
-0.205 |
|
|
|
| 3 |
C |
-0.030 |
|
|
|
| 4 |
O |
-0.340 |
|
|
|
| 5 |
H |
0.115 |
|
|
|
| 6 |
H |
0.104 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
H |
0.116 |
|
|
|
| 9 |
H |
0.108 |
|
|
|
| 10 |
H |
0.072 |
|
|
|
| 11 |
H |
0.082 |
|
|
|
| 12 |
H |
0.207 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.704 |
0.851 |
0.958 |
1.462 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.951 |
-0.320 |
1.965 |
| y |
-0.320 |
-27.032 |
-0.561 |
| z |
1.965 |
-0.561 |
-26.722 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.926 |
-0.320 |
1.965 |
| y |
-0.320 |
-2.196 |
-0.561 |
| z |
1.965 |
-0.561 |
-1.730 |
|
| Polar |
|---|
| 3z2-r2 | -3.460 |
|---|
| x2-y2 | 4.082 |
|---|
| xy | -0.320 |
|---|
| xz | 1.965 |
|---|
| yz | -0.561 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.866 |
0.099 |
0.137 |
| y |
0.099 |
6.128 |
-0.010 |
| z |
0.137 |
-0.010 |
5.765 |
<r2> (average value of r
2) Å
2
| <r2> |
94.090 |
| (<r2>)1/2 |
9.700 |
