return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (1-Butanol)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-233.265889
Energy at 298.15K-233.277114
HF Energy-232.238184
Nuclear repulsion energy186.414855
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3884 3687 35.87      
2 A' 3166 3006 28.03      
3 A' 3096 2940 45.03      
4 A' 3089 2933 20.96      
5 A' 3081 2925 12.30      
6 A' 3048 2894 45.54      
7 A' 1552 1474 2.78      
8 A' 1534 1456 5.85      
9 A' 1520 1443 1.22      
10 A' 1509 1433 0.08      
11 A' 1476 1401 5.32      
12 A' 1425 1353 3.32      
13 A' 1405 1333 2.42      
14 A' 1320 1253 20.55      
15 A' 1262 1198 30.04      
16 A' 1140 1082 4.31      
17 A' 1103 1047 9.14      
18 A' 1091 1036 83.75      
19 A' 1037 985 0.30      
20 A' 928 881 11.86      
21 A' 444 422 12.40      
22 A' 399 379 0.11      
23 A' 183 174 2.33      
24 A" 3161 3001 42.45      
25 A" 3138 2979 32.04      
26 A" 3105 2948 2.44      
27 A" 3076 2921 32.29      
28 A" 1528 1451 7.52      
29 A" 1343 1275 0.01      
30 A" 1338 1270 0.77      
31 A" 1267 1203 0.31      
32 A" 1204 1143 1.62      
33 A" 968 919 0.04      
34 A" 824 782 0.77      
35 A" 758 720 1.92      
36 A" 274 260 83.83      
37 A" 264 251 24.06      
38 A" 121 115 2.38      
39 A" 109 104 4.50      

Unscaled Zero Point Vibrational Energy (zpe) 30585.4 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 29037.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.63299 0.06720 0.06359

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.336 -0.355 0.000
C2 0.000 0.338 0.000
C3 -1.160 -0.637 0.000
C4 -2.503 0.069 0.000
O5 2.346 0.642 0.000
H6 1.416 -0.993 0.881
H7 1.416 -0.993 -0.881
H8 -0.056 0.983 0.874
H9 -0.056 0.983 -0.874
H10 -1.089 -1.284 0.872
H11 -1.089 -1.284 -0.872
H12 -3.327 -0.638 0.000
H13 -2.605 0.702 0.877
H14 -2.605 0.702 -0.877
H15 3.197 0.199 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.50512.51223.86271.41881.09071.09072.11942.11942.73892.73894.67144.17294.17291.9414
C21.50511.51572.51802.36522.13372.13371.08751.08752.13942.13943.46722.77212.77213.1996
C32.51221.51571.51693.73192.74602.74602.14632.14631.08831.08832.16642.15532.15534.4364
C43.86272.51801.51694.88294.15504.15502.75492.75492.14272.14271.08521.08621.08625.7015
O51.41882.36523.73194.88292.07672.07672.57862.57864.03274.03275.81525.02775.02770.9591
H61.09072.13372.74604.15502.07671.76252.46373.02512.52133.07094.83684.36284.70372.3169
H71.09072.13372.74604.15502.07671.76253.02512.46373.07092.52134.83684.70374.36282.3169
H82.11941.08752.14632.75492.57862.46373.02511.74822.49093.04193.75372.56393.10483.4581
H92.11941.08752.14632.75492.57863.02512.46371.74823.04192.49093.75373.10482.56393.4581
H102.73892.13941.08832.14274.03272.52133.07092.49093.04191.74392.48722.49803.04934.6176
H112.73892.13941.08832.14274.03273.07092.52133.04192.49091.74392.48723.04932.49804.6176
H124.67143.46722.16641.08525.81524.83684.83683.75373.75372.48722.48721.75671.75676.5769
H134.17292.77212.15531.08625.02774.36284.70372.56393.10482.49803.04931.75671.75395.8885
H144.17292.77212.15531.08625.02774.70374.36283.10482.56393.04932.49801.75671.75395.8885
H151.94143.19964.43645.70150.95912.31692.31693.45813.45814.61764.61766.57695.88855.8885

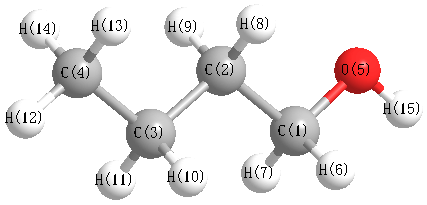
picture of 1-Butanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.535 C1 C2 H8 108.593
C1 C2 H9 108.593 C1 O5 H15 107.896
C2 C1 O5 107.946 C2 C1 H6 109.533
C2 C1 H7 109.533 C2 C3 C4 112.261
C2 C3 H10 109.395 C2 C3 H11 109.395
C3 C2 H8 109.984 C3 C2 H9 109.984
C3 C4 H12 111.648 C3 C4 H13 110.689
C3 C4 H14 110.689 C4 C3 H10 109.567
C4 C3 H11 109.567 O5 C1 H6 111.015
O5 C1 H7 111.015 H6 C1 H7 107.789
H8 C2 H9 106.980 H10 C3 H11 106.489
H12 C4 H13 107.998 H12 C4 H14 107.998
H13 C4 H14 107.671
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability