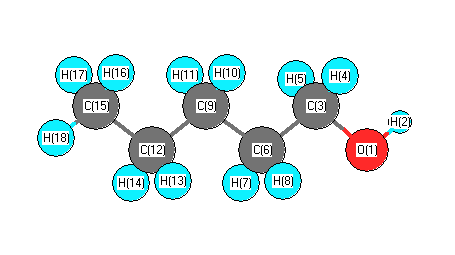
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3890 |
3723 |
30.59 |
|
|
|
| 2 |
A' |
3126 |
2992 |
37.45 |
|
|
|
| 3 |
A' |
3055 |
2924 |
60.01 |
|
|
|
| 4 |
A' |
3049 |
2918 |
34.93 |
|
|
|
| 5 |
A' |
3043 |
2912 |
19.74 |
|
|
|
| 6 |
A' |
3026 |
2896 |
15.46 |
|
|
|
| 7 |
A' |
2990 |
2862 |
53.50 |
|
|
|
| 8 |
A' |
1528 |
1463 |
3.26 |
|
|
|
| 9 |
A' |
1511 |
1446 |
6.93 |
|
|
|
| 10 |
A' |
1501 |
1436 |
0.54 |
|
|
|
| 11 |
A' |
1490 |
1426 |
0.71 |
|
|
|
| 12 |
A' |
1486 |
1422 |
0.05 |
|
|
|
| 13 |
A' |
1458 |
1396 |
5.51 |
|
|
|
| 14 |
A' |
1409 |
1349 |
3.44 |
|
|
|
| 15 |
A' |
1405 |
1344 |
0.35 |
|
|
|
| 16 |
A' |
1355 |
1297 |
9.86 |
|
|
|
| 17 |
A' |
1290 |
1235 |
26.70 |
|
|
|
| 18 |
A' |
1235 |
1182 |
22.11 |
|
|
|
| 19 |
A' |
1134 |
1086 |
6.98 |
|
|
|
| 20 |
A' |
1092 |
1045 |
83.78 |
|
|
|
| 21 |
A' |
1091 |
1045 |
12.87 |
|
|
|
| 22 |
A' |
1056 |
1010 |
1.13 |
|
|
|
| 23 |
A' |
1018 |
974 |
11.95 |
|
|
|
| 24 |
A' |
908 |
869 |
4.01 |
|
|
|
| 25 |
A' |
489 |
468 |
11.45 |
|
|
|
| 26 |
A' |
359 |
344 |
0.06 |
|
|
|
| 27 |
A' |
306 |
293 |
4.11 |
|
|
|
| 28 |
A' |
129 |
124 |
1.30 |
|
|
|
| 29 |
A" |
3118 |
2985 |
66.19 |
|
|
|
| 30 |
A" |
3101 |
2968 |
54.08 |
|
|
|
| 31 |
A" |
3078 |
2946 |
8.08 |
|
|
|
| 32 |
A" |
3052 |
2921 |
2.35 |
|
|
|
| 33 |
A" |
3020 |
2890 |
44.12 |
|
|
|
| 34 |
A" |
1498 |
1434 |
7.45 |
|
|
|
| 35 |
A" |
1331 |
1274 |
0.51 |
|
|
|
| 36 |
A" |
1321 |
1264 |
0.83 |
|
|
|
| 37 |
A" |
1298 |
1242 |
0.01 |
|
|
|
| 38 |
A" |
1238 |
1185 |
0.39 |
|
|
|
| 39 |
A" |
1189 |
1138 |
1.02 |
|
|
|
| 40 |
A" |
987 |
944 |
0.96 |
|
|
|
| 41 |
A" |
857 |
820 |
0.00 |
|
|
|
| 42 |
A" |
769 |
736 |
0.16 |
|
|
|
| 43 |
A" |
734 |
702 |
3.67 |
|
|
|
| 44 |
A" |
271 |
260 |
108.71 |
|
|
|
| 45 |
A" |
245 |
234 |
0.84 |
|
|
|
| 46 |
A" |
149 |
142 |
1.31 |
|
|
|
| 47 |
A" |
100 |
95 |
3.77 |
|
|
|
| 48 |
A" |
74 |
71 |
1.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36428.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34865.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.332 |
|
|
|
| 2 |
H |
0.202 |
|
|
|
| 3 |
C |
-0.028 |
|
|
|
| 4 |
H |
0.070 |
|
|
|
| 5 |
H |
0.070 |
|
|
|
| 6 |
C |
-0.209 |
|
|
|
| 7 |
H |
0.118 |
|
|
|
| 8 |
H |
0.118 |
|
|
|
| 9 |
C |
-0.223 |
|
|
|
| 10 |
H |
0.108 |
|
|
|
| 11 |
H |
0.108 |
|
|
|
| 12 |
C |
-0.204 |
|
|
|
| 13 |
H |
0.112 |
|
|
|
| 14 |
H |
0.112 |
|
|
|
| 15 |
C |
-0.364 |
|
|
|
| 16 |
H |
0.111 |
|
|
|
| 17 |
H |
0.111 |
|
|
|
| 18 |
H |
0.119 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.198 |
0.709 |
0.000 |
1.392 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.021 |
-3.840 |
0.000 |
| y |
-3.840 |
-43.174 |
0.000 |
| z |
0.000 |
0.000 |
-39.505 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.319 |
-3.840 |
0.000 |
| y |
-3.840 |
-5.911 |
0.000 |
| z |
0.000 |
0.000 |
-0.408 |
|
| Polar |
|---|
| 3z2-r2 | -0.815 |
|---|
| x2-y2 | 8.153 |
|---|
| xy | -3.840 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.892 |
-1.254 |
0.000 |
| y |
-1.254 |
11.155 |
0.000 |
| z |
0.000 |
0.000 |
8.432 |
<r2> (average value of r
2) Å
2
| <r2> |
298.061 |
| (<r2>)1/2 |
17.264 |