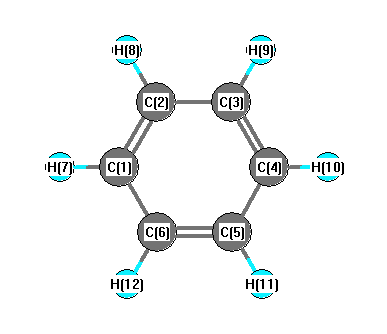
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3256 |
3091 |
0.00 |
392.39 |
0.12 |
0.21 |
| 2 |
A1g |
1024 |
972 |
0.00 |
61.80 |
0.08 |
0.14 |
| 3 |
A2g |
1378 |
1309 |
0.00 |
0.00 |
0.00 |
0.00 |
| 4 |
A2u |
708 |
672 |
108.04 |
0.00 |
0.00 |
0.00 |
| 5 |
B1u |
3201 |
3039 |
0.00 |
0.00 |
0.75 |
0.86 |
| 6 |
B1u |
1024 |
972 |
0.00 |
0.00 |
0.00 |
0.00 |
| 7 |
B2g |
1001 |
951 |
0.00 |
0.00 |
0.75 |
0.86 |
| 8 |
B2g |
721 |
685 |
0.00 |
0.00 |
0.75 |
0.86 |
| 9 |
B2u |
1472 |
1398 |
0.00 |
0.00 |
0.75 |
0.86 |
| 10 |
B2u |
1191 |
1131 |
0.00 |
0.00 |
0.75 |
0.86 |
| 11 |
E1g |
878 |
834 |
0.00 |
0.15 |
0.75 |
0.86 |
| 11 |
E1g |
878 |
834 |
0.00 |
0.15 |
0.75 |
0.86 |
| 12 |
E1u |
3241 |
3077 |
24.09 |
0.00 |
0.00 |
0.00 |
| 12 |
E1u |
3241 |
3077 |
24.09 |
0.00 |
0.00 |
0.00 |
| 13 |
E1u |
1518 |
1442 |
8.15 |
0.00 |
0.00 |
0.00 |
| 13 |
E1u |
1518 |
1442 |
8.15 |
0.00 |
0.00 |
0.00 |
| 14 |
E1u |
1071 |
1017 |
4.65 |
0.00 |
0.00 |
0.00 |
| 14 |
E1u |
1071 |
1017 |
4.65 |
0.00 |
0.00 |
0.00 |
| 15 |
E2g |
3219 |
3056 |
0.00 |
121.62 |
0.75 |
0.86 |
| 15 |
E2g |
3219 |
3056 |
0.00 |
121.62 |
0.75 |
0.86 |
| 16 |
E2g |
1652 |
1568 |
0.00 |
8.18 |
0.75 |
0.86 |
| 16 |
E2g |
1652 |
1568 |
0.00 |
8.18 |
0.75 |
0.86 |
| 17 |
E2g |
1213 |
1151 |
0.00 |
6.16 |
0.75 |
0.86 |
| 17 |
E2g |
1213 |
1151 |
0.00 |
6.16 |
0.75 |
0.86 |
| 18 |
E2g |
611 |
580 |
0.00 |
3.52 |
0.75 |
0.86 |
| 18 |
E2g |
611 |
580 |
0.00 |
3.52 |
0.75 |
0.86 |
| 19 |
E2u |
985 |
935 |
0.00 |
0.00 |
0.00 |
0.00 |
| 19 |
E2u |
985 |
935 |
0.00 |
0.00 |
0.00 |
0.00 |
| 20 |
E2u |
412 |
391 |
0.00 |
0.00 |
0.00 |
0.00 |
| 20 |
E2u |
412 |
391 |
0.00 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 22288.2 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 21160.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.