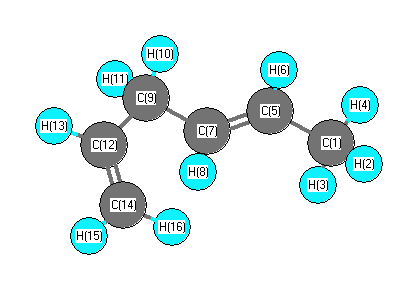
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3248 |
3109 |
17.39 |
|
|
|
| 2 |
A |
3166 |
3030 |
7.44 |
|
|
|
| 3 |
A |
3153 |
3018 |
21.76 |
|
|
|
| 4 |
A |
3149 |
3014 |
51.27 |
|
|
|
| 5 |
A |
3142 |
3007 |
0.87 |
|
|
|
| 6 |
A |
3126 |
2992 |
5.05 |
|
|
|
| 7 |
A |
3095 |
2962 |
17.83 |
|
|
|
| 8 |
A |
3055 |
2924 |
12.94 |
|
|
|
| 9 |
A |
3042 |
2912 |
31.02 |
|
|
|
| 10 |
A |
3020 |
2890 |
25.63 |
|
|
|
| 11 |
A |
1765 |
1690 |
1.35 |
|
|
|
| 12 |
A |
1729 |
1654 |
16.69 |
|
|
|
| 13 |
A |
1491 |
1427 |
11.14 |
|
|
|
| 14 |
A |
1478 |
1415 |
6.05 |
|
|
|
| 15 |
A |
1462 |
1400 |
14.86 |
|
|
|
| 16 |
A |
1448 |
1386 |
1.61 |
|
|
|
| 17 |
A |
1410 |
1350 |
3.17 |
|
|
|
| 18 |
A |
1370 |
1311 |
0.30 |
|
|
|
| 19 |
A |
1334 |
1277 |
1.65 |
|
|
|
| 20 |
A |
1330 |
1273 |
1.56 |
|
|
|
| 21 |
A |
1313 |
1256 |
2.80 |
|
|
|
| 22 |
A |
1223 |
1170 |
1.11 |
|
|
|
| 23 |
A |
1143 |
1094 |
4.58 |
|
|
|
| 24 |
A |
1117 |
1069 |
6.42 |
|
|
|
| 25 |
A |
1069 |
1023 |
0.66 |
|
|
|
| 26 |
A |
1055 |
1010 |
8.47 |
|
|
|
| 27 |
A |
1041 |
996 |
5.72 |
|
|
|
| 28 |
A |
1012 |
968 |
32.14 |
|
|
|
| 29 |
A |
966 |
925 |
29.77 |
|
|
|
| 30 |
A |
952 |
911 |
16.76 |
|
|
|
| 31 |
A |
934 |
894 |
1.91 |
|
|
|
| 32 |
A |
899 |
861 |
3.17 |
|
|
|
| 33 |
A |
789 |
755 |
0.31 |
|
|
|
| 34 |
A |
599 |
573 |
5.18 |
|
|
|
| 35 |
A |
553 |
529 |
7.18 |
|
|
|
| 36 |
A |
467 |
447 |
0.30 |
|
|
|
| 37 |
A |
318 |
304 |
0.32 |
|
|
|
| 38 |
A |
280 |
268 |
1.91 |
|
|
|
| 39 |
A |
217 |
208 |
0.85 |
|
|
|
| 40 |
A |
158 |
151 |
0.69 |
|
|
|
| 41 |
A |
141 |
135 |
0.09 |
|
|
|
| 42 |
A |
79 |
75 |
0.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31167.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29830.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.332 |
|
|
|
| 2 |
H |
0.116 |
|
|
|
| 3 |
H |
0.111 |
|
|
|
| 4 |
H |
0.117 |
|
|
|
| 5 |
C |
-0.122 |
|
|
|
| 6 |
H |
0.124 |
|
|
|
| 7 |
C |
-0.188 |
|
|
|
| 8 |
H |
0.149 |
|
|
|
| 9 |
C |
-0.164 |
|
|
|
| 10 |
H |
0.114 |
|
|
|
| 11 |
H |
0.118 |
|
|
|
| 12 |
C |
-0.108 |
|
|
|
| 13 |
H |
0.136 |
|
|
|
| 14 |
C |
-0.335 |
|
|
|
| 15 |
H |
0.133 |
|
|
|
| 16 |
H |
0.132 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.332 |
-0.313 |
-0.044 |
0.458 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.628 |
1.342 |
-0.385 |
| y |
1.342 |
-37.699 |
0.758 |
| z |
-0.385 |
0.758 |
-39.165 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.804 |
1.342 |
-0.385 |
| y |
1.342 |
0.197 |
0.758 |
| z |
-0.385 |
0.758 |
-2.001 |
|
| Polar |
|---|
| 3z2-r2 | -4.003 |
|---|
| x2-y2 | 1.071 |
|---|
| xy | 1.342 |
|---|
| xz | -0.385 |
|---|
| yz | 0.758 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.310 |
0.221 |
-0.691 |
| y |
0.221 |
10.444 |
0.221 |
| z |
-0.691 |
0.221 |
8.124 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |