Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.662134 |
| Energy at 298.15K | -79.668102 |
| HF Energy | -79.260005 |
| Nuclear repulsion energy | 42.518380 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3099 |
2942 |
0.00 |
274.76 |
0.01 |
0.01 |
| 2 |
A1g |
1441 |
1368 |
0.00 |
0.54 |
0.58 |
0.74 |
| 3 |
A1g |
1038 |
985 |
0.00 |
11.66 |
0.21 |
0.35 |
| 4 |
A1u |
343 |
325 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
3101 |
2944 |
47.22 |
0.00 |
0.01 |
0.01 |
| 6 |
A2u |
1418 |
1346 |
1.74 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3158 |
2998 |
0.00 |
118.20 |
0.75 |
0.86 |
| 7 |
Eg |
3158 |
2998 |
0.00 |
118.20 |
0.75 |
0.86 |
| 8 |
Eg |
1536 |
1458 |
0.00 |
16.30 |
0.75 |
0.86 |
| 8 |
Eg |
1536 |
1458 |
0.00 |
16.30 |
0.75 |
0.86 |
| 9 |
Eg |
1239 |
1176 |
0.00 |
0.25 |
0.75 |
0.86 |
| 9 |
Eg |
1239 |
1176 |
0.00 |
0.25 |
0.75 |
0.86 |
| 10 |
Eu |
3180 |
3019 |
46.39 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3180 |
3019 |
46.39 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1535 |
1457 |
8.85 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1535 |
1457 |
8.85 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
834 |
792 |
3.49 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
834 |
792 |
3.49 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16700.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 15855.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.759 |
| C2 |
0.000 |
0.000 |
-0.759 |
| H3 |
0.000 |
1.012 |
1.151 |
| H4 |
-0.876 |
-0.506 |
1.151 |
| H5 |
0.876 |
-0.506 |
1.151 |
| H6 |
0.000 |
-1.012 |
-1.151 |
| H7 |
-0.876 |
0.506 |
-1.151 |
| H8 |
0.876 |
0.506 |
-1.151 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5178 | 1.0849 | 1.0849 | 1.0849 | 2.1610 | 2.1610 | 2.1610 |
C2 | 1.5178 | | 2.1610 | 2.1610 | 2.1610 | 1.0849 | 1.0849 | 1.0849 | H3 | 1.0849 | 2.1610 | | 1.7524 | 1.7524 | 3.0644 | 2.5139 | 2.5139 | H4 | 1.0849 | 2.1610 | 1.7524 | | 1.7524 | 2.5139 | 2.5139 | 3.0644 | H5 | 1.0849 | 2.1610 | 1.7524 | 1.7524 | | 2.5139 | 3.0644 | 2.5139 | H6 | 2.1610 | 1.0849 | 3.0644 | 2.5139 | 2.5139 | | 1.7524 | 1.7524 | H7 | 2.1610 | 1.0849 | 2.5139 | 2.5139 | 3.0644 | 1.7524 | | 1.7524 | H8 | 2.1610 | 1.0849 | 2.5139 | 3.0644 | 2.5139 | 1.7524 | 1.7524 | |
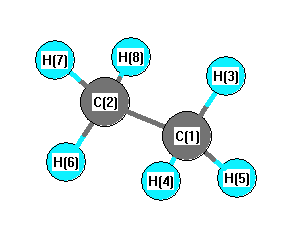 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.166 |
|
C1 |
C2 |
H7 |
111.166 |
| C1 |
C2 |
H8 |
111.166 |
|
C2 |
C1 |
H3 |
111.166 |
| C2 |
C1 |
H4 |
111.166 |
|
C2 |
C1 |
H5 |
111.166 |
| H3 |
C1 |
H4 |
107.724 |
|
H3 |
C1 |
H5 |
107.724 |
| H4 |
C1 |
H5 |
107.724 |
|
H6 |
C2 |
H7 |
107.724 |
| H6 |
C2 |
H8 |
107.724 |
|
H7 |
C2 |
H8 |
107.724 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.657250 |
| Energy at 298.15K | |
| HF Energy | -79.255177 |
| Nuclear repulsion energy | 42.346611 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3113 |
2956 |
0.00 |
|
|
|
| 2 |
A1' |
1451 |
1378 |
0.00 |
|
|
|
| 3 |
A1' |
1034 |
981 |
0.00 |
|
|
|
| 4 |
A1" |
309i |
294i |
0.00 |
|
|
|
| 5 |
A2" |
3107 |
2950 |
48.72 |
|
|
|
| 6 |
A2" |
1415 |
1343 |
1.30 |
|
|
|
| 7 |
E' |
3187 |
3026 |
38.68 |
|
|
|
| 7 |
E' |
3187 |
3026 |
38.68 |
|
|
|
| 8 |
E' |
1542 |
1464 |
10.40 |
|
|
|
| 8 |
E' |
1542 |
1464 |
10.40 |
|
|
|
| 9 |
E' |
905 |
859 |
2.55 |
|
|
|
| 9 |
E' |
905 |
859 |
2.55 |
|
|
|
| 10 |
E" |
3165 |
3005 |
0.00 |
|
|
|
| 10 |
E" |
3165 |
3005 |
0.00 |
|
|
|
| 11 |
E" |
1532 |
1454 |
0.00 |
|
|
|
| 11 |
E" |
1532 |
1454 |
0.00 |
|
|
|
| 12 |
E" |
1183 |
1123 |
0.00 |
|
|
|
| 12 |
E" |
1183 |
1123 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16419.5 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 15588.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.766 |
| C2 |
0.000 |
0.000 |
-0.766 |
| H3 |
0.000 |
1.008 |
1.165 |
| H4 |
-0.873 |
-0.504 |
1.165 |
| H5 |
0.873 |
-0.504 |
1.165 |
| H6 |
0.000 |
1.008 |
-1.165 |
| H7 |
0.873 |
-0.504 |
-1.165 |
| H8 |
-0.873 |
-0.504 |
-1.165 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5322 | 1.0838 | 1.0838 | 1.0838 | 2.1785 | 2.1785 | 2.1785 |
C2 | 1.5322 | | 2.1785 | 2.1785 | 2.1785 | 1.0838 | 1.0838 | 1.0838 | H3 | 1.0838 | 2.1785 | | 1.7451 | 1.7451 | 2.3308 | 2.9117 | 2.9117 | H4 | 1.0838 | 2.1785 | 1.7451 | | 1.7451 | 2.9117 | 2.9117 | 2.3308 | H5 | 1.0838 | 2.1785 | 1.7451 | 1.7451 | | 2.9117 | 2.3308 | 2.9117 | H6 | 2.1785 | 1.0838 | 2.3308 | 2.9117 | 2.9117 | | 1.7451 | 1.7451 | H7 | 2.1785 | 1.0838 | 2.9117 | 2.9117 | 2.3308 | 1.7451 | | 1.7451 | H8 | 2.1785 | 1.0838 | 2.9117 | 2.3308 | 2.9117 | 1.7451 | 1.7451 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.618 |
|
C1 |
C2 |
H7 |
111.618 |
| C1 |
C2 |
H8 |
111.618 |
|
C2 |
C1 |
H3 |
111.618 |
| C2 |
C1 |
H4 |
111.618 |
|
C2 |
C1 |
H5 |
111.618 |
| H3 |
C1 |
H4 |
107.241 |
|
H3 |
C1 |
H5 |
107.241 |
| H4 |
C1 |
H5 |
107.241 |
|
H6 |
C2 |
H7 |
107.241 |
| H6 |
C2 |
H8 |
107.241 |
|
H7 |
C2 |
H8 |
107.241 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
