Jump to
S1C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.707461 |
| Energy at 298.15K | -79.713413 |
| HF Energy | -79.259974 |
| Nuclear repulsion energy | 42.387511 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3062 |
2949 |
|
|
|
|
| 2 |
A1g |
1436 |
1383 |
|
|
|
|
| 3 |
A1g |
1023 |
985 |
|
|
|
|
| 4 |
A1u |
336 |
323 |
|
|
|
|
| 5 |
A2u |
3062 |
2948 |
|
|
|
|
| 6 |
A2u |
1414 |
1361 |
|
|
|
|
| 7 |
Eg |
3108 |
2992 |
|
|
|
|
| 7 |
Eg |
3108 |
2992 |
|
|
|
|
| 8 |
Eg |
1525 |
1469 |
|
|
|
|
| 8 |
Eg |
1524 |
1468 |
|
|
|
|
| 9 |
Eg |
1234 |
1188 |
|
|
|
|
| 9 |
Eg |
1233 |
1188 |
|
|
|
|
| 10 |
Eu |
3131 |
3015 |
|
|
|
|
| 10 |
Eu |
3131 |
3015 |
|
|
|
|
| 11 |
Eu |
1525 |
1469 |
|
|
|
|
| 11 |
Eu |
1525 |
1469 |
|
|
|
|
| 12 |
Eu |
827 |
797 |
|
|
|
|
| 12 |
Eu |
827 |
796 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16515.3 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 15902.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.761 |
| C2 |
0.000 |
0.000 |
-0.761 |
| H3 |
0.000 |
1.015 |
1.154 |
| H4 |
-0.879 |
-0.507 |
1.154 |
| H5 |
0.879 |
-0.507 |
1.154 |
| H6 |
0.000 |
-1.015 |
-1.154 |
| H7 |
-0.879 |
0.507 |
-1.154 |
| H8 |
0.879 |
0.507 |
-1.154 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5230 | 1.0880 | 1.0880 | 1.0880 | 2.1675 | 2.1675 | 2.1675 |
C2 | 1.5230 | | 2.1675 | 2.1675 | 2.1675 | 1.0880 | 1.0880 | 1.0880 | H3 | 1.0880 | 2.1675 | | 1.7577 | 1.7577 | 3.0731 | 2.5208 | 2.5208 | H4 | 1.0880 | 2.1675 | 1.7577 | | 1.7577 | 2.5208 | 2.5208 | 3.0731 | H5 | 1.0880 | 2.1675 | 1.7577 | 1.7577 | | 2.5208 | 3.0731 | 2.5208 | H6 | 2.1675 | 1.0880 | 3.0731 | 2.5208 | 2.5208 | | 1.7577 | 1.7577 | H7 | 2.1675 | 1.0880 | 2.5208 | 2.5208 | 3.0731 | 1.7577 | | 1.7577 | H8 | 2.1675 | 1.0880 | 2.5208 | 3.0731 | 2.5208 | 1.7577 | 1.7577 | |
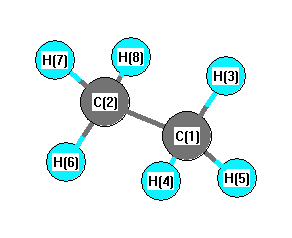 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.132 |
|
C1 |
C2 |
H7 |
111.132 |
| C1 |
C2 |
H8 |
111.132 |
|
C2 |
C1 |
H3 |
111.132 |
| C2 |
C1 |
H4 |
111.132 |
|
C2 |
C1 |
H5 |
111.132 |
| H3 |
C1 |
H4 |
107.761 |
|
H3 |
C1 |
H5 |
107.761 |
| H4 |
C1 |
H5 |
107.761 |
|
H6 |
C2 |
H7 |
107.761 |
| H6 |
C2 |
H8 |
107.761 |
|
H7 |
C2 |
H8 |
107.761 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.702708 |
| Energy at 298.15K | |
| HF Energy | -79.255141 |
| Nuclear repulsion energy | 42.217013 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3076 |
2961 |
|
|
|
|
| 2 |
A1' |
1446 |
1392 |
|
|
|
|
| 3 |
A1' |
1019 |
981 |
|
|
|
|
| 4 |
A1" |
306i |
295i |
|
|
|
|
| 5 |
A2" |
3067 |
2954 |
|
|
|
|
| 6 |
A2" |
1410 |
1358 |
|
|
|
|
| 7 |
E' |
3137 |
3021 |
|
|
|
|
| 7 |
E' |
3137 |
3020 |
|
|
|
|
| 8 |
E' |
1533 |
1476 |
|
|
|
|
| 8 |
E' |
1533 |
1476 |
|
|
|
|
| 9 |
E' |
897 |
864 |
|
|
|
|
| 9 |
E' |
897 |
863 |
|
|
|
|
| 10 |
E" |
3114 |
2999 |
|
|
|
|
| 10 |
E" |
3114 |
2999 |
|
|
|
|
| 11 |
E" |
1521 |
1465 |
|
|
|
|
| 11 |
E" |
1521 |
1464 |
|
|
|
|
| 12 |
E" |
1179 |
1135 |
|
|
|
|
| 12 |
E" |
1179 |
1135 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16236.7 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 15634.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.769 |
| C2 |
0.000 |
0.000 |
-0.769 |
| H3 |
0.000 |
1.011 |
1.168 |
| H4 |
-0.875 |
-0.505 |
1.168 |
| H5 |
0.875 |
-0.505 |
1.168 |
| H6 |
0.000 |
1.011 |
-1.168 |
| H7 |
0.875 |
-0.505 |
-1.168 |
| H8 |
-0.875 |
-0.505 |
-1.168 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5372 | 1.0869 | 1.0869 | 1.0869 | 2.1849 | 2.1849 | 2.1849 |
C2 | 1.5372 | | 2.1849 | 2.1849 | 2.1849 | 1.0869 | 1.0869 | 1.0869 | H3 | 1.0869 | 2.1849 | | 1.7506 | 1.7506 | 2.3369 | 2.9199 | 2.9199 | H4 | 1.0869 | 2.1849 | 1.7506 | | 1.7506 | 2.9199 | 2.9199 | 2.3369 | H5 | 1.0869 | 2.1849 | 1.7506 | 1.7506 | | 2.9199 | 2.3369 | 2.9199 | H6 | 2.1849 | 1.0869 | 2.3369 | 2.9199 | 2.9199 | | 1.7506 | 1.7506 | H7 | 2.1849 | 1.0869 | 2.9199 | 2.9199 | 2.3369 | 1.7506 | | 1.7506 | H8 | 2.1849 | 1.0869 | 2.9199 | 2.3369 | 2.9199 | 1.7506 | 1.7506 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.584 |
|
C1 |
C2 |
H7 |
111.584 |
| C1 |
C2 |
H8 |
111.584 |
|
C2 |
C1 |
H3 |
111.584 |
| C2 |
C1 |
H4 |
111.584 |
|
C2 |
C1 |
H5 |
111.584 |
| H3 |
C1 |
H4 |
107.278 |
|
H3 |
C1 |
H5 |
107.278 |
| H4 |
C1 |
H5 |
107.278 |
|
H6 |
C2 |
H7 |
107.278 |
| H6 |
C2 |
H8 |
107.278 |
|
H7 |
C2 |
H8 |
107.278 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
