Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.797440 |
| Energy at 298.15K | -79.803349 |
| HF Energy | -79.797440 |
| Nuclear repulsion energy | 42.432379 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3056 |
2925 |
0.00 |
293.49 |
0.01 |
0.01 |
| 2 |
A1g |
1422 |
1361 |
0.00 |
1.08 |
0.75 |
0.86 |
| 3 |
A1g |
1022 |
978 |
0.00 |
10.90 |
0.22 |
0.36 |
| 4 |
A1u |
308 |
295 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
3057 |
2926 |
58.47 |
0.00 |
0.00 |
0.00 |
| 6 |
A2u |
1405 |
1344 |
1.43 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3112 |
2978 |
0.00 |
135.48 |
0.75 |
0.86 |
| 7 |
Eg |
3112 |
2978 |
0.00 |
135.48 |
0.75 |
0.86 |
| 8 |
Eg |
1503 |
1439 |
0.00 |
17.73 |
0.75 |
0.86 |
| 8 |
Eg |
1503 |
1439 |
0.00 |
17.73 |
0.75 |
0.86 |
| 9 |
Eg |
1219 |
1166 |
0.00 |
0.26 |
0.75 |
0.86 |
| 9 |
Eg |
1219 |
1166 |
0.00 |
0.26 |
0.75 |
0.86 |
| 10 |
Eu |
3137 |
3002 |
61.24 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3137 |
3002 |
61.24 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1505 |
1440 |
9.18 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1505 |
1440 |
9.18 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
821 |
786 |
4.39 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
821 |
786 |
4.39 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 16431.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 15726.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
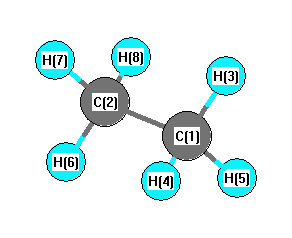
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.759 |
| C2 |
0.000 |
0.000 |
-0.759 |
| H3 |
0.000 |
1.014 |
1.155 |
| H4 |
-0.878 |
-0.507 |
1.155 |
| H5 |
0.878 |
-0.507 |
1.155 |
| H6 |
0.000 |
-1.014 |
-1.155 |
| H7 |
-0.878 |
0.507 |
-1.155 |
| H8 |
0.878 |
0.507 |
-1.155 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5177 | 1.0884 | 1.0884 | 1.0884 | 2.1661 | 2.1661 | 2.1661 |
C2 | 1.5177 | | 2.1661 | 2.1661 | 2.1661 | 1.0884 | 1.0884 | 1.0884 | H3 | 1.0884 | 2.1661 | | 1.7556 | 1.7556 | 3.0741 | 2.5234 | 2.5234 | H4 | 1.0884 | 2.1661 | 1.7556 | | 1.7556 | 2.5234 | 2.5234 | 3.0741 | H5 | 1.0884 | 2.1661 | 1.7556 | 1.7556 | | 2.5234 | 3.0741 | 2.5234 | H6 | 2.1661 | 1.0884 | 3.0741 | 2.5234 | 2.5234 | | 1.7556 | 1.7556 | H7 | 2.1661 | 1.0884 | 2.5234 | 2.5234 | 3.0741 | 1.7556 | | 1.7556 | H8 | 2.1661 | 1.0884 | 2.5234 | 3.0741 | 2.5234 | 1.7556 | 1.7556 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.370 |
|
C1 |
C2 |
H7 |
111.370 |
| C1 |
C2 |
H8 |
111.370 |
|
C2 |
C1 |
H3 |
111.370 |
| C2 |
C1 |
H4 |
111.370 |
|
C2 |
C1 |
H5 |
111.370 |
| H3 |
C1 |
H4 |
107.508 |
|
H3 |
C1 |
H5 |
107.508 |
| H4 |
C1 |
H5 |
107.508 |
|
H6 |
C2 |
H7 |
107.508 |
| H6 |
C2 |
H8 |
107.508 |
|
H7 |
C2 |
H8 |
107.508 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.338 |
|
|
|
| 2 |
C |
-0.338 |
|
|
|
| 3 |
H |
0.113 |
|
|
|
| 4 |
H |
0.113 |
|
|
|
| 5 |
H |
0.113 |
|
|
|
| 6 |
H |
0.113 |
|
|
|
| 7 |
H |
0.113 |
|
|
|
| 8 |
H |
0.113 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.824 |
0.000 |
0.000 |
| y |
0.000 |
-14.824 |
0.000 |
| z |
0.000 |
0.000 |
-15.489 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.333 |
0.000 |
0.000 |
| y |
0.000 |
0.333 |
0.000 |
| z |
0.000 |
0.000 |
-0.666 |
|
| Polar |
|---|
| 3z2-r2 | -1.331 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.830 |
0.000 |
0.000 |
| y |
0.000 |
3.830 |
0.000 |
| z |
0.000 |
0.000 |
4.314 |
<r2> (average value of r
2) Å
2
| <r2> |
30.483 |
| (<r2>)1/2 |
5.521 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.793150 |
| Energy at 298.15K | |
| HF Energy | -79.793150 |
| Nuclear repulsion energy | 42.275121 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3072 |
2941 |
0.00 |
|
|
|
| 2 |
A1' |
1438 |
1376 |
0.00 |
|
|
|
| 3 |
A1' |
1021 |
977 |
0.00 |
|
|
|
| 4 |
A1" |
300i |
287i |
0.00 |
|
|
|
| 5 |
A2" |
3065 |
2934 |
59.81 |
|
|
|
| 6 |
A2" |
1402 |
1342 |
1.03 |
|
|
|
| 7 |
E' |
3144 |
3009 |
51.29 |
|
|
|
| 7 |
E' |
3144 |
3009 |
51.29 |
|
|
|
| 8 |
E' |
1510 |
1445 |
10.54 |
|
|
|
| 8 |
E' |
1510 |
1445 |
10.54 |
|
|
|
| 9 |
E' |
897 |
858 |
3.25 |
|
|
|
| 9 |
E' |
897 |
858 |
3.25 |
|
|
|
| 10 |
E" |
3122 |
2988 |
0.00 |
|
|
|
| 10 |
E" |
3122 |
2988 |
0.00 |
|
|
|
| 11 |
E" |
1502 |
1438 |
0.00 |
|
|
|
| 11 |
E" |
1502 |
1438 |
0.00 |
|
|
|
| 12 |
E" |
1169 |
1119 |
0.00 |
|
|
|
| 12 |
E" |
1169 |
1119 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16193.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 15498.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.765 |
| C2 |
0.000 |
0.000 |
-0.765 |
| H3 |
0.000 |
1.009 |
1.170 |
| H4 |
-0.874 |
-0.505 |
1.170 |
| H5 |
0.874 |
-0.505 |
1.170 |
| H6 |
0.000 |
1.009 |
-1.170 |
| H7 |
0.874 |
-0.505 |
-1.170 |
| H8 |
-0.874 |
-0.505 |
-1.170 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5307 | 1.0874 | 1.0874 | 1.0874 | 2.1825 | 2.1825 | 2.1825 |
C2 | 1.5307 | | 2.1825 | 2.1825 | 2.1825 | 1.0874 | 1.0874 | 1.0874 | H3 | 1.0874 | 2.1825 | | 1.7483 | 1.7483 | 2.3394 | 2.9206 | 2.9206 | H4 | 1.0874 | 2.1825 | 1.7483 | | 1.7483 | 2.9206 | 2.9206 | 2.3394 | H5 | 1.0874 | 2.1825 | 1.7483 | 1.7483 | | 2.9206 | 2.3394 | 2.9206 | H6 | 2.1825 | 1.0874 | 2.3394 | 2.9206 | 2.9206 | | 1.7483 | 1.7483 | H7 | 2.1825 | 1.0874 | 2.9206 | 2.9206 | 2.3394 | 1.7483 | | 1.7483 | H8 | 2.1825 | 1.0874 | 2.9206 | 2.3394 | 2.9206 | 1.7483 | 1.7483 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.830 |
|
C1 |
C2 |
H7 |
111.830 |
| C1 |
C2 |
H8 |
111.830 |
|
C2 |
C1 |
H3 |
111.830 |
| C2 |
C1 |
H4 |
111.830 |
|
C2 |
C1 |
H5 |
111.830 |
| H3 |
C1 |
H4 |
107.013 |
|
H3 |
C1 |
H5 |
107.013 |
| H4 |
C1 |
H5 |
107.013 |
|
H6 |
C2 |
H7 |
107.013 |
| H6 |
C2 |
H8 |
107.013 |
|
H7 |
C2 |
H8 |
107.013 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.343 |
|
|
|
| 2 |
C |
-0.343 |
|
|
|
| 3 |
H |
0.114 |
|
|
|
| 4 |
H |
0.114 |
|
|
|
| 5 |
H |
0.114 |
|
|
|
| 6 |
H |
0.114 |
|
|
|
| 7 |
H |
0.114 |
|
|
|
| 8 |
H |
0.114 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.813 |
0.000 |
0.000 |
| y |
0.000 |
-14.813 |
0.000 |
| z |
0.000 |
0.000 |
-15.464 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.325 |
0.000 |
0.000 |
| y |
0.000 |
0.325 |
0.000 |
| z |
0.000 |
0.000 |
-0.651 |
|
| Polar |
|---|
| 3z2-r2 | -1.301 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.790 |
0.000 |
0.000 |
| y |
0.000 |
3.790 |
0.000 |
| z |
0.000 |
0.000 |
4.275 |
<r2> (average value of r
2) Å
2
| <r2> |
30.740 |
| (<r2>)1/2 |
5.544 |
