return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8 (Propane)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-118.900187
Energy at 298.15K-118.908638
HF Energy-118.307882
Nuclear repulsion energy83.182302
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3166 3006 35.68 83.02 0.55 0.71
2 A1 3096 2940 32.33 57.45 0.26 0.42
3 A1 3087 2931 8.03 335.32 0.00 0.01
4 A1 1540 1462 5.02 2.57 0.75 0.86
5 A1 1518 1441 0.02 18.38 0.75 0.86
6 A1 1431 1359 3.87 0.35 0.11 0.20
7 A1 1196 1136 0.81 2.07 0.14 0.25
8 A1 906 860 0.90 10.57 0.16 0.28
9 A1 370 351 0.05 0.32 0.25 0.40
10 A2 3153 2994 0.00 7.00 0.75 0.86
11 A2 1518 1442 0.00 15.21 0.75 0.86
12 A2 1338 1270 0.00 3.89 0.75 0.86
13 A2 921 875 0.00 0.10 0.75 0.86
14 A2 236 224 0.00 0.02 0.75 0.86
15 B1 3162 3002 70.09 25.92 0.75 0.86
16 B1 3119 2961 2.47 138.02 0.75 0.86
17 B1 1536 1459 13.88 0.00 0.75 0.86
18 B1 1229 1167 0.10 0.02 0.75 0.86
19 B1 763 724 2.79 0.11 0.75 0.86
20 B1 293 278 0.00 0.01 0.75 0.86
21 B2 3164 3004 23.20 48.61 0.75 0.86
22 B2 3087 2931 27.54 1.19 0.75 0.86
23 B2 1524 1447 2.12 0.20 0.75 0.86
24 B2 1419 1347 5.41 0.10 0.75 0.86
25 B2 1380 1311 0.97 0.24 0.75 0.86
26 B2 1099 1043 0.52 5.09 0.75 0.86
27 B2 943 895 1.82 0.02 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 23096.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 21927.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.99103 0.28744 0.25388

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.589
C2 0.000 1.258 -0.259
C3 0.000 -1.258 -0.259
H4 0.871 0.000 1.239
H5 -0.871 0.000 1.239
H6 0.000 2.155 0.351
H7 0.000 -2.155 0.351
H8 0.877 1.288 -0.900
H9 -0.877 1.288 -0.900
H10 -0.877 -1.288 -0.900
H11 0.877 -1.288 -0.900

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10 H11
C11.51711.51711.08691.08692.16822.16822.15532.15532.15532.1553
C21.51712.51562.14152.14151.08523.46711.08651.08652.76782.7678
C31.51712.51562.14152.14153.46711.08522.76782.76781.08651.0865
H41.08692.14152.14151.74172.48842.48842.49723.04813.04812.4972
H51.08692.14152.14151.74172.48842.48843.04812.49722.49723.0481
H62.16821.08523.46712.48842.48844.31031.75671.75673.76693.7669
H72.16823.46711.08522.48842.48844.31033.76693.76691.75671.7567
H82.15531.08652.76782.49723.04811.75673.76691.75383.11642.5761
H92.15531.08652.76783.04812.49721.75673.76691.75382.57613.1164
H102.15532.76781.08653.04812.49723.76691.75673.11642.57611.7538
H112.15532.76781.08652.49723.04813.76691.75672.57613.11641.7538

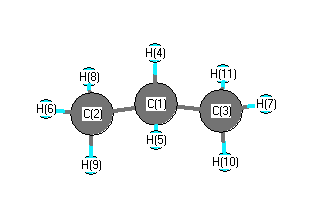
picture of Propane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H6 111.785 C1 C2 H8 110.661
C1 C2 H9 110.661 C1 C3 H7 111.785
C1 C3 H10 110.661 C1 C3 H11 110.661
C2 C1 C3 112.012 C2 C1 H4 109.545
C2 C1 H5 109.545 C3 C1 H4 109.545
C3 C1 H5 109.545 H4 C1 H5 106.493
H6 C2 H8 107.980 H6 C2 H9 107.980
H7 C3 H10 107.980 H7 C3 H11 107.980
H8 C2 H9 107.621 H10 C3 H11 107.621
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability