Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -134.929315 |
| Energy at 298.15K | -134.937531 |
| HF Energy | -134.304248 |
| Nuclear repulsion energy | 83.344941 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3539 |
3360 |
0.81 |
|
|
|
| 2 |
A' |
3158 |
2998 |
33.37 |
|
|
|
| 3 |
A' |
3103 |
2946 |
30.54 |
|
|
|
| 4 |
A' |
3082 |
2926 |
16.63 |
|
|
|
| 5 |
A' |
1665 |
1581 |
20.99 |
|
|
|
| 6 |
A' |
1531 |
1454 |
2.33 |
|
|
|
| 7 |
A' |
1513 |
1437 |
0.16 |
|
|
|
| 8 |
A' |
1418 |
1346 |
11.67 |
|
|
|
| 9 |
A' |
1391 |
1321 |
8.87 |
|
|
|
| 10 |
A' |
1173 |
1113 |
8.33 |
|
|
|
| 11 |
A' |
1105 |
1049 |
25.48 |
|
|
|
| 12 |
A' |
914 |
868 |
26.71 |
|
|
|
| 13 |
A' |
876 |
832 |
143.41 |
|
|
|
| 14 |
A' |
403 |
382 |
7.01 |
|
|
|
| 15 |
A" |
3630 |
3447 |
1.32 |
|
|
|
| 16 |
A" |
3161 |
3001 |
42.50 |
|
|
|
| 17 |
A" |
3129 |
2970 |
5.00 |
|
|
|
| 18 |
A" |
1521 |
1444 |
8.05 |
|
|
|
| 19 |
A" |
1407 |
1336 |
0.00 |
|
|
|
| 20 |
A" |
1287 |
1222 |
0.14 |
|
|
|
| 21 |
A" |
1017 |
965 |
1.10 |
|
|
|
| 22 |
A" |
788 |
748 |
0.78 |
|
|
|
| 23 |
A" |
303 |
287 |
21.19 |
|
|
|
| 24 |
A" |
265 |
251 |
24.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20688.3 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19641.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.297 |
-0.094 |
0.000 |
| C2 |
0.000 |
0.571 |
0.000 |
| C3 |
1.203 |
-0.346 |
0.000 |
| H4 |
2.135 |
0.214 |
0.000 |
| H5 |
1.199 |
-0.986 |
0.878 |
| H6 |
1.199 |
-0.986 |
-0.878 |
| H7 |
0.033 |
1.222 |
-0.868 |
| H8 |
0.033 |
1.222 |
0.868 |
| H9 |
-1.367 |
-0.690 |
0.810 |
| H10 |
-1.367 |
-0.690 |
-0.810 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4579 | 2.5126 | 3.4464 | 2.7925 | 2.7925 | 2.0629 | 2.0629 | 1.0084 | 1.0084 |
C2 | 1.4579 | | 1.5125 | 2.1650 | 2.1527 | 2.1527 | 1.0856 | 1.0856 | 2.0290 | 2.0290 | C3 | 2.5126 | 1.5125 | | 1.0879 | 1.0865 | 1.0865 | 2.1401 | 2.1401 | 2.7164 | 2.7164 | H4 | 3.4464 | 2.1650 | 1.0879 | | 1.7574 | 1.7574 | 2.4876 | 2.4876 | 3.7069 | 3.7069 | H5 | 2.7925 | 2.1527 | 1.0865 | 1.7574 | | 1.7552 | 3.0470 | 2.4972 | 2.5840 | 3.0857 | H6 | 2.7925 | 2.1527 | 1.0865 | 1.7574 | 1.7552 | | 2.4972 | 3.0470 | 3.0857 | 2.5840 | H7 | 2.0629 | 1.0856 | 2.1401 | 2.4876 | 3.0470 | 2.4972 | | 1.7368 | 2.9042 | 2.3707 | H8 | 2.0629 | 1.0856 | 2.1401 | 2.4876 | 2.4972 | 3.0470 | 1.7368 | | 2.3707 | 2.9042 | H9 | 1.0084 | 2.0290 | 2.7164 | 3.7069 | 2.5840 | 3.0857 | 2.9042 | 2.3707 | | 1.6205 | H10 | 1.0084 | 2.0290 | 2.7164 | 3.7069 | 3.0857 | 2.5840 | 2.3707 | 2.9042 | 1.6205 | |
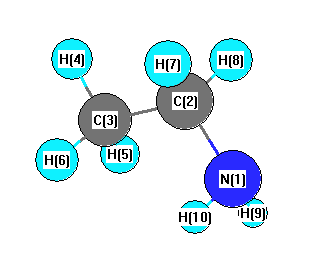 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.521 |
|
N1 |
C2 |
H7 |
107.499 |
| N1 |
C2 |
H8 |
107.499 |
|
C2 |
N1 |
H9 |
109.354 |
| C2 |
N1 |
H10 |
109.354 |
|
C2 |
C3 |
H4 |
111.678 |
| C2 |
C3 |
H5 |
110.782 |
|
C2 |
C3 |
H6 |
110.782 |
| C3 |
C2 |
H7 |
109.823 |
|
C3 |
C2 |
H8 |
109.823 |
| H4 |
C3 |
H5 |
107.844 |
|
H4 |
C3 |
H6 |
107.844 |
| H5 |
C3 |
H6 |
107.753 |
|
H7 |
C2 |
H8 |
106.244 |
| H9 |
N1 |
H10 |
106.927 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -134.929420 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
