Jump to
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -135.191007 |
| Energy at 298.15K | -135.199156 |
| Nuclear repulsion energy | 83.075890 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3522 |
3379 |
0.96 |
|
|
|
| 2 |
A' |
3112 |
2985 |
43.03 |
|
|
|
| 3 |
A' |
3050 |
2926 |
35.57 |
|
|
|
| 4 |
A' |
3037 |
2913 |
21.70 |
|
|
|
| 5 |
A' |
1666 |
1598 |
21.60 |
|
|
|
| 6 |
A' |
1506 |
1444 |
2.85 |
|
|
|
| 7 |
A' |
1487 |
1426 |
0.15 |
|
|
|
| 8 |
A' |
1408 |
1350 |
10.81 |
|
|
|
| 9 |
A' |
1383 |
1326 |
10.09 |
|
|
|
| 10 |
A' |
1160 |
1113 |
9.43 |
|
|
|
| 11 |
A' |
1083 |
1039 |
27.99 |
|
|
|
| 12 |
A' |
901 |
864 |
21.02 |
|
|
|
| 13 |
A' |
861 |
826 |
154.87 |
|
|
|
| 14 |
A' |
400 |
384 |
6.36 |
|
|
|
| 15 |
A" |
3602 |
3455 |
0.18 |
|
|
|
| 16 |
A" |
3115 |
2988 |
54.19 |
|
|
|
| 17 |
A" |
3079 |
2953 |
6.24 |
|
|
|
| 18 |
A" |
1493 |
1432 |
8.00 |
|
|
|
| 19 |
A" |
1395 |
1338 |
0.05 |
|
|
|
| 20 |
A" |
1274 |
1222 |
0.04 |
|
|
|
| 21 |
A" |
1009 |
967 |
0.92 |
|
|
|
| 22 |
A" |
779 |
747 |
1.48 |
|
|
|
| 23 |
A" |
284 |
273 |
26.85 |
|
|
|
| 24 |
A" |
248 |
238 |
17.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20426.6 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 19593.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.301 |
-0.083 |
0.000 |
| C2 |
0.000 |
0.570 |
0.000 |
| C3 |
1.208 |
-0.356 |
0.000 |
| H4 |
2.144 |
0.206 |
0.000 |
| H5 |
1.205 |
-1.000 |
0.882 |
| H6 |
1.205 |
-1.000 |
-0.882 |
| H7 |
0.041 |
1.228 |
-0.871 |
| H8 |
0.041 |
1.228 |
0.871 |
| H9 |
-1.388 |
-0.684 |
0.809 |
| H10 |
-1.388 |
-0.684 |
-0.809 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4556 | 2.5232 | 3.4568 | 2.8102 | 2.8102 | 2.0685 | 2.0685 | 1.0117 | 1.0117 |
C2 | 1.4556 | | 1.5219 | 2.1746 | 2.1669 | 2.1669 | 1.0923 | 1.0923 | 2.0384 | 2.0384 | C3 | 2.5232 | 1.5219 | | 1.0919 | 1.0919 | 1.0919 | 2.1511 | 2.1511 | 2.7388 | 2.7388 | H4 | 3.4568 | 2.1746 | 1.0919 | | 1.7645 | 1.7645 | 2.4947 | 2.4947 | 3.7313 | 3.7313 | H5 | 2.8102 | 2.1669 | 1.0919 | 1.7645 | | 1.7634 | 3.0643 | 2.5133 | 2.6134 | 3.1119 | H6 | 2.8102 | 2.1669 | 1.0919 | 1.7645 | 1.7634 | | 2.5133 | 3.0643 | 3.1119 | 2.6134 | H7 | 2.0685 | 1.0923 | 2.1511 | 2.4947 | 3.0643 | 2.5133 | | 1.7429 | 2.9193 | 2.3878 | H8 | 2.0685 | 1.0923 | 2.1511 | 2.4947 | 2.5133 | 3.0643 | 1.7429 | | 2.3878 | 2.9193 | H9 | 1.0117 | 2.0384 | 2.7388 | 3.7313 | 2.6134 | 3.1119 | 2.9193 | 2.3878 | | 1.6184 | H10 | 1.0117 | 2.0384 | 2.7388 | 3.7313 | 3.1119 | 2.6134 | 2.3878 | 2.9193 | 1.6184 | |
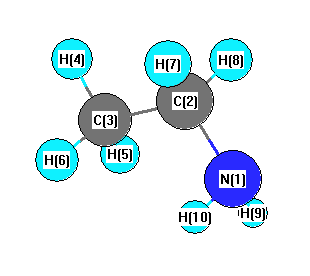 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.846 |
|
N1 |
C2 |
H7 |
107.697 |
| N1 |
C2 |
H8 |
107.697 |
|
C2 |
N1 |
H9 |
110.110 |
| C2 |
N1 |
H10 |
110.110 |
|
C2 |
C3 |
H4 |
111.543 |
| C2 |
C3 |
H5 |
110.923 |
|
C2 |
C3 |
H6 |
110.923 |
| C3 |
C2 |
H7 |
109.641 |
|
C3 |
C2 |
H8 |
109.641 |
| H4 |
C3 |
H5 |
107.794 |
|
H4 |
C3 |
H6 |
107.794 |
| H5 |
C3 |
H6 |
107.700 |
|
H7 |
C2 |
H8 |
105.834 |
| H9 |
N1 |
H10 |
106.228 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.299 |
|
|
|
| 2 |
C |
-0.066 |
|
|
|
| 3 |
C |
-0.257 |
|
|
|
| 4 |
H |
0.076 |
|
|
|
| 5 |
H |
0.077 |
|
|
|
| 6 |
H |
0.077 |
|
|
|
| 7 |
H |
0.081 |
|
|
|
| 8 |
H |
0.081 |
|
|
|
| 9 |
H |
0.115 |
|
|
|
| 10 |
H |
0.115 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.848 |
-0.899 |
0.000 |
1.236 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.729 |
2.511 |
0.000 |
| y |
2.511 |
-20.320 |
0.000 |
| z |
0.000 |
0.000 |
-19.053 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.043 |
2.511 |
0.000 |
| y |
2.511 |
1.071 |
0.000 |
| z |
0.000 |
0.000 |
2.972 |
|
| Polar |
|---|
| 3z2-r2 | 5.943 |
|---|
| x2-y2 | -3.409 |
|---|
| xy | 2.511 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.615 |
0.084 |
0.000 |
| y |
0.084 |
5.036 |
0.000 |
| z |
0.000 |
0.000 |
4.838 |
<r2> (average value of r
2) Å
2
| <r2> |
58.228 |
| (<r2>)1/2 |
7.631 |
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -135.191968 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
