Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -135.141614 |
| Energy at 298.15K | -135.149747 |
| Nuclear repulsion energy | 83.212867 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3521 |
3370 |
0.97 |
|
|
|
| 2 |
A' |
3117 |
2984 |
44.42 |
|
|
|
| 3 |
A' |
3056 |
2925 |
35.93 |
|
|
|
| 4 |
A' |
3042 |
2912 |
22.71 |
|
|
|
| 5 |
A' |
1663 |
1592 |
21.31 |
|
|
|
| 6 |
A' |
1505 |
1440 |
2.51 |
|
|
|
| 7 |
A' |
1486 |
1422 |
0.13 |
|
|
|
| 8 |
A' |
1404 |
1344 |
10.52 |
|
|
|
| 9 |
A' |
1379 |
1319 |
10.90 |
|
|
|
| 10 |
A' |
1158 |
1109 |
9.06 |
|
|
|
| 11 |
A' |
1084 |
1037 |
28.23 |
|
|
|
| 12 |
A' |
899 |
860 |
16.20 |
|
|
|
| 13 |
A' |
860 |
823 |
158.42 |
|
|
|
| 14 |
A' |
393 |
376 |
6.66 |
|
|
|
| 15 |
A" |
3602 |
3448 |
0.15 |
|
|
|
| 16 |
A" |
3121 |
2987 |
56.46 |
|
|
|
| 17 |
A" |
3085 |
2953 |
5.91 |
|
|
|
| 18 |
A" |
1492 |
1428 |
7.80 |
|
|
|
| 19 |
A" |
1391 |
1332 |
0.03 |
|
|
|
| 20 |
A" |
1270 |
1216 |
0.06 |
|
|
|
| 21 |
A" |
1005 |
962 |
0.91 |
|
|
|
| 22 |
A" |
775 |
741 |
1.28 |
|
|
|
| 23 |
A" |
282 |
270 |
27.98 |
|
|
|
| 24 |
A" |
247 |
236 |
15.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20419.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 19543.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.298 |
-0.084 |
0.000 |
| C2 |
0.000 |
0.571 |
0.000 |
| C3 |
1.205 |
-0.355 |
0.000 |
| H4 |
2.140 |
0.205 |
0.000 |
| H5 |
1.200 |
-0.997 |
0.880 |
| H6 |
1.200 |
-0.997 |
-0.880 |
| H7 |
0.040 |
1.227 |
-0.870 |
| H8 |
0.040 |
1.227 |
0.870 |
| H9 |
-1.382 |
-0.685 |
0.808 |
| H10 |
-1.382 |
-0.685 |
-0.808 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4543 | 2.5176 | 3.4506 | 2.8019 | 2.8019 | 2.0657 | 2.0657 | 1.0106 | 1.0106 |
C2 | 1.4543 | | 1.5196 | 2.1715 | 2.1624 | 2.1624 | 1.0902 | 1.0902 | 2.0352 | 2.0352 | C3 | 2.5176 | 1.5196 | | 1.0899 | 1.0899 | 1.0899 | 2.1482 | 2.1482 | 2.7303 | 2.7303 | H4 | 3.4506 | 2.1715 | 1.0899 | | 1.7616 | 1.7616 | 2.4925 | 2.4925 | 3.7218 | 3.7218 | H5 | 2.8019 | 2.1624 | 1.0899 | 1.7616 | | 1.7604 | 3.0588 | 2.5085 | 2.6021 | 3.1009 | H6 | 2.8019 | 2.1624 | 1.0899 | 1.7616 | 1.7604 | | 2.5085 | 3.0588 | 3.1009 | 2.6021 | H7 | 2.0657 | 1.0902 | 2.1482 | 2.4925 | 3.0588 | 2.5085 | | 1.7403 | 2.9146 | 2.3838 | H8 | 2.0657 | 1.0902 | 2.1482 | 2.4925 | 2.5085 | 3.0588 | 1.7403 | | 2.3838 | 2.9146 | H9 | 1.0106 | 2.0352 | 2.7303 | 3.7218 | 2.6021 | 3.1009 | 2.9146 | 2.3838 | | 1.6161 | H10 | 1.0106 | 2.0352 | 2.7303 | 3.7218 | 3.1009 | 2.6021 | 2.3838 | 2.9146 | 1.6161 | |
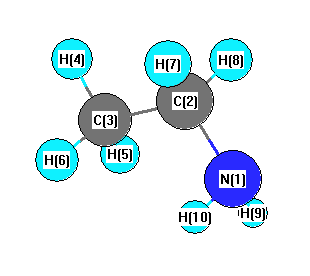 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.668 |
|
N1 |
C2 |
H7 |
107.694 |
| N1 |
C2 |
H8 |
107.694 |
|
C2 |
N1 |
H9 |
110.009 |
| C2 |
N1 |
H10 |
110.009 |
|
C2 |
C3 |
H4 |
111.586 |
| C2 |
C3 |
H5 |
110.852 |
|
C2 |
C3 |
H6 |
110.852 |
| C3 |
C2 |
H7 |
109.705 |
|
C3 |
C2 |
H8 |
109.705 |
| H4 |
C3 |
H5 |
107.835 |
|
H4 |
C3 |
H6 |
107.835 |
| H5 |
C3 |
H6 |
107.723 |
|
H7 |
C2 |
H8 |
105.910 |
| H9 |
N1 |
H10 |
106.185 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.344 |
|
|
|
| 2 |
C |
-0.131 |
|
|
|
| 3 |
C |
-0.346 |
|
|
|
| 4 |
H |
0.105 |
|
|
|
| 5 |
H |
0.105 |
|
|
|
| 6 |
H |
0.105 |
|
|
|
| 7 |
H |
0.112 |
|
|
|
| 8 |
H |
0.112 |
|
|
|
| 9 |
H |
0.141 |
|
|
|
| 10 |
H |
0.141 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.855 |
-0.887 |
0.000 |
1.232 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.758 |
2.481 |
0.000 |
| y |
2.481 |
-20.341 |
0.000 |
| z |
0.000 |
0.000 |
-19.094 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.040 |
2.481 |
0.000 |
| y |
2.481 |
1.085 |
0.000 |
| z |
0.000 |
0.000 |
2.955 |
|
| Polar |
|---|
| 3z2-r2 | 5.911 |
|---|
| x2-y2 | -3.417 |
|---|
| xy | 2.481 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.610 |
0.088 |
0.000 |
| y |
0.088 |
5.047 |
0.000 |
| z |
0.000 |
0.000 |
4.847 |
<r2> (average value of r
2) Å
2
| <r2> |
58.062 |
| (<r2>)1/2 |
7.620 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -135.141374 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
