Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -477.409328 |
| Energy at 298.15K | |
| HF Energy | -476.804696 |
| Nuclear repulsion energy | 107.373838 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3118 |
3040 |
|
|
|
|
| 2 |
A' |
3070 |
2993 |
|
|
|
|
| 3 |
A' |
3041 |
2964 |
|
|
|
|
| 4 |
A' |
2706 |
2637 |
|
|
|
|
| 5 |
A' |
1510 |
1472 |
|
|
|
|
| 6 |
A' |
1496 |
1458 |
|
|
|
|
| 7 |
A' |
1414 |
1378 |
|
|
|
|
| 8 |
A' |
1302 |
1269 |
|
|
|
|
| 9 |
A' |
1115 |
1087 |
|
|
|
|
| 10 |
A' |
1002 |
977 |
|
|
|
|
| 11 |
A' |
861 |
839 |
|
|
|
|
| 12 |
A' |
685 |
667 |
|
|
|
|
| 13 |
A' |
301 |
294 |
|
|
|
|
| 14 |
A" |
3132 |
3053 |
|
|
|
|
| 15 |
A" |
3109 |
3030 |
|
|
|
|
| 16 |
A" |
1500 |
1462 |
|
|
|
|
| 17 |
A" |
1272 |
1240 |
|
|
|
|
| 18 |
A" |
1046 |
1020 |
|
|
|
|
| 19 |
A" |
789 |
769 |
|
|
|
|
| 20 |
A" |
254 |
248 |
|
|
|
|
| 21 |
A" |
176 |
171 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16448.6 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 16034.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.517 |
0.682 |
0.000 |
| C2 |
0.000 |
0.836 |
0.000 |
| S3 |
-0.757 |
-0.835 |
0.000 |
| H4 |
1.997 |
1.663 |
0.000 |
| H5 |
1.852 |
0.137 |
0.884 |
| H6 |
1.852 |
0.137 |
-0.884 |
| H7 |
-0.331 |
1.377 |
0.886 |
| H8 |
-0.331 |
1.377 |
-0.886 |
| H9 |
-2.037 |
-0.434 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5251 | 2.7336 | 1.0916 | 1.0916 | 1.0916 | 2.1641 | 2.1641 | 3.7249 |
C2 | 1.5251 | | 1.8344 | 2.1613 | 2.1684 | 2.1684 | 1.0896 | 1.0896 | 2.4000 | S3 | 2.7336 | 1.8344 | | 3.7177 | 2.9211 | 2.9211 | 2.4207 | 2.4207 | 1.3415 | H4 | 1.0916 | 2.1613 | 3.7177 | | 1.7696 | 1.7696 | 2.5070 | 2.5070 | 4.5458 | H5 | 1.0916 | 2.1684 | 2.9211 | 1.7696 | | 1.7685 | 2.5107 | 3.0722 | 4.0288 | H6 | 1.0916 | 2.1684 | 2.9211 | 1.7696 | 1.7685 | | 3.0722 | 2.5107 | 4.0288 | H7 | 2.1641 | 1.0896 | 2.4207 | 2.5070 | 2.5107 | 3.0722 | | 1.7725 | 2.6407 | H8 | 2.1641 | 1.0896 | 2.4207 | 2.5070 | 3.0722 | 2.5107 | 1.7725 | | 2.6407 | H9 | 3.7249 | 2.4000 | 1.3415 | 4.5458 | 4.0288 | 4.0288 | 2.6407 | 2.6407 | |
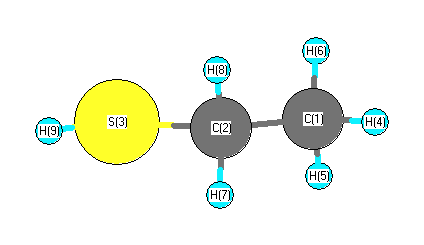 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.564 |
|
C1 |
C2 |
H7 |
110.614 |
| C1 |
C2 |
H8 |
110.614 |
|
C2 |
C1 |
H4 |
110.268 |
| C2 |
C1 |
H5 |
110.840 |
|
C2 |
C1 |
H6 |
110.840 |
| C2 |
S3 |
H9 |
96.944 |
|
S3 |
C2 |
H7 |
109.084 |
| S3 |
C2 |
H8 |
109.084 |
|
H4 |
C1 |
H5 |
108.299 |
| H4 |
C1 |
H6 |
108.299 |
|
H5 |
C1 |
H6 |
108.204 |
| H7 |
C2 |
H8 |
108.850 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -477.409076 |
| Energy at 298.15K | |
| HF Energy | -476.804012 |
| Nuclear repulsion energy | 108.293012 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C1
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
