Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -478.051599 |
| Energy at 298.15K | |
| HF Energy | -478.051599 |
| Nuclear repulsion energy | 107.993828 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3135 |
3001 |
22.19 |
|
|
|
| 2 |
A' |
3085 |
2953 |
20.21 |
|
|
|
| 3 |
A' |
3061 |
2929 |
19.64 |
|
|
|
| 4 |
A' |
2712 |
2596 |
4.08 |
|
|
|
| 5 |
A' |
1503 |
1439 |
2.59 |
|
|
|
| 6 |
A' |
1490 |
1426 |
2.67 |
|
|
|
| 7 |
A' |
1412 |
1351 |
4.16 |
|
|
|
| 8 |
A' |
1296 |
1240 |
28.59 |
|
|
|
| 9 |
A' |
1118 |
1070 |
1.47 |
|
|
|
| 10 |
A' |
1006 |
963 |
3.78 |
|
|
|
| 11 |
A' |
859 |
822 |
0.87 |
|
|
|
| 12 |
A' |
691 |
661 |
1.12 |
|
|
|
| 13 |
A' |
295 |
282 |
2.10 |
|
|
|
| 14 |
A" |
3149 |
3013 |
25.99 |
|
|
|
| 15 |
A" |
3124 |
2990 |
0.37 |
|
|
|
| 16 |
A" |
1492 |
1428 |
9.11 |
|
|
|
| 17 |
A" |
1268 |
1214 |
0.53 |
|
|
|
| 18 |
A" |
1044 |
999 |
0.32 |
|
|
|
| 19 |
A" |
788 |
754 |
3.46 |
|
|
|
| 20 |
A" |
249 |
239 |
0.72 |
|
|
|
| 21 |
A" |
168 |
161 |
15.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16471.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 15765.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.507 |
0.680 |
0.000 |
| C2 |
0.000 |
0.828 |
0.000 |
| S3 |
-0.751 |
-0.830 |
0.000 |
| H4 |
1.982 |
1.659 |
0.000 |
| H5 |
1.847 |
0.139 |
0.881 |
| H6 |
1.847 |
0.139 |
-0.881 |
| H7 |
-0.333 |
1.367 |
0.883 |
| H8 |
-0.333 |
1.367 |
-0.883 |
| H9 |
-2.031 |
-0.433 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5138 | 2.7162 | 1.0882 | 1.0880 | 1.0880 | 2.1525 | 2.1525 | 3.7088 |
C2 | 1.5138 | | 1.8203 | 2.1489 | 2.1589 | 2.1589 | 1.0865 | 1.0865 | 2.3909 | S3 | 2.7162 | 1.8203 | | 3.6966 | 2.9095 | 2.9095 | 2.4047 | 2.4047 | 1.3403 | H4 | 1.0882 | 2.1489 | 3.6966 | | 1.7616 | 1.7616 | 2.4940 | 2.4940 | 4.5257 | H5 | 1.0880 | 2.1589 | 2.9095 | 1.7616 | | 1.7611 | 2.5017 | 3.0606 | 4.0180 | H6 | 1.0880 | 2.1589 | 2.9095 | 1.7616 | 1.7611 | | 3.0606 | 2.5017 | 4.0180 | H7 | 2.1525 | 1.0865 | 2.4047 | 2.4940 | 2.5017 | 3.0606 | | 1.7652 | 2.6279 | H8 | 2.1525 | 1.0865 | 2.4047 | 2.4940 | 3.0606 | 2.5017 | 1.7652 | | 2.6279 | H9 | 3.7088 | 2.3909 | 1.3403 | 4.5257 | 4.0180 | 4.0180 | 2.6279 | 2.6279 | |
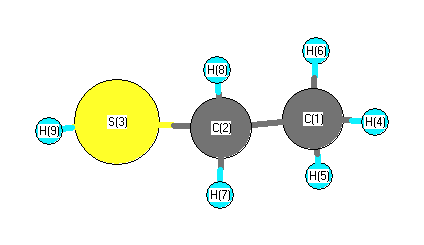 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.765 |
|
C1 |
C2 |
H7 |
110.680 |
| C1 |
C2 |
H8 |
110.680 |
|
C2 |
C1 |
H4 |
110.285 |
| C2 |
C1 |
H5 |
111.099 |
|
C2 |
C1 |
H6 |
111.099 |
| C2 |
S3 |
H9 |
97.140 |
|
S3 |
C2 |
H7 |
109.016 |
| S3 |
C2 |
H8 |
109.016 |
|
H4 |
C1 |
H5 |
108.091 |
| H4 |
C1 |
H6 |
108.091 |
|
H5 |
C1 |
H6 |
108.061 |
| H7 |
C2 |
H8 |
108.652 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.353 |
|
|
|
| 2 |
C |
-0.204 |
|
|
|
| 3 |
S |
-0.189 |
|
|
|
| 4 |
H |
0.120 |
|
|
|
| 5 |
H |
0.127 |
|
|
|
| 6 |
H |
0.127 |
|
|
|
| 7 |
H |
0.132 |
|
|
|
| 8 |
H |
0.132 |
|
|
|
| 9 |
H |
0.109 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.056 |
1.639 |
0.000 |
1.640 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.869 |
-0.238 |
0.000 |
| y |
-0.238 |
-28.011 |
0.000 |
| z |
0.000 |
0.000 |
-28.694 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.483 |
-0.238 |
0.000 |
| y |
-0.238 |
-1.229 |
0.000 |
| z |
0.000 |
0.000 |
-2.254 |
|
| Polar |
|---|
| 3z2-r2 | -4.508 |
|---|
| x2-y2 | 3.141 |
|---|
| xy | -0.238 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.286 |
0.893 |
0.000 |
| y |
0.893 |
6.833 |
0.000 |
| z |
0.000 |
0.000 |
5.202 |
<r2> (average value of r
2) Å
2
| <r2> |
82.467 |
| (<r2>)1/2 |
9.081 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -478.052429 |
| Energy at 298.15K | -478.058703 |
| HF Energy | -478.052429 |
| Nuclear repulsion energy | 107.677567 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3150 |
3015 |
25.27 |
|
|
|
| 2 |
A |
3128 |
2994 |
11.58 |
|
|
|
| 3 |
A |
3120 |
2986 |
17.90 |
|
|
|
| 4 |
A |
3085 |
2953 |
12.34 |
|
|
|
| 5 |
A |
3053 |
2922 |
25.58 |
|
|
|
| 6 |
A |
2715 |
2598 |
3.97 |
|
|
|
| 7 |
A |
1498 |
1434 |
2.84 |
|
|
|
| 8 |
A |
1489 |
1425 |
9.92 |
|
|
|
| 9 |
A |
1478 |
1414 |
1.60 |
|
|
|
| 10 |
A |
1409 |
1349 |
4.11 |
|
|
|
| 11 |
A |
1305 |
1249 |
17.33 |
|
|
|
| 12 |
A |
1281 |
1226 |
3.28 |
|
|
|
| 13 |
A |
1123 |
1075 |
5.88 |
|
|
|
| 14 |
A |
1073 |
1027 |
0.46 |
|
|
|
| 15 |
A |
995 |
953 |
6.67 |
|
|
|
| 16 |
A |
875 |
838 |
5.80 |
|
|
|
| 17 |
A |
736 |
705 |
1.91 |
|
|
|
| 18 |
A |
675 |
646 |
2.97 |
|
|
|
| 19 |
A |
323 |
309 |
1.60 |
|
|
|
| 20 |
A |
256 |
245 |
0.94 |
|
|
|
| 21 |
A |
188 |
180 |
14.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16477.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 15770.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.629 |
-0.348 |
-0.053 |
| C2 |
0.494 |
0.641 |
0.091 |
| S3 |
-1.157 |
-0.100 |
-0.080 |
| H4 |
2.591 |
0.160 |
0.014 |
| H5 |
1.594 |
-1.099 |
0.734 |
| H6 |
1.576 |
-0.864 |
-1.008 |
| H7 |
0.545 |
1.173 |
1.037 |
| H8 |
0.529 |
1.389 |
-0.699 |
| H9 |
-1.070 |
-0.920 |
0.977 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5124 | 2.7968 | 1.0900 | 1.0883 | 1.0870 | 2.1627 | 2.1550 | 2.9448 |
C2 | 1.5124 | | 1.8171 | 2.1525 | 2.1565 | 2.1549 | 1.0867 | 1.0879 | 2.3806 | S3 | 2.7968 | 1.8171 | | 3.7575 | 3.0372 | 2.9857 | 2.4002 | 2.3326 | 1.3409 | H4 | 1.0900 | 2.1525 | 3.7575 | | 1.7607 | 1.7671 | 2.5018 | 2.5031 | 3.9364 | H5 | 1.0883 | 2.1565 | 3.0372 | 1.7607 | | 1.7584 | 2.5206 | 3.0619 | 2.6805 | H6 | 1.0870 | 2.1549 | 2.9857 | 1.7671 | 1.7584 | | 3.0653 | 2.5035 | 3.3088 | H7 | 2.1627 | 1.0867 | 2.4002 | 2.5018 | 2.5206 | 3.0653 | | 1.7489 | 2.6436 | H8 | 2.1550 | 1.0879 | 2.3326 | 2.5031 | 3.0619 | 2.5035 | 1.7489 | | 3.2705 | H9 | 2.9448 | 2.3806 | 1.3409 | 3.9364 | 2.6805 | 3.3088 | 2.6436 | 3.2705 | |
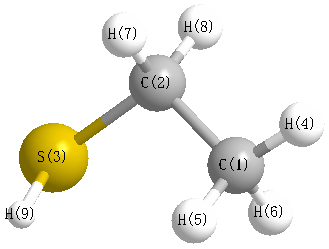 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.968 |
|
C1 |
C2 |
H7 |
111.578 |
| C1 |
C2 |
H8 |
110.886 |
|
C2 |
C1 |
H4 |
110.560 |
| C2 |
C1 |
H5 |
110.985 |
|
C2 |
C1 |
H6 |
110.931 |
| C2 |
S3 |
H9 |
96.687 |
|
S3 |
C2 |
H7 |
108.880 |
| S3 |
C2 |
H8 |
103.985 |
|
H4 |
C1 |
H5 |
107.859 |
| H4 |
C1 |
H6 |
108.525 |
|
H5 |
C1 |
H6 |
107.868 |
| H7 |
C2 |
H8 |
107.070 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.342 |
|
|
|
| 2 |
C |
-0.212 |
|
|
|
| 3 |
S |
-0.186 |
|
|
|
| 4 |
H |
0.117 |
|
|
|
| 5 |
H |
0.118 |
|
|
|
| 6 |
H |
0.130 |
|
|
|
| 7 |
H |
0.137 |
|
|
|
| 8 |
H |
0.130 |
|
|
|
| 9 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.507 |
0.143 |
0.675 |
1.657 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.923 |
1.267 |
-0.665 |
| y |
1.267 |
-26.997 |
-1.617 |
| z |
-0.665 |
-1.617 |
-26.775 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.038 |
1.267 |
-0.665 |
| y |
1.267 |
0.852 |
-1.617 |
| z |
-0.665 |
-1.617 |
1.186 |
|
| Polar |
|---|
| 3z2-r2 | 2.371 |
|---|
| x2-y2 | -1.926 |
|---|
| xy | 1.267 |
|---|
| xz | -0.665 |
|---|
| yz | -1.617 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.852 |
0.376 |
0.027 |
| y |
0.376 |
5.979 |
-0.262 |
| z |
0.027 |
-0.262 |
5.661 |
<r2> (average value of r
2) Å
2
| <r2> |
83.188 |
| (<r2>)1/2 |
9.121 |
