Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.642587 |
| Energy at 298.15K | |
| HF Energy | -169.001079 |
| Nuclear repulsion energy | 71.152600 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3759 |
3566 |
|
|
|
|
| 2 |
A' |
3614 |
3429 |
|
|
|
|
| 3 |
A' |
2980 |
2827 |
|
|
|
|
| 4 |
A' |
1802 |
1710 |
|
|
|
|
| 5 |
A' |
1618 |
1535 |
|
|
|
|
| 6 |
A' |
1424 |
1351 |
|
|
|
|
| 7 |
A' |
1276 |
1210 |
|
|
|
|
| 8 |
A' |
1051 |
997 |
|
|
|
|
| 9 |
A' |
563 |
534 |
|
|
|
|
| 10 |
A" |
1050 |
996 |
|
|
|
|
| 11 |
A" |
637 |
605 |
|
|
|
|
| 12 |
A" |
133i |
126i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9820.4 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 9315.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.420 |
0.000 |
| O2 |
1.200 |
0.233 |
0.000 |
| N3 |
-0.941 |
-0.563 |
0.000 |
| H4 |
-0.450 |
1.427 |
0.000 |
| H5 |
-0.646 |
-1.524 |
0.000 |
| H6 |
-1.919 |
-0.343 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2147 | 1.3610 | 1.1027 | 2.0489 | 2.0653 |
O2 | 1.2147 | | 2.2847 | 2.0366 | 2.5494 | 3.1723 | N3 | 1.3610 | 2.2847 | | 2.0496 | 1.0051 | 1.0025 | H4 | 1.1027 | 2.0366 | 2.0496 | | 2.9574 | 2.3001 | H5 | 2.0489 | 2.5494 | 1.0051 | 2.9574 | | 1.7363 | H6 | 2.0653 | 3.1723 | 1.0025 | 2.3001 | 1.7363 | |
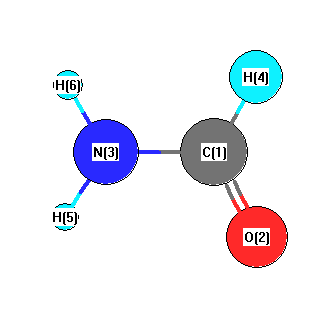 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.211 |
|
C1 |
N3 |
H6 |
121.068 |
| O2 |
C1 |
N3 |
124.906 |
|
O2 |
C1 |
H4 |
122.928 |
| N3 |
C1 |
H4 |
112.166 |
|
H5 |
N3 |
H6 |
119.721 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.642608 |
| Energy at 298.15K | -169.646268 |
| HF Energy | -169.000997 |
| Nuclear repulsion energy | 71.127540 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3748 |
3556 |
|
|
|
|
| 2 |
A |
3607 |
3422 |
|
|
|
|
| 3 |
A |
2981 |
2828 |
|
|
|
|
| 4 |
A |
1803 |
1710 |
|
|
|
|
| 5 |
A |
1620 |
1537 |
|
|
|
|
| 6 |
A |
1424 |
1351 |
|
|
|
|
| 7 |
A |
1274 |
1209 |
|
|
|
|
| 8 |
A |
1062 |
1008 |
|
|
|
|
| 9 |
A |
1042 |
989 |
|
|
|
|
| 10 |
A |
630 |
597 |
|
|
|
|
| 11 |
A |
561 |
533 |
|
|
|
|
| 12 |
A |
201 |
191 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9976.8 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 9464.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.161 |
0.388 |
0.001 |
| O2 |
1.197 |
-0.246 |
0.005 |
| N3 |
-1.087 |
-0.159 |
-0.033 |
| H4 |
0.134 |
1.490 |
0.001 |
| H5 |
-1.175 |
-1.156 |
0.070 |
| H6 |
-1.895 |
0.417 |
0.116 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2144 | 1.3638 | 1.1026 | 2.0433 | 2.0598 |
O2 | 1.2144 | | 2.2864 | 2.0359 | 2.5419 | 3.1643 | N3 | 1.3638 | 2.2864 | | 2.0527 | 1.0058 | 1.0033 | H4 | 1.1026 | 2.0359 | 2.0527 | | 2.9531 | 2.2983 | H5 | 2.0433 | 2.5419 | 1.0058 | 2.9531 | | 1.7300 | H6 | 2.0598 | 3.1643 | 1.0033 | 2.2983 | 1.7300 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.376 |
|
C1 |
N3 |
H6 |
120.195 |
| O2 |
C1 |
N3 |
124.858 |
|
O2 |
C1 |
H4 |
122.895 |
| N3 |
C1 |
H4 |
112.231 |
|
H5 |
N3 |
H6 |
118.876 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
