Jump to
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.916376 |
| Energy at 298.15K | -169.920079 |
| HF Energy | -169.916376 |
| Nuclear repulsion energy | 71.554527 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3768 |
3614 |
43.76 |
|
|
|
| 2 |
A' |
3626 |
3478 |
35.53 |
|
|
|
| 3 |
A' |
2959 |
2838 |
100.47 |
|
|
|
| 4 |
A' |
1838 |
1763 |
398.57 |
|
|
|
| 5 |
A' |
1622 |
1556 |
67.44 |
|
|
|
| 6 |
A' |
1427 |
1369 |
5.05 |
|
|
|
| 7 |
A' |
1286 |
1234 |
101.02 |
|
|
|
| 8 |
A' |
1063 |
1020 |
3.19 |
|
|
|
| 9 |
A' |
569 |
546 |
11.33 |
|
|
|
| 10 |
A" |
1054 |
1011 |
1.43 |
|
|
|
| 11 |
A" |
652 |
625 |
15.49 |
|
|
|
| 12 |
A" |
218 |
209 |
202.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10040.8 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 9631.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| O2 |
1.192 |
0.235 |
0.000 |
| N3 |
-0.933 |
-0.562 |
0.000 |
| H4 |
-0.452 |
1.424 |
0.000 |
| H5 |
-0.640 |
-1.522 |
0.000 |
| H6 |
-1.911 |
-0.347 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2056 | 1.3516 | 1.1049 | 2.0410 | 2.0581 |
O2 | 1.2056 | | 2.2694 | 2.0288 | 2.5381 | 3.1574 | N3 | 1.3516 | 2.2694 | | 2.0436 | 1.0042 | 1.0014 | H4 | 1.1049 | 2.0288 | 2.0436 | | 2.9524 | 2.2955 | H5 | 2.0410 | 2.5381 | 1.0042 | 2.9524 | | 1.7312 | H6 | 2.0581 | 3.1574 | 1.0014 | 2.2955 | 1.7312 | |
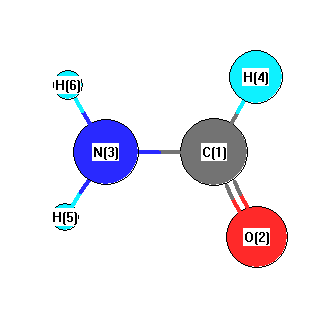 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.350 |
|
C1 |
N3 |
H6 |
121.297 |
| O2 |
C1 |
N3 |
125.023 |
|
O2 |
C1 |
H4 |
122.772 |
| N3 |
C1 |
H4 |
112.205 |
|
H5 |
N3 |
H6 |
119.354 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.180 |
|
|
|
| 2 |
O |
-0.326 |
|
|
|
| 3 |
N |
-0.183 |
|
|
|
| 4 |
H |
0.021 |
|
|
|
| 5 |
H |
0.158 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.852 |
-0.723 |
0.000 |
3.919 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.047 |
-0.053 |
0.000 |
| y |
-0.053 |
-14.683 |
0.000 |
| z |
0.000 |
0.000 |
-18.596 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.408 |
-0.053 |
0.000 |
| y |
-0.053 |
3.639 |
0.000 |
| z |
0.000 |
0.000 |
-2.231 |
|
| Polar |
|---|
| 3z2-r2 | -4.462 |
|---|
| x2-y2 | -3.365 |
|---|
| xy | -0.053 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.515 |
0.277 |
0.000 |
| y |
0.277 |
3.672 |
0.000 |
| z |
0.000 |
0.000 |
2.199 |
<r2> (average value of r
2) Å
2
| <r2> |
40.567 |
| (<r2>)1/2 |
6.369 |
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.916375 |
| Energy at 298.15K | -169.920071 |
| HF Energy | -169.916375 |
| Nuclear repulsion energy | 71.556015 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3768 |
3615 |
43.78 |
|
|
|
| 2 |
A |
3626 |
3478 |
35.54 |
|
|
|
| 3 |
A |
2960 |
2839 |
100.39 |
|
|
|
| 4 |
A |
1838 |
1763 |
398.58 |
|
|
|
| 5 |
A |
1622 |
1556 |
67.34 |
|
|
|
| 6 |
A |
1426 |
1368 |
4.99 |
|
|
|
| 7 |
A |
1286 |
1234 |
100.99 |
|
|
|
| 8 |
A |
1063 |
1020 |
3.14 |
|
|
|
| 9 |
A |
1055 |
1012 |
1.44 |
|
|
|
| 10 |
A |
651 |
625 |
15.82 |
|
|
|
| 11 |
A |
569 |
546 |
11.33 |
|
|
|
| 12 |
A |
213 |
204 |
202.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10038.5 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 9628.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.161 |
0.384 |
0.000 |
| O2 |
-1.190 |
-0.244 |
0.000 |
| N3 |
1.078 |
-0.157 |
-0.000 |
| H4 |
-0.134 |
1.488 |
-0.000 |
| H5 |
1.178 |
-1.156 |
0.001 |
| H6 |
1.897 |
0.418 |
0.001 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2056 | 1.3515 | 1.1048 | 2.0408 | 2.0582 |
O2 | 1.2056 | | 2.2693 | 2.0287 | 2.5377 | 3.1574 | N3 | 1.3515 | 2.2693 | | 2.0437 | 1.0042 | 1.0014 | H4 | 1.1048 | 2.0287 | 2.0437 | | 2.9524 | 2.2959 | H5 | 2.0408 | 2.5377 | 1.0042 | 2.9524 | | 1.7311 | H6 | 2.0582 | 3.1574 | 1.0014 | 2.2959 | 1.7311 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.340 |
|
C1 |
N3 |
H6 |
121.314 |
| O2 |
C1 |
N3 |
125.013 |
|
O2 |
C1 |
H4 |
122.761 |
| N3 |
C1 |
H4 |
112.225 |
|
H5 |
N3 |
H6 |
119.346 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.180 |
|
|
|
| 2 |
O |
-0.326 |
|
|
|
| 3 |
N |
-0.183 |
|
|
|
| 4 |
H |
0.021 |
|
|
|
| 5 |
H |
0.158 |
|
|
|
| 6 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.832 |
0.823 |
0.003 |
3.920 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.582 |
-1.163 |
0.005 |
| y |
-1.163 |
-15.148 |
-0.001 |
| z |
0.005 |
-0.001 |
-18.596 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.710 |
-1.163 |
0.005 |
| y |
-1.163 |
2.941 |
-0.001 |
| z |
0.005 |
-0.001 |
-2.231 |
|
| Polar |
|---|
| 3z2-r2 | -4.463 |
|---|
| x2-y2 | -2.434 |
|---|
| xy | -1.163 |
|---|
| xz | 0.005 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.586 |
0.107 |
0.000 |
| y |
0.107 |
3.600 |
0.000 |
| z |
0.000 |
0.000 |
2.199 |
<r2> (average value of r
2) Å
2
| <r2> |
40.565 |
| (<r2>)1/2 |
6.369 |
