Jump to
S1C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.690991 |
| Energy at 298.15K | -169.694478 |
| HF Energy | -169.001183 |
| Nuclear repulsion energy | 71.222043 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3765 |
3625 |
|
|
|
|
| 2 |
A' |
3626 |
3491 |
|
|
|
|
| 3 |
A' |
2986 |
2875 |
|
|
|
|
| 4 |
A' |
1788 |
1721 |
|
|
|
|
| 5 |
A' |
1629 |
1569 |
|
|
|
|
| 6 |
A' |
1428 |
1375 |
|
|
|
|
| 7 |
A' |
1281 |
1233 |
|
|
|
|
| 8 |
A' |
1047 |
1009 |
|
|
|
|
| 9 |
A' |
562 |
541 |
|
|
|
|
| 10 |
A" |
1059 |
1020 |
|
|
|
|
| 11 |
A" |
646 |
622 |
|
|
|
|
| 12 |
A" |
90 |
86 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9953.2 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 9583.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.418 |
0.000 |
| O2 |
1.201 |
0.229 |
0.000 |
| N3 |
-0.942 |
-0.559 |
0.000 |
| H4 |
-0.447 |
1.422 |
0.000 |
| H5 |
-0.652 |
-1.520 |
0.000 |
| H6 |
-1.918 |
-0.336 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2162 | 1.3571 | 1.0986 | 2.0448 | 2.0612 |
O2 | 1.2162 | | 2.2835 | 2.0347 | 2.5483 | 3.1702 | N3 | 1.3571 | 2.2835 | | 2.0413 | 1.0037 | 1.0010 | H4 | 1.0986 | 2.0347 | 2.0413 | | 2.9486 | 2.2924 | H5 | 2.0448 | 2.5483 | 1.0037 | 2.9486 | | 1.7326 | H6 | 2.0612 | 3.1702 | 1.0010 | 2.2924 | 1.7326 | |
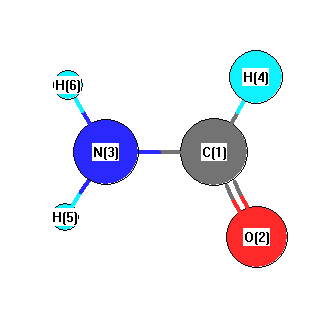 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.272 |
|
C1 |
N3 |
H6 |
121.134 |
| O2 |
C1 |
N3 |
125.002 |
|
O2 |
C1 |
H4 |
122.968 |
| N3 |
C1 |
H4 |
112.029 |
|
H5 |
N3 |
H6 |
119.595 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.690990 |
| Energy at 298.15K | -169.694487 |
| HF Energy | -169.001194 |
| Nuclear repulsion energy | 71.222124 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3766 |
3626 |
|
|
|
|
| 2 |
A |
3628 |
3493 |
|
|
|
|
| 3 |
A |
2985 |
2875 |
|
|
|
|
| 4 |
A |
1788 |
1721 |
|
|
|
|
| 5 |
A |
1630 |
1570 |
|
|
|
|
| 6 |
A |
1428 |
1375 |
|
|
|
|
| 7 |
A |
1280 |
1233 |
|
|
|
|
| 8 |
A |
1059 |
1020 |
|
|
|
|
| 9 |
A |
1047 |
1008 |
|
|
|
|
| 10 |
A |
644 |
620 |
|
|
|
|
| 11 |
A |
560 |
539 |
|
|
|
|
| 12 |
A |
96 |
93 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9955.7 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 9586.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.159 |
0.387 |
0.000 |
| O2 |
1.199 |
-0.244 |
0.001 |
| N3 |
-1.083 |
-0.159 |
-0.005 |
| H4 |
0.127 |
1.485 |
0.001 |
| H5 |
-1.181 |
-1.158 |
0.012 |
| H6 |
-1.902 |
0.416 |
0.018 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2162 | 1.3571 | 1.0986 | 2.0452 | 2.0614 |
O2 | 1.2162 | | 2.2835 | 2.0345 | 2.5493 | 3.1703 | N3 | 1.3571 | 2.2835 | | 2.0415 | 1.0035 | 1.0009 | H4 | 1.0986 | 2.0345 | 2.0415 | | 2.9488 | 2.2933 | H5 | 2.0452 | 2.5493 | 1.0035 | 2.9488 | | 1.7310 | H6 | 2.0614 | 3.1703 | 1.0009 | 2.2933 | 1.7310 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.325 |
|
C1 |
N3 |
H6 |
121.164 |
| O2 |
C1 |
N3 |
125.009 |
|
O2 |
C1 |
H4 |
122.943 |
| N3 |
C1 |
H4 |
112.047 |
|
H5 |
N3 |
H6 |
119.443 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
