Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.892102 |
| Energy at 298.15K | -169.895801 |
| HF Energy | -169.892102 |
| Nuclear repulsion energy | 71.633820 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3766 |
3604 |
43.92 |
|
|
|
| 2 |
A' |
3622 |
3467 |
36.57 |
|
|
|
| 3 |
A' |
2961 |
2834 |
102.24 |
|
|
|
| 4 |
A' |
1841 |
1762 |
401.17 |
|
|
|
| 5 |
A' |
1619 |
1549 |
68.89 |
|
|
|
| 6 |
A' |
1426 |
1364 |
4.51 |
|
|
|
| 7 |
A' |
1284 |
1229 |
101.48 |
|
|
|
| 8 |
A' |
1060 |
1015 |
2.95 |
|
|
|
| 9 |
A' |
565 |
541 |
11.45 |
|
|
|
| 10 |
A" |
1053 |
1008 |
1.30 |
|
|
|
| 11 |
A" |
649 |
622 |
15.56 |
|
|
|
| 12 |
A" |
217 |
208 |
201.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10031.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 9601.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| O2 |
1.190 |
0.234 |
0.000 |
| N3 |
-0.932 |
-0.561 |
0.000 |
| H4 |
-0.451 |
1.423 |
0.000 |
| H5 |
-0.638 |
-1.520 |
0.000 |
| H6 |
-1.910 |
-0.348 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2043 | 1.3504 | 1.1027 | 2.0388 | 2.0566 |
O2 | 1.2043 | | 2.2667 | 2.0261 | 2.5343 | 3.1541 | N3 | 1.3504 | 2.2667 | | 2.0411 | 1.0031 | 1.0004 | H4 | 1.1027 | 2.0261 | 2.0411 | | 2.9487 | 2.2941 | H5 | 2.0388 | 2.5343 | 1.0031 | 2.9487 | | 1.7292 | H6 | 2.0566 | 3.1541 | 1.0004 | 2.2941 | 1.7292 | |
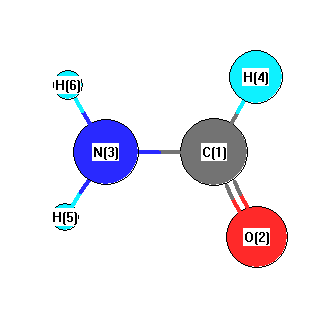 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.324 |
|
C1 |
N3 |
H6 |
121.350 |
| O2 |
C1 |
N3 |
124.969 |
|
O2 |
C1 |
H4 |
122.800 |
| N3 |
C1 |
H4 |
112.231 |
|
H5 |
N3 |
H6 |
119.327 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.154 |
|
|
|
| 2 |
O |
-0.328 |
|
|
|
| 3 |
N |
-0.226 |
|
|
|
| 4 |
H |
0.045 |
|
|
|
| 5 |
H |
0.180 |
|
|
|
| 6 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.837 |
-0.719 |
0.000 |
3.904 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.045 |
-0.047 |
0.000 |
| y |
-0.047 |
-14.696 |
0.000 |
| z |
0.000 |
0.000 |
-18.577 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.408 |
-0.047 |
0.000 |
| y |
-0.047 |
3.615 |
0.000 |
| z |
0.000 |
0.000 |
-2.207 |
|
| Polar |
|---|
| 3z2-r2 | -4.413 |
|---|
| x2-y2 | -3.348 |
|---|
| xy | -0.047 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.507 |
0.274 |
0.000 |
| y |
0.274 |
3.670 |
0.000 |
| z |
0.000 |
0.000 |
2.189 |
<r2> (average value of r
2) Å
2
| <r2> |
40.497 |
| (<r2>)1/2 |
6.364 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.892103 |
| Energy at 298.15K | -169.895798 |
| HF Energy | -169.892103 |
| Nuclear repulsion energy | 71.635628 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3766 |
3604 |
43.92 |
|
|
|
| 2 |
A |
3623 |
3467 |
36.58 |
|
|
|
| 3 |
A |
2962 |
2835 |
102.22 |
|
|
|
| 4 |
A |
1841 |
1762 |
401.22 |
|
|
|
| 5 |
A |
1618 |
1549 |
68.78 |
|
|
|
| 6 |
A |
1425 |
1364 |
4.48 |
|
|
|
| 7 |
A |
1285 |
1230 |
101.50 |
|
|
|
| 8 |
A |
1060 |
1014 |
2.89 |
|
|
|
| 9 |
A |
1053 |
1008 |
1.32 |
|
|
|
| 10 |
A |
650 |
622 |
16.44 |
|
|
|
| 11 |
A |
565 |
541 |
11.42 |
|
|
|
| 12 |
A |
214 |
205 |
200.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10030.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 9600.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.161 |
0.384 |
-0.000 |
| O2 |
1.188 |
-0.244 |
0.000 |
| N3 |
-1.076 |
-0.157 |
-0.000 |
| H4 |
0.134 |
1.486 |
0.000 |
| H5 |
-1.176 |
-1.155 |
0.001 |
| H6 |
-1.896 |
0.417 |
0.001 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2043 | 1.3502 | 1.1027 | 2.0387 | 2.0566 |
O2 | 1.2043 | | 2.2666 | 2.0260 | 2.5342 | 3.1541 | N3 | 1.3502 | 2.2666 | | 2.0411 | 1.0031 | 1.0003 | H4 | 1.1027 | 2.0260 | 2.0411 | | 2.9486 | 2.2943 | H5 | 2.0387 | 2.5342 | 1.0031 | 2.9486 | | 1.7289 | H6 | 2.0566 | 3.1541 | 1.0003 | 2.2943 | 1.7289 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.328 |
|
C1 |
N3 |
H6 |
121.366 |
| O2 |
C1 |
N3 |
124.966 |
|
O2 |
C1 |
H4 |
122.793 |
| N3 |
C1 |
H4 |
112.242 |
|
H5 |
N3 |
H6 |
119.306 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.154 |
|
|
|
| 2 |
O |
-0.328 |
|
|
|
| 3 |
N |
-0.225 |
|
|
|
| 4 |
H |
0.045 |
|
|
|
| 5 |
H |
0.180 |
|
|
|
| 6 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.818 |
0.820 |
0.004 |
3.905 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.577 |
1.161 |
-0.008 |
| y |
1.161 |
-15.163 |
-0.001 |
| z |
-0.008 |
-0.001 |
-18.577 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.707 |
1.161 |
-0.008 |
| y |
1.161 |
2.914 |
-0.001 |
| z |
-0.008 |
-0.001 |
-2.207 |
|
| Polar |
|---|
| 3z2-r2 | -4.414 |
|---|
| x2-y2 | -2.414 |
|---|
| xy | 1.161 |
|---|
| xz | -0.008 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.577 |
-0.106 |
0.000 |
| y |
-0.106 |
3.600 |
0.000 |
| z |
0.000 |
0.000 |
2.189 |
<r2> (average value of r
2) Å
2
| <r2> |
40.495 |
| (<r2>)1/2 |
6.364 |
