return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6 (Cyclopropane)

using model chemistry: MP4=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D3H 1A1'
Energy calculated at MP4=FULL/cc-pVTZ
 hartrees
Energy at 0K-117.720176
Energy at 298.15K-117.726555
HF Energy-117.101344
Nuclear repulsion energy76.004214
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1' 3180 3062        
2 A1' 1536 1479        
3 A1' 1227 1181        
4 A1" 1168 1124        
5 A2' 1092 1051        
6 A2" 3257 3136        
7 A2" 863 831        
8 E' 3170 3052        
8 E' 3170 3052        
9 E' 1487 1432        
9 E' 1486 1431        
10 E' 1062 1023        
10 E' 1062 1023        
11 E' 901 868        
11 E' 901 868        
12 E" 3237 3117        
12 E" 3236 3116        
13 E" 1221 1176        
13 E" 1221 1176        
14 E" 744 717        
14 E" 744 716        

Unscaled Zero Point Vibrational Energy (zpe) 17981.6 cm-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 17314.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4=FULL/cc-pVTZ
ABC
0.67701 0.67701 0.42323

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4=FULL/cc-pVTZ

Point Group is D3h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.869 0.000
C2 0.752 -0.434 0.000
C3 -0.752 -0.434 0.000
H4 0.000 1.448 0.908
H5 1.254 -0.724 0.908
H6 -1.254 -0.724 0.908
H7 0.000 1.448 -0.908
H8 1.254 -0.724 -0.908
H9 -1.254 -0.724 -0.908

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9
C11.50431.50431.07702.22092.22091.07702.22092.2209
C21.50431.50432.22091.07702.22092.22091.07702.2209
C31.50431.50432.22092.22091.07702.22092.22091.0770
H41.07702.22092.22092.50762.50761.81593.09613.0961
H52.22091.07702.22092.50762.50763.09611.81593.0961
H62.22092.22091.07702.50762.50763.09613.09611.8159
H71.07702.22092.22091.81593.09613.09612.50762.5076
H82.22091.07702.22093.09611.81593.09612.50762.5076
H92.22092.22091.07703.09613.09611.81592.50762.5076

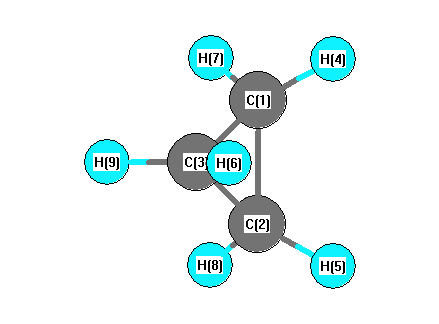
picture of Cyclopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 60.000 C1 C2 H5 117.761
C1 C2 H8 117.761 C1 C3 C2 60.000
C1 C3 H6 117.761 C1 C3 H9 117.761
C2 C1 C3 60.000 C2 C1 H4 117.761
C2 C1 H7 117.761 C2 C3 H6 117.761
C2 C3 H9 117.761 C3 C1 H4 117.761
C3 C1 H7 117.761 C3 C2 H5 117.761
C3 C2 H8 117.761 H4 C1 H7 114.926
H5 C2 H8 114.926 H6 C3 H9 114.926
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability