return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHBrCH3 (i-propyl bromide)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-2691.048753
Energy at 298.15K-2691.059356
HF Energy-2690.192062
Nuclear repulsion energy254.181189
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3189 3027 13.41      
2 A' 3167 3007 24.58      
3 A' 3145 2986 0.56      
4 A' 3089 2933 21.76      
5 A' 1527 1450 8.24      
6 A' 1518 1442 8.79      
7 A' 1427 1355 6.31      
8 A' 1283 1218 37.34      
9 A' 1199 1138 18.99      
10 A' 1075 1021 9.08      
11 A' 926 879 5.10      
12 A' 591 561 10.97      
13 A' 407 387 0.82      
14 A' 313 297 0.49      
15 A' 286 272 0.71      
16 A" 3184 3023 9.00      
17 A" 3159 2999 0.47      
18 A" 3086 2930 9.55      
19 A" 1506 1430 0.04      
20 A" 1505 1429 3.27      
21 A" 1414 1342 11.43      
22 A" 1371 1302 0.22      
23 A" 1175 1115 0.77      
24 A" 969 920 0.06      
25 A" 946 898 1.22      
26 A" 294 279 0.38      
27 A" 267 254 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 21009.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19946.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.27362 0.09958 0.07839

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.904 -0.406 0.000
Br2 -0.669 0.736 0.000
H3 1.735 0.287 0.000
C4 0.904 -1.240 1.256
C5 0.904 -1.240 -1.256
H6 0.029 -1.882 1.276
H7 1.793 -1.865 1.276
H8 0.894 -0.618 2.144
H9 0.029 -1.882 -1.276
H10 1.793 -1.865 -1.276
H11 0.894 -0.618 -2.144

Atom - Atom Distances (Å)
  C1 Br2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.94391.08231.50731.50732.13802.13212.15422.13802.13212.1542
Br21.94392.44592.82032.82032.99513.80202.97912.99513.80202.9791
H31.08232.44592.14442.14443.04012.50192.47423.04012.50192.4742
C41.50732.82032.14442.51201.08551.08761.08352.75452.75523.4560
C51.50732.82032.14442.51202.75452.75523.45601.08551.08761.0835
H62.13802.99513.04011.08552.75451.76431.76052.55143.10193.7468
H72.13213.80202.50191.08762.75521.76431.76493.10192.55123.7487
H82.15422.97912.47421.08353.45601.76051.76493.74683.74874.2875
H92.13802.99513.04012.75451.08552.55143.10193.74681.76431.7605
H102.13213.80202.50192.75521.08763.10192.55123.74871.76431.7649
H112.15422.97912.47423.45601.08353.74683.74874.28751.76051.7649

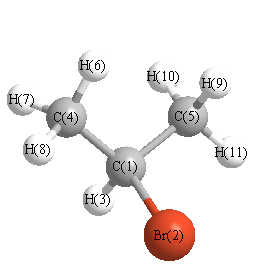
picture of i-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H6 110.032 C1 C4 H7 109.436
C1 C4 H8 111.459 C1 C5 H9 110.032
C1 C5 H10 109.436 C1 C5 H11 111.459
Br2 C1 H3 104.194 Br2 C1 C4 108.955
Br2 C1 C5 108.955 H3 C1 C4 110.735
H3 C1 C5 110.735 C4 C1 C5 112.876
H6 C4 H7 108.564 H6 C4 H8 108.526
H7 C4 H8 108.765 H9 C5 H10 108.564
H9 C5 H11 108.526 H10 C5 H11 108.765
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability