return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH3 (Propane, 2-chloro-)

using model chemistry: CCSD(T)/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCSD(T)/cc-pVTZ
 hartrees
Energy at 0K-578.053179
Energy at 298.15K-578.060875
HF Energy-577.249713
Nuclear repulsion energy166.021818
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3137 3058        
2 A' 3118 3040        
3 A' 3092 3014        
4 A' 3040 2963        
5 A' 1511 1473        
6 A' 1498 1460        
7 A' 1420 1384        
8 A' 1292 1259        
9 A' 1191 1161        
10 A' 1083 1055        
11 A' 909 886        
12 A' 642 626        
13 A' 418 408        
14 A' 333 325        
15 A' 269 262        
16 A" 3133 3054        
17 A" 3111 3032        
18 A" 3037 2960        
19 A" 1491 1454        
20 A" 1487 1449        
21 A" 1407 1372        
22 A" 1363 1329        
23 A" 1158 1129        
24 A" 955 931        
25 A" 937 914        
26 A" 320 311        
27 A" 233 227        

Unscaled Zero Point Vibrational Energy (zpe) 20791.6 cm-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 20267.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD(T)/cc-pVTZ
ABC
0.27002 0.15166 0.10680

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD(T)/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.892 1.013 0.000
C2 0.572 -0.060 0.000
C3 0.572 -0.900 1.266
C4 0.572 -0.900 -1.266
H5 1.416 0.629 0.000
H6 1.472 -1.520 1.293
H7 1.472 -1.520 -1.293
H8 0.555 -0.270 2.156
H9 0.555 -0.270 -2.156
H10 -0.302 -1.554 1.286
H11 -0.302 -1.554 -1.286

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
Cl11.81502.72112.72112.34023.69803.69802.89592.89592.93052.9305
C21.81501.51941.51941.08962.14782.14782.16602.16602.15612.1561
C32.72111.51942.53272.15701.09342.78301.08993.47961.09162.7760
C42.72111.51942.53272.15702.78301.09343.47961.08992.77601.0916
H52.34021.08962.15702.15702.50822.50822.48942.48943.06093.0609
H63.69802.14781.09342.78302.50822.58611.77413.78121.77453.1306
H73.69802.14782.78301.09342.50822.58613.78121.77413.13061.7745
H82.89592.16601.08993.47962.48941.77413.78124.31151.77143.7720
H92.89592.16603.47961.08992.48943.78121.77414.31153.77201.7714
H102.93052.15611.09162.77603.06091.77453.13061.77143.77202.5722
H112.93052.15612.77601.09163.06093.13061.77453.77201.77142.5722

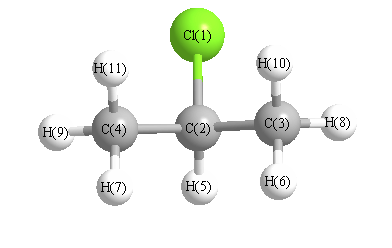
picture of Propane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 109.062 Cl1 C2 C4 109.062
Cl1 C3 H5 55.905 C2 C3 H6 109.495
C2 C3 H8 111.151 C2 C3 H10 110.251
C2 C4 H7 109.495 C2 C4 H9 111.151
C2 C4 H11 110.251 C3 C2 C4 112.903
C3 C2 H5 110.447 C4 C2 H5 110.447
H6 C3 H8 108.703 H6 C3 H10 108.604
H7 C4 H9 108.703 H7 C4 H11 108.604
H8 C3 H10 108.581 H9 C4 H11 108.581
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability