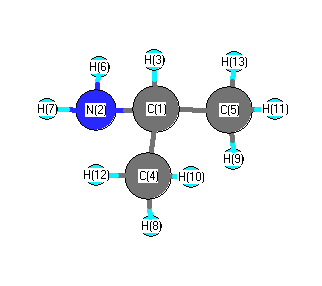
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3528 |
3385 |
0.57 |
|
|
|
| 2 |
A' |
3133 |
3005 |
51.97 |
|
|
|
| 3 |
A' |
3116 |
2989 |
45.30 |
|
|
|
| 4 |
A' |
3048 |
2923 |
5.03 |
|
|
|
| 5 |
A' |
2948 |
2827 |
71.50 |
|
|
|
| 6 |
A' |
1657 |
1590 |
32.44 |
|
|
|
| 7 |
A' |
1509 |
1447 |
6.24 |
|
|
|
| 8 |
A' |
1494 |
1433 |
6.44 |
|
|
|
| 9 |
A' |
1414 |
1356 |
17.70 |
|
|
|
| 10 |
A' |
1380 |
1323 |
16.60 |
|
|
|
| 11 |
A' |
1211 |
1162 |
7.42 |
|
|
|
| 12 |
A' |
1167 |
1120 |
17.12 |
|
|
|
| 13 |
A' |
1004 |
963 |
3.08 |
|
|
|
| 14 |
A' |
849 |
815 |
112.56 |
|
|
|
| 15 |
A' |
831 |
797 |
12.53 |
|
|
|
| 16 |
A' |
472 |
452 |
10.89 |
|
|
|
| 17 |
A' |
357 |
343 |
0.14 |
|
|
|
| 18 |
A' |
259 |
248 |
0.26 |
|
|
|
| 19 |
A" |
3612 |
3465 |
0.03 |
|
|
|
| 20 |
A" |
3127 |
3000 |
0.00 |
|
|
|
| 21 |
A" |
3114 |
2987 |
19.30 |
|
|
|
| 22 |
A" |
3044 |
2920 |
40.65 |
|
|
|
| 23 |
A" |
1489 |
1429 |
0.15 |
|
|
|
| 24 |
A" |
1484 |
1424 |
0.41 |
|
|
|
| 25 |
A" |
1419 |
1361 |
17.28 |
|
|
|
| 26 |
A" |
1386 |
1330 |
1.09 |
|
|
|
| 27 |
A" |
1271 |
1220 |
0.12 |
|
|
|
| 28 |
A" |
1044 |
1001 |
0.95 |
|
|
|
| 29 |
A" |
965 |
926 |
0.39 |
|
|
|
| 30 |
A" |
930 |
892 |
0.35 |
|
|
|
| 31 |
A" |
403 |
387 |
7.10 |
|
|
|
| 32 |
A" |
281 |
270 |
29.46 |
|
|
|
| 33 |
A" |
214 |
205 |
5.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26580.3 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 25495.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.031 |
0.000 |
|
|
| 2 |
N |
-0.300 |
0.000 |
|
|
| 3 |
H |
0.057 |
0.000 |
|
|
| 4 |
C |
-0.247 |
0.000 |
|
|
| 5 |
C |
-0.247 |
0.000 |
|
|
| 6 |
H |
0.113 |
0.000 |
|
|
| 7 |
H |
0.113 |
0.000 |
|
|
| 8 |
H |
0.089 |
0.000 |
|
|
| 9 |
H |
0.089 |
0.000 |
|
|
| 10 |
H |
0.077 |
0.000 |
|
|
| 11 |
H |
0.077 |
0.000 |
|
|
| 12 |
H |
0.075 |
0.000 |
|
|
| 13 |
H |
0.075 |
0.000 |
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.140 |
1.224 |
0.000 |
1.232 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.056 |
-2.143 |
0.000 |
| y |
-2.143 |
-28.549 |
0.000 |
| z |
0.000 |
0.000 |
-26.456 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.446 |
-2.143 |
0.000 |
| y |
-2.143 |
-2.292 |
0.000 |
| z |
0.000 |
0.000 |
0.846 |
|
| Polar |
|---|
| 3z2-r2 | 1.692 |
|---|
| x2-y2 | 2.492 |
|---|
| xy | -2.143 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.498 |
-0.443 |
0.000 |
| y |
-0.443 |
6.911 |
0.000 |
| z |
0.000 |
0.000 |
7.346 |
<r2> (average value of r
2) Å
2
| <r2> |
92.619 |
| (<r2>)1/2 |
9.624 |