return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (Propylene oxide)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-192.791962
Energy at 298.15K-192.798933
HF Energy-191.981409
Nuclear repulsion energy125.497387
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3250 3086 18.81      
2 A 3188 3027 28.84      
3 A 3171 3010 7.77      
4 A 3168 3008 11.76      
5 A 3164 3004 17.58      
6 A 3094 2937 13.08      
7 A 1558 1480 4.87      
8 A 1524 1447 6.40      
9 A 1508 1432 5.26      
10 A 1470 1395 19.12      
11 A 1413 1342 4.27      
12 A 1317 1250 3.99      
13 A 1208 1147 0.43      
14 A 1187 1127 3.53      
15 A 1171 1112 0.53      
16 A 1145 1087 5.17      
17 A 1060 1006 9.94      
18 A 992 942 11.15      
19 A 921 875 1.69      
20 A 876 832 37.40      
21 A 792 752 5.72      
22 A 414 393 4.76      
23 A 373 354 3.35      
24 A 224 213 0.48      

Unscaled Zero Point Vibrational Energy (zpe) 19094.9 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18128.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.60755 0.22683 0.20177

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.821 -0.783 -0.240
C2 -1.492 0.097 -0.148
H3 -1.384 0.319 -1.204
H4 -2.054 -0.826 -0.044
H5 -2.055 0.897 0.325
C6 -0.149 -0.042 0.486
H7 -0.151 -0.244 1.547
C8 1.028 0.613 -0.063
H9 0.916 1.220 -0.949
H10 1.840 0.888 0.593

Atom - Atom Distances (Å)
  O1 C2 H3 H4 H5 C6 H7 C8 H9 H10
O12.47712.64742.88183.37871.42032.10471.42292.12732.1273
C22.47711.08471.08551.08601.49162.18832.57342.77553.5036
H32.64741.08471.76171.76672.12383.06722.68442.48393.7344
H42.88181.08551.76171.76192.12682.54793.40073.71824.3014
H53.37871.08601.76671.76192.13102.53433.11993.24923.9040
C61.42031.49162.12382.12682.13101.08091.45382.18742.1980
H72.10472.18833.06722.54792.53431.08092.17153.08442.4811
C81.42292.57342.68443.40073.11991.45382.17151.07981.0792
H92.12732.77552.48393.71823.24922.18743.08441.07981.8272
H102.12733.50363.73444.30143.90402.19802.48111.07921.8272

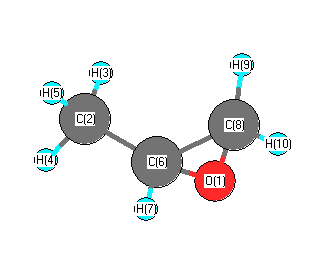
picture of Propylene oxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C6 C2 116.554 O1 C6 H7 113.907
O1 C6 C8 59.336 O1 C8 C6 59.157
O1 C8 H9 115.745 O1 C8 H10 115.788
C2 C6 H7 115.649 C2 C6 C8 121.778
H3 C2 H4 108.545 H3 C2 H5 108.961
H3 C2 C6 110.048 H4 C2 H5 108.457
H4 C2 C6 110.240 H5 C2 C6 110.540
C6 O1 C8 61.507 C6 C8 H9 118.644
C6 C8 H10 119.659 H7 C6 C8 117.140
H9 C8 H10 115.634
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability