return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (Ethanol, 1,1-dimethyl-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-233.279474
Energy at 298.15K-233.290757
HF Energy-232.245702
Nuclear repulsion energy201.052336
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3840 3646 18.59      
2 A' 3179 3019 32.54      
3 A' 3173 3012 4.16      
4 A' 3154 2994 42.11      
5 A' 3095 2939 9.34      
6 A' 3082 2926 23.25      
7 A' 1540 1462 9.94      
8 A' 1529 1452 4.17      
9 A' 1509 1433 0.05      
10 A' 1428 1356 6.84      
11 A' 1414 1342 36.11      
12 A' 1372 1303 17.49      
13 A' 1256 1192 40.67      
14 A' 1172 1113 60.73      
15 A' 1037 984 9.20      
16 A' 960 911 34.68      
17 A' 941 894 0.89      
18 A' 774 735 2.71      
19 A' 456 433 9.40      
20 A' 409 388 0.16      
21 A' 344 326 1.09      
22 A' 287 272 0.07      
23 A" 3177 3016 25.37      
24 A" 3171 3011 3.10      
25 A" 3149 2990 2.32      
26 A" 3079 2923 13.67      
27 A" 1522 1445 3.27      
28 A" 1505 1429 0.06      
29 A" 1495 1420 0.02      
30 A" 1402 1331 22.60      
31 A" 1286 1221 15.44      
32 A" 1053 999 0.89      
33 A" 959 910 0.00      
34 A" 928 881 0.12      
35 A" 464 440 11.78      
36 A" 347 329 15.54      
37 A" 312 296 66.34      
38 A" 271 258 18.18      
39 A" 211 200 1.94      

Unscaled Zero Point Vibrational Energy (zpe) 30138.9 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 28613.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.15966 0.15873 0.15349

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.003 0.020 0.000
O2 -0.478 1.361 0.000
C3 1.502 0.138 0.000
C4 -0.478 -0.701 1.245
C5 -0.478 -0.701 -1.245
H6 -1.438 1.326 0.000
H7 1.959 -0.846 0.000
H8 1.828 0.680 -0.881
H9 1.828 0.680 0.881
H10 -0.086 -1.713 1.285
H11 -0.086 -1.713 -1.285
H12 -0.153 -0.161 2.128
H13 -1.564 -0.758 1.255
H14 -0.153 -0.161 -2.128
H15 -1.564 -0.758 -1.255

Atom - Atom Distances (Å)
  C1 O2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.42231.50891.51521.51521.94022.14482.13612.13612.15922.15922.14082.14902.14082.1490
O21.42232.32712.40832.40830.96053.28822.56102.56103.35463.35462.63612.69132.63612.6913
C31.50892.32712.48452.48453.17071.08531.08501.08502.75622.75622.71183.43172.71183.4317
C41.51522.40832.48452.48972.56492.74063.42712.71221.08632.75271.08501.08773.43112.7258
C51.51522.40832.48452.48972.56492.74062.71223.42712.75271.08633.43112.72581.08501.0877
H61.94020.96053.17072.56492.56494.03233.44393.44393.56573.56572.89662.43582.89662.4358
H72.14483.28821.08532.74062.74064.03231.76741.76742.56592.56593.07533.74113.07533.7411
H82.13612.56101.08503.42712.71223.44391.76741.76293.75253.09053.69944.25882.48653.7032
H92.13612.56101.08502.71223.42713.44391.76741.76293.09053.75252.48653.70323.69944.2588
H102.15923.35462.75621.08632.75273.56572.56593.75253.09052.56961.76761.76023.74973.0896
H112.15923.35462.75622.75271.08633.56572.56593.09053.75252.56963.74973.08961.76761.7602
H122.14082.63612.71181.08503.43112.89663.07533.69942.48651.76763.74971.76414.25573.7135
H132.14902.69133.43171.08772.72582.43583.74114.25883.70321.76023.08961.76413.71352.5090
H142.14082.63612.71183.43111.08502.89663.07532.48653.69943.74971.76764.25573.71351.7641
H152.14902.69133.43172.72581.08772.43583.74113.70324.25883.08961.76023.71352.50901.7641

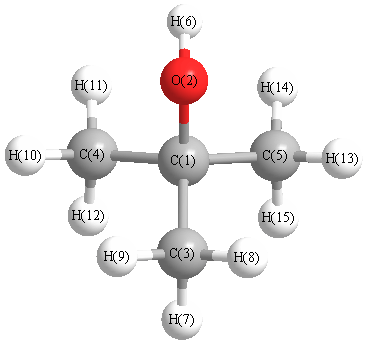
picture of Ethanol, 1,1-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H2 33.682 C1 C5 H3 34.679
C1 C5 H4 34.757 C1 C9 H6 30.640
C1 C9 H7 65.871 C1 C9 H8 65.629
C1 C13 H10 66.189 C1 C13 H11 44.314
C1 C13 H12 65.475 C1 O15 H14 65.475
H2 C5 H3 56.775 H2 C5 H4 58.876
H3 C5 H4 59.930 C5 C1 C9 139.014
C5 C1 C13 94.542 C5 C1 O15 28.354
H6 C9 H7 96.013 H6 C9 H8 75.170
H7 C9 H8 60.084 C9 C1 C13 119.582
C9 C1 O15 167.274 H10 C13 H11 56.260
H10 C13 H12 60.205 H11 C13 H12 97.391
C13 C1 O15 71.431
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability