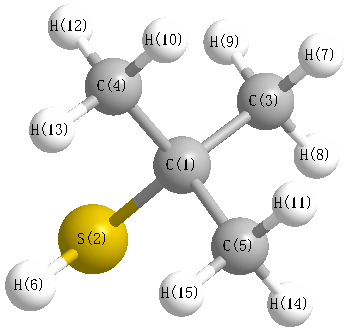
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3143 |
3008 |
23.95 |
|
|
|
| 2 |
A' |
3127 |
2993 |
25.12 |
|
|
|
| 3 |
A' |
3116 |
2983 |
41.88 |
|
|
|
| 4 |
A' |
3054 |
2923 |
23.16 |
|
|
|
| 5 |
A' |
3045 |
2915 |
36.04 |
|
|
|
| 6 |
A' |
2719 |
2602 |
5.68 |
|
|
|
| 7 |
A' |
1509 |
1444 |
7.36 |
|
|
|
| 8 |
A' |
1495 |
1431 |
9.91 |
|
|
|
| 9 |
A' |
1482 |
1418 |
0.28 |
|
|
|
| 10 |
A' |
1421 |
1360 |
1.31 |
|
|
|
| 11 |
A' |
1394 |
1334 |
12.03 |
|
|
|
| 12 |
A' |
1264 |
1210 |
0.46 |
|
|
|
| 13 |
A' |
1197 |
1146 |
38.64 |
|
|
|
| 14 |
A' |
1055 |
1010 |
1.68 |
|
|
|
| 15 |
A' |
945 |
904 |
0.37 |
|
|
|
| 16 |
A' |
882 |
844 |
2.92 |
|
|
|
| 17 |
A' |
835 |
799 |
1.14 |
|
|
|
| 18 |
A' |
594 |
568 |
4.15 |
|
|
|
| 19 |
A' |
369 |
353 |
0.97 |
|
|
|
| 20 |
A' |
351 |
336 |
0.18 |
|
|
|
| 21 |
A' |
285 |
272 |
0.32 |
|
|
|
| 22 |
A' |
269 |
257 |
0.54 |
|
|
|
| 23 |
A" |
3141 |
3007 |
22.07 |
|
|
|
| 24 |
A" |
3136 |
3002 |
8.03 |
|
|
|
| 25 |
A" |
3113 |
2979 |
0.96 |
|
|
|
| 26 |
A" |
3042 |
2911 |
18.89 |
|
|
|
| 27 |
A" |
1497 |
1433 |
8.46 |
|
|
|
| 28 |
A" |
1478 |
1415 |
0.20 |
|
|
|
| 29 |
A" |
1470 |
1407 |
0.01 |
|
|
|
| 30 |
A" |
1395 |
1335 |
12.70 |
|
|
|
| 31 |
A" |
1252 |
1198 |
2.93 |
|
|
|
| 32 |
A" |
1045 |
1000 |
0.20 |
|
|
|
| 33 |
A" |
961 |
920 |
0.01 |
|
|
|
| 34 |
A" |
939 |
898 |
0.13 |
|
|
|
| 35 |
A" |
382 |
366 |
0.90 |
|
|
|
| 36 |
A" |
303 |
290 |
7.78 |
|
|
|
| 37 |
A" |
271 |
259 |
0.56 |
|
|
|
| 38 |
A" |
260 |
249 |
4.99 |
|
|
|
| 39 |
A" |
217 |
207 |
1.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28725.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 27492.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.022 |
|
|
|
| 2 |
S |
-0.187 |
|
|
|
| 3 |
C |
-0.353 |
|
|
|
| 4 |
C |
-0.345 |
|
|
|
| 5 |
C |
-0.345 |
|
|
|
| 6 |
H |
0.109 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.133 |
|
|
|
| 9 |
H |
0.133 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.118 |
|
|
|
| 12 |
H |
0.135 |
|
|
|
| 13 |
H |
0.125 |
|
|
|
| 14 |
H |
0.135 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.521 |
-0.694 |
0.000 |
1.672 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.208 |
1.994 |
0.000 |
| y |
1.994 |
-39.133 |
0.000 |
| z |
0.000 |
0.000 |
-41.956 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.664 |
1.994 |
0.000 |
| y |
1.994 |
2.950 |
0.000 |
| z |
0.000 |
0.000 |
-1.286 |
|
| Polar |
|---|
| 3z2-r2 | -2.571 |
|---|
| x2-y2 | -3.076 |
|---|
| xy | 1.994 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.156 |
0.108 |
0.000 |
| y |
0.108 |
9.795 |
0.000 |
| z |
0.000 |
0.000 |
9.281 |
<r2> (average value of r
2) Å
2
| <r2> |
163.080 |
| (<r2>)1/2 |
12.770 |