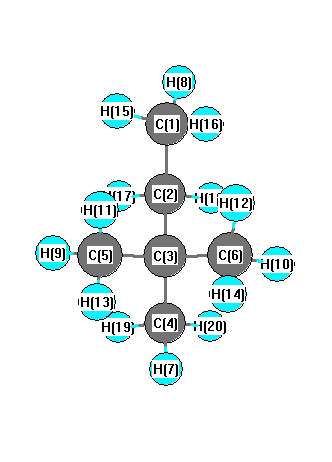
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3130 |
2996 |
47.69 |
|
|
|
| 2 |
A' |
3124 |
2990 |
56.71 |
|
|
|
| 3 |
A' |
3117 |
2983 |
69.16 |
|
|
|
| 4 |
A' |
3112 |
2979 |
1.46 |
|
|
|
| 5 |
A' |
3060 |
2929 |
21.49 |
|
|
|
| 6 |
A' |
3049 |
2918 |
11.17 |
|
|
|
| 7 |
A' |
3039 |
2909 |
35.63 |
|
|
|
| 8 |
A' |
3032 |
2902 |
32.61 |
|
|
|
| 9 |
A' |
1518 |
1453 |
10.68 |
|
|
|
| 10 |
A' |
1508 |
1444 |
9.31 |
|
|
|
| 11 |
A' |
1505 |
1441 |
6.70 |
|
|
|
| 12 |
A' |
1487 |
1423 |
0.33 |
|
|
|
| 13 |
A' |
1476 |
1413 |
0.14 |
|
|
|
| 14 |
A' |
1423 |
1362 |
4.35 |
|
|
|
| 15 |
A' |
1410 |
1349 |
7.88 |
|
|
|
| 16 |
A' |
1393 |
1333 |
8.61 |
|
|
|
| 17 |
A' |
1368 |
1309 |
0.48 |
|
|
|
| 18 |
A' |
1285 |
1230 |
2.07 |
|
|
|
| 19 |
A' |
1248 |
1194 |
5.62 |
|
|
|
| 20 |
A' |
1098 |
1051 |
1.34 |
|
|
|
| 21 |
A' |
1040 |
996 |
2.79 |
|
|
|
| 22 |
A' |
1007 |
964 |
2.89 |
|
|
|
| 23 |
A' |
942 |
902 |
0.94 |
|
|
|
| 24 |
A' |
886 |
848 |
0.22 |
|
|
|
| 25 |
A' |
723 |
692 |
0.34 |
|
|
|
| 26 |
A' |
476 |
455 |
0.32 |
|
|
|
| 27 |
A' |
389 |
372 |
0.12 |
|
|
|
| 28 |
A' |
346 |
331 |
0.01 |
|
|
|
| 29 |
A' |
274 |
262 |
0.03 |
|
|
|
| 30 |
A' |
242 |
231 |
0.05 |
|
|
|
| 31 |
A" |
3134 |
2999 |
24.60 |
|
|
|
| 32 |
A" |
3121 |
2987 |
33.05 |
|
|
|
| 33 |
A" |
3114 |
2980 |
33.50 |
|
|
|
| 34 |
A" |
3112 |
2979 |
0.14 |
|
|
|
| 35 |
A" |
3067 |
2936 |
17.87 |
|
|
|
| 36 |
A" |
3043 |
2912 |
40.93 |
|
|
|
| 37 |
A" |
1515 |
1450 |
0.00 |
|
|
|
| 38 |
A" |
1500 |
1436 |
11.27 |
|
|
|
| 39 |
A" |
1481 |
1418 |
0.40 |
|
|
|
| 40 |
A" |
1478 |
1415 |
0.12 |
|
|
|
| 41 |
A" |
1395 |
1335 |
11.38 |
|
|
|
| 42 |
A" |
1336 |
1279 |
0.26 |
|
|
|
| 43 |
A" |
1252 |
1198 |
1.14 |
|
|
|
| 44 |
A" |
1101 |
1054 |
0.18 |
|
|
|
| 45 |
A" |
999 |
956 |
0.52 |
|
|
|
| 46 |
A" |
951 |
911 |
0.06 |
|
|
|
| 47 |
A" |
943 |
902 |
0.21 |
|
|
|
| 48 |
A" |
783 |
750 |
2.39 |
|
|
|
| 49 |
A" |
398 |
381 |
0.10 |
|
|
|
| 50 |
A" |
333 |
319 |
0.00 |
|
|
|
| 51 |
A" |
292 |
280 |
0.00 |
|
|
|
| 52 |
A" |
256 |
245 |
0.00 |
|
|
|
| 53 |
A" |
200 |
192 |
0.01 |
|
|
|
| 54 |
A" |
83 |
80 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41296.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 39524.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.398 |
|
|
|
| 2 |
C |
-0.202 |
|
|
|
| 3 |
C |
0.030 |
|
|
|
| 4 |
C |
-0.365 |
|
|
|
| 5 |
C |
-0.385 |
|
|
|
| 6 |
C |
-0.385 |
|
|
|
| 7 |
H |
0.118 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.124 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
H |
0.126 |
|
|
|
| 12 |
H |
0.126 |
|
|
|
| 13 |
H |
0.122 |
|
|
|
| 14 |
H |
0.122 |
|
|
|
| 15 |
H |
0.122 |
|
|
|
| 16 |
H |
0.122 |
|
|
|
| 17 |
H |
0.119 |
|
|
|
| 18 |
H |
0.119 |
|
|
|
| 19 |
H |
0.119 |
|
|
|
| 20 |
H |
0.119 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.055 |
0.010 |
0.000 |
0.056 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.783 |
0.215 |
0.000 |
| y |
0.215 |
-42.384 |
0.000 |
| z |
0.000 |
0.000 |
-41.894 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.356 |
0.215 |
0.000 |
| y |
0.215 |
-0.546 |
0.000 |
| z |
0.000 |
0.000 |
0.190 |
|
| Polar |
|---|
| 3z2-r2 | 0.380 |
|---|
| x2-y2 | 0.601 |
|---|
| xy | 0.215 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.317 |
-0.497 |
0.000 |
| y |
-0.497 |
10.880 |
0.000 |
| z |
0.000 |
0.000 |
10.519 |
<r2> (average value of r
2) Å
2
| <r2> |
193.634 |
| (<r2>)1/2 |
13.915 |