All results from a given calculation for CF3CHO (trifluoroacetaldehyde)
using model chemistry: QCISD(T)/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -450.996077 |
| Energy at 298.15K | |
| HF Energy | -449.656809 |
| Nuclear repulsion energy | 252.603287 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.019 |
0.362 |
0.000 |
| C2 |
0.502 |
-1.098 |
0.000 |
| O3 |
-0.258 |
-2.031 |
0.000 |
| F4 |
-1.300 |
0.457 |
0.000 |
| F5 |
0.502 |
0.986 |
1.083 |
| F6 |
0.502 |
0.986 |
-1.083 |
| H7 |
1.599 |
-1.198 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
F4 |
F5 |
F6 |
H7 |
| C1 | | 1.5378 | 2.4095 | 1.3225 | 1.3398 | 1.3398 | 2.2208 |
C2 | 1.5378 | | 1.2035 | 2.3801 | 2.3487 | 2.3487 | 1.1022 | O3 | 2.4095 | 1.2035 | | 2.6980 | 3.2948 | 3.2948 | 2.0355 | F4 | 1.3225 | 2.3801 | 2.6980 | | 2.1676 | 2.1676 | 3.3387 | F5 | 1.3398 | 2.3487 | 3.2948 | 2.1676 | | 2.1660 | 2.6739 | F6 | 1.3398 | 2.3487 | 3.2948 | 2.1676 | 2.1660 | | 2.6739 | H7 | 2.2208 | 1.1022 | 2.0355 | 3.3387 | 2.6739 | 2.6739 | |
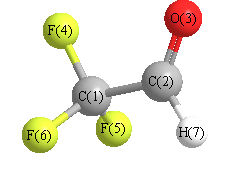 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
122.568 |
|
C1 |
C2 |
H7 |
113.514 |
| C2 |
C1 |
F4 |
112.416 |
|
C2 |
C1 |
F5 |
109.222 |
| C2 |
C1 |
F6 |
109.222 |
|
O3 |
C2 |
H7 |
123.918 |
| F4 |
C1 |
F5 |
109.009 |
|
F4 |
C1 |
F6 |
109.009 |
| F5 |
C1 |
F6 |
107.857 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
