return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((Z)-hex-3-ene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-235.389868
Energy at 298.15K-235.402296
HF Energy-234.259606
Nuclear repulsion energy233.807517
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3199 3038 34.93      
2 A 3168 3007 1.81      
3 A 3164 3004 52.67      
4 A 3140 2981 1.31      
5 A 3091 2935 2.13      
6 A 3089 2933 2.74      
7 A 1728 1641 1.16      
8 A 1536 1458 9.64      
9 A 1524 1447 3.21      
10 A 1519 1442 0.02      
11 A 1416 1344 0.11      
12 A 1357 1288 0.04      
13 A 1319 1252 0.06      
14 A 1284 1219 0.09      
15 A 1106 1050 0.69      
16 A 1086 1031 0.35      
17 A 1041 989 0.38      
18 A 1017 966 0.80      
19 A 866 822 1.06      
20 A 791 751 0.66      
21 A 539 512 0.28      
22 A 330 313 0.01      
23 A 221 210 0.01      
24 A 196 186 0.01      
25 A 39 37 0.00      
26 B 3168 3008 58.39      
27 B 3165 3005 2.96      
28 B 3160 3000 0.71      
29 B 3129 2971 4.63      
30 B 3090 2933 58.89      
31 B 3088 2932 29.86      
32 B 1530 1453 2.89      
33 B 1524 1447 9.70      
34 B 1501 1425 1.45      
35 B 1453 1379 3.74      
36 B 1416 1344 5.37      
37 B 1342 1274 5.12      
38 B 1306 1240 0.08      
39 B 1192 1132 0.11      
40 B 1110 1054 6.76      
41 B 1071 1016 2.29      
42 B 928 881 6.51      
43 B 823 781 3.61      
44 B 780 741 35.52      
45 B 566 537 4.32      
46 B 362 343 1.06      
47 B 251 239 0.04      
48 B 39 37 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 36879.9 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 35013.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.26436 0.05410 0.05140

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.008 0.667 0.892
C2 -0.008 -0.667 0.892
C3 -0.008 1.564 -0.298
C4 0.008 -1.564 -0.298
C5 -1.344 2.283 -0.436
C6 1.344 -2.283 -0.436
H7 0.002 1.166 1.851
H8 -0.002 -1.166 1.851
H9 0.204 1.006 -1.204
H10 -0.204 -1.006 -1.204
H11 0.784 2.303 -0.196
H12 -0.784 -2.303 -0.196
H13 -1.337 2.974 -1.274
H14 -2.145 1.566 -0.588
H15 -1.571 2.847 0.464
H16 1.337 -2.974 -1.274
H17 2.145 -1.566 -0.588
H18 1.571 -2.847 0.464

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33471.49022.52882.49093.50071.08072.06892.13262.69062.11233.26103.43822.76302.72534.44033.42713.8697
C21.33472.52881.49023.50072.49092.06891.08072.69062.13263.26102.11234.44033.42713.86973.43822.76302.7253
C31.49022.52883.12781.52364.08002.18543.47431.08522.73191.08843.94552.16932.15642.16094.83243.80994.7462
C42.52881.49023.12784.08001.52363.47432.18542.73191.08523.94551.08844.83243.80994.74622.16932.15642.1609
C52.49093.50071.52364.08005.29872.87944.35102.14893.56462.14214.62651.08571.08551.08615.96015.19725.9685
C63.50072.49094.08001.52365.29874.35102.87943.56462.14894.62652.14215.96015.19725.96851.08571.08551.0861
H71.08072.06892.18543.47432.87944.35102.33223.06603.75412.46844.10383.85033.27372.68745.35604.24354.5265
H82.06891.08073.47432.18544.35102.87942.33223.75413.06604.10382.46845.35604.24354.52653.85033.27372.6874
H92.13262.69061.08522.73192.14893.56463.06603.75412.05271.74263.59772.50042.49213.05334.13833.28024.4150
H102.69062.13262.73191.08523.56462.14893.75413.06602.05273.59771.74264.13833.28024.41502.50042.49213.0533
H112.11233.26101.08843.94552.14214.62652.46844.10381.74263.59774.86602.47213.04582.50595.41434.12015.2510
H123.26102.11233.94551.08844.62652.14214.10382.46843.59771.74264.86605.41434.12015.25102.47213.04582.5059
H133.43824.44032.16934.83241.08575.96013.85035.35602.50044.13832.47215.41431.76221.75836.52085.76206.7345
H142.76303.42712.15643.80991.08555.19723.27374.24352.49213.28023.04584.12011.76221.75375.76205.31135.8641
H152.72533.86972.16094.74621.08615.96852.68744.52653.05334.41502.50595.25101.75831.75376.73455.86416.5029
H164.44033.43824.83242.16935.96011.08575.35603.85034.13832.50045.41432.47216.52085.76206.73451.76221.7583
H173.42712.76303.80992.15645.19721.08554.24353.27373.28022.49214.12013.04585.76205.31135.86411.76221.7537
H183.86972.72534.74622.16095.96851.08614.52652.68744.41503.05335.25102.50596.73455.86416.50291.75831.7537

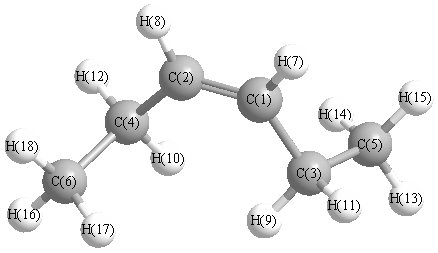
picture of (Z)-hex-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 126.977 C1 C2 H8 117.482
C1 C3 C5 111.476 C1 C3 H9 110.821
C1 C3 H11 109.004 C2 C1 C3 126.977
C2 C1 H7 117.482 C2 C4 C6 111.476
C2 C4 H10 110.821 C2 C4 H12 109.004
C3 C1 H7 115.512 C3 C5 H13 111.375
C3 C5 H14 110.347 C3 C5 H15 110.671
C4 C2 H8 115.512 C4 C6 H16 111.375
C4 C6 H17 110.347 C4 C6 H18 110.671
C5 C3 H9 109.769 C5 C3 H11 109.046
C6 C4 H10 109.769 C6 C4 H12 109.046
H9 C3 H11 106.583 H10 C4 H12 106.583
H13 C5 H14 108.506 H13 C5 H15 108.111
H14 C5 H15 107.714 H16 C6 H17 108.506
H16 C6 H18 108.111 H17 C6 H18 107.714
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability