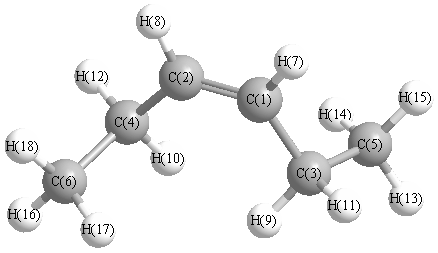
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3158 |
3022 |
53.16 |
|
|
|
| 2 |
A |
3133 |
2999 |
7.67 |
|
|
|
| 3 |
A |
3130 |
2996 |
60.22 |
|
|
|
| 4 |
A |
3109 |
2976 |
0.08 |
|
|
|
| 5 |
A |
3054 |
2923 |
0.14 |
|
|
|
| 6 |
A |
3042 |
2911 |
5.00 |
|
|
|
| 7 |
A |
1751 |
1675 |
1.70 |
|
|
|
| 8 |
A |
1508 |
1443 |
10.32 |
|
|
|
| 9 |
A |
1497 |
1432 |
3.10 |
|
|
|
| 10 |
A |
1488 |
1424 |
0.27 |
|
|
|
| 11 |
A |
1406 |
1346 |
0.28 |
|
|
|
| 12 |
A |
1348 |
1290 |
0.19 |
|
|
|
| 13 |
A |
1306 |
1250 |
0.05 |
|
|
|
| 14 |
A |
1274 |
1219 |
0.16 |
|
|
|
| 15 |
A |
1094 |
1047 |
0.41 |
|
|
|
| 16 |
A |
1074 |
1028 |
0.86 |
|
|
|
| 17 |
A |
1038 |
994 |
0.01 |
|
|
|
| 18 |
A |
1015 |
972 |
1.15 |
|
|
|
| 19 |
A |
857 |
820 |
1.42 |
|
|
|
| 20 |
A |
780 |
746 |
0.81 |
|
|
|
| 21 |
A |
516 |
494 |
0.27 |
|
|
|
| 22 |
A |
310 |
297 |
0.00 |
|
|
|
| 23 |
A |
207 |
198 |
0.00 |
|
|
|
| 24 |
A |
197 |
189 |
0.04 |
|
|
|
| 25 |
A |
61 |
58 |
0.01 |
|
|
|
| 26 |
B |
3135 |
3001 |
44.41 |
|
|
|
| 27 |
B |
3131 |
2997 |
34.33 |
|
|
|
| 28 |
B |
3126 |
2992 |
1.23 |
|
|
|
| 29 |
B |
3098 |
2965 |
3.69 |
|
|
|
| 30 |
B |
3054 |
2923 |
71.38 |
|
|
|
| 31 |
B |
3040 |
2910 |
40.73 |
|
|
|
| 32 |
B |
1506 |
1441 |
2.91 |
|
|
|
| 33 |
B |
1497 |
1433 |
9.39 |
|
|
|
| 34 |
B |
1479 |
1416 |
1.34 |
|
|
|
| 35 |
B |
1444 |
1382 |
4.58 |
|
|
|
| 36 |
B |
1405 |
1345 |
3.43 |
|
|
|
| 37 |
B |
1327 |
1270 |
8.74 |
|
|
|
| 38 |
B |
1287 |
1231 |
0.25 |
|
|
|
| 39 |
B |
1181 |
1130 |
0.08 |
|
|
|
| 40 |
B |
1096 |
1049 |
7.10 |
|
|
|
| 41 |
B |
1051 |
1006 |
3.65 |
|
|
|
| 42 |
B |
918 |
878 |
8.53 |
|
|
|
| 43 |
B |
810 |
776 |
5.74 |
|
|
|
| 44 |
B |
757 |
725 |
31.74 |
|
|
|
| 45 |
B |
566 |
542 |
6.18 |
|
|
|
| 46 |
B |
351 |
336 |
0.93 |
|
|
|
| 47 |
B |
235 |
225 |
0.03 |
|
|
|
| 48 |
B |
54 |
52 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36448.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34885.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.150 |
|
|
|
| 2 |
C |
-0.150 |
|
|
|
| 3 |
C |
-0.203 |
|
|
|
| 4 |
C |
-0.203 |
|
|
|
| 5 |
C |
-0.349 |
|
|
|
| 6 |
C |
-0.349 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.126 |
|
|
|
| 9 |
H |
0.112 |
|
|
|
| 10 |
H |
0.112 |
|
|
|
| 11 |
H |
0.119 |
|
|
|
| 12 |
H |
0.119 |
|
|
|
| 13 |
H |
0.115 |
|
|
|
| 14 |
H |
0.118 |
|
|
|
| 15 |
H |
0.112 |
|
|
|
| 16 |
H |
0.115 |
|
|
|
| 17 |
H |
0.118 |
|
|
|
| 18 |
H |
0.112 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.288 |
0.288 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.571 |
0.581 |
0.000 |
| y |
0.581 |
-38.452 |
0.000 |
| z |
0.000 |
0.000 |
-39.152 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.769 |
0.581 |
0.000 |
| y |
0.581 |
1.909 |
0.000 |
| z |
0.000 |
0.000 |
0.860 |
|
| Polar |
|---|
| 3z2-r2 | 1.720 |
|---|
| x2-y2 | -3.118 |
|---|
| xy | 0.581 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.147 |
-1.202 |
0.000 |
| y |
-1.202 |
14.110 |
0.000 |
| z |
0.000 |
0.000 |
9.750 |
<r2> (average value of r
2) Å
2
| <r2> |
255.411 |
| (<r2>)1/2 |
15.982 |