return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7N (Cyclopropylamine)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-172.937968
Energy at 298.15K-172.946448
HF Energy-172.149535
Nuclear repulsion energy126.093664
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3545 3366 0.00      
2 A' 3300 3133 7.48      
3 A' 3212 3049 3.56      
4 A' 3158 2998 34.46      
5 A' 1662 1578 15.13      
6 A' 1528 1450 17.62      
7 A' 1436 1364 18.65      
8 A' 1270 1206 0.68      
9 A' 1197 1137 4.56      
10 A' 1053 1000 13.81      
11 A' 1034 982 9.60      
12 A' 865 821 72.40      
13 A' 831 789 71.65      
14 A' 784 744 3.33      
15 A' 411 390 5.30      
16 A" 3635 3451 3.86      
17 A" 3288 3122 0.27      
18 A" 3205 3043 11.56      
19 A" 1474 1400 2.69      
20 A" 1279 1214 0.86      
21 A" 1210 1149 0.25      
22 A" 1149 1091 0.81      
23 A" 1070 1015 1.81      
24 A" 926 879 11.01      
25 A" 857 814 2.98      
26 A" 406 386 6.68      
27 A" 272 259 29.46      

Unscaled Zero Point Vibrational Energy (zpe) 22028.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 20913.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.55153 0.22829 0.19692

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.310 0.408 0.000
H2 -1.259 0.930 0.000
N3 0.887 1.190 0.000
C4 -0.310 -0.884 0.749
C5 -0.310 -0.884 -0.749
H6 0.927 1.778 0.818
H7 0.927 1.778 -0.818
H8 -1.208 -1.170 1.268
H9 -1.208 -1.170 -1.268
H10 0.597 -1.160 1.255
H11 0.597 -1.160 -1.255

Atom - Atom Distances (Å)
  C1 H2 N3 C4 C5 H6 H7 H8 H9 H10 H11
C11.08241.42971.49341.49342.01832.01832.21472.21472.20412.2041
H21.08242.16162.17922.17922.48292.48292.45342.45343.06373.0637
N31.42972.16162.50882.50881.00771.00773.40073.40072.67962.6796
C41.49342.17922.50881.49722.93583.32671.07582.22611.07532.2169
C51.49342.17922.50881.49723.32672.93582.22611.07582.21691.0753
H62.01832.48291.00772.93583.32671.63503.66714.19472.98813.6102
H72.01832.48291.00773.32672.93581.63504.19473.66713.61022.9881
H82.21472.45343.40071.07582.22613.66714.19472.53691.80513.1026
H92.21472.45343.40072.22611.07584.19473.66712.53693.10261.8051
H102.20413.06372.67961.07532.21692.98813.61021.80513.10262.5102
H112.20413.06372.67962.21691.07533.61022.98813.10261.80512.5102

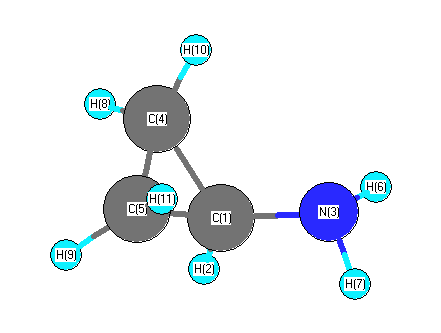
picture of Cyclopropylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 H6 110.606 C1 N3 H7 110.606
C1 C4 C5 59.917 C1 C4 H8 118.180
C1 C4 H10 117.265 C1 C5 C4 59.917
C1 C5 H9 118.180 C1 C5 H11 117.265
H2 C1 N3 118.075 H2 C1 C4 114.626
H2 C1 C5 114.626 N3 C1 C4 118.230
N3 C1 C5 118.230 C4 C1 C5 60.166
C4 C5 H9 118.898 C4 C5 H11 118.101
C5 C4 H8 118.898 C5 C4 H10 118.101
H6 N3 H7 108.441 H8 C4 H10 114.091
H9 C5 H11 114.091
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability