return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Methyl cyclopropyl ketone)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-270.086828
Energy at 298.15K-270.095547
HF Energy-268.929009
Nuclear repulsion energy232.214735
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3302 3135 1.35      
2 A' 3243 3079 4.67      
3 A' 3209 3046 6.33      
4 A' 3206 3044 6.21      
5 A' 3092 2935 2.14      
6 A' 1769 1679 90.89      
7 A' 1512 1436 11.88      
8 A' 1488 1413 16.13      
9 A' 1445 1371 113.63      
10 A' 1387 1317 14.67      
11 A' 1246 1183 2.60      
12 A' 1213 1152 77.71      
13 A' 1124 1067 12.17      
14 A' 1062 1009 10.16      
15 A' 984 934 43.81      
16 A' 947 899 11.45      
17 A' 821 780 1.99      
18 A' 767 728 0.55      
19 A' 598 568 11.10      
20 A' 377 358 3.95      
21 A' 243 231 3.90      
22 A" 3289 3122 0.49      
23 A" 3203 3041 7.53      
24 A" 3159 3000 4.40      
25 A" 1502 1426 9.50      
26 A" 1474 1399 4.43      
27 A" 1223 1161 0.36      
28 A" 1141 1083 2.20      
29 A" 1096 1041 2.19      
30 A" 1053 999 3.31      
31 A" 894 849 0.15      
32 A" 857 814 5.94      
33 A" 609 578 0.97      
34 A" 263 250 0.34      
35 A" 151 143 0.02      
36 A" 69 66 1.70      

Unscaled Zero Point Vibrational Energy (zpe) 26507.8 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 25166.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.24263 0.08963 0.07815

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -1.474 -0.539 0.000
C2 -0.275 -0.749 0.000
C3 0.295 -2.135 0.000
C4 0.701 0.360 0.000
C5 0.295 1.612 0.741
C6 0.295 1.612 -0.741
H7 -0.502 -2.866 0.000
H8 0.926 -2.271 -0.875
H9 0.926 -2.271 0.875
H10 1.744 0.099 0.000
H11 -0.661 1.561 1.230
H12 1.074 2.147 1.254
H13 -0.661 1.561 -1.230
H14 1.074 2.147 -1.254

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
O11.21792.38312.35422.88272.88272.52203.08663.08663.28072.56613.90892.56613.9089
C21.21791.49891.47752.53952.53952.12942.12702.12702.18932.64513.43162.64513.4316
C32.38311.49892.52833.81993.81991.08171.08691.08692.66274.01084.52924.01084.5292
C42.35421.47752.52831.51011.51013.44362.78222.78221.07462.19292.21412.19292.2141
C52.88272.53953.81991.51011.48164.60874.25303.93652.22151.07541.07452.19132.2068
C62.88272.53953.81991.51011.48164.60873.93654.25302.22152.19132.20681.07541.0745
H72.52202.12941.08173.44364.60874.60871.77721.77723.71974.59735.40234.59735.4023
H83.08662.12701.08692.78224.25303.93651.77721.74942.65564.65104.90624.16264.4367
H93.08662.12701.08692.78223.93654.25301.77721.74942.65564.16264.43674.65104.9062
H103.28072.18932.66271.07462.22152.22153.71972.65562.65563.07082.49273.07082.4927
H112.56612.64514.01082.19291.07542.19134.59734.65104.16263.07081.83092.46043.0858
H123.90893.43164.52922.21411.07452.20685.40234.90624.43672.49271.83093.08582.5076
H132.56612.64514.01082.19292.19131.07544.59734.16264.65103.07082.46043.08581.8309
H143.90893.43164.52922.21412.20681.07455.40234.43674.90622.49273.08582.50761.8309

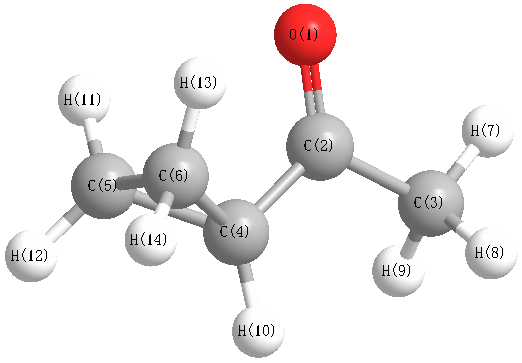
picture of Methyl cyclopropyl ketone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 122.272 O1 C2 C4 121.424
C2 C3 H7 110.157 C2 C3 H8 109.660
C2 C3 H9 109.660 C2 C4 C5 116.419
C2 C4 C6 116.419 C2 C4 H10 117.274
C3 C2 C4 116.304 C4 C5 C6 60.623
C4 C5 H11 114.985 C4 C5 H12 116.877
C4 C6 C5 60.623 C4 C6 H13 114.985
C4 C6 H14 116.877 C5 C4 C6 58.754
C5 C4 H10 117.521 C5 C6 H13 117.071
C5 C6 H14 118.517 C6 C4 H10 117.521
C6 C5 H11 117.071 C6 C5 H12 118.517
H7 C3 H8 110.071 H7 C3 H9 110.071
H8 C3 H9 107.172 H11 C5 H12 116.778
H13 C6 H14 116.778
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability