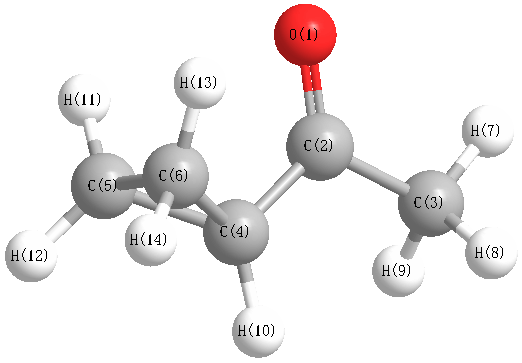
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3265 |
3125 |
4.44 |
|
|
|
| 2 |
A' |
3202 |
3064 |
8.31 |
|
|
|
| 3 |
A' |
3179 |
3043 |
9.60 |
|
|
|
| 4 |
A' |
3164 |
3028 |
9.59 |
|
|
|
| 5 |
A' |
3057 |
2926 |
2.41 |
|
|
|
| 6 |
A' |
1807 |
1729 |
141.57 |
|
|
|
| 7 |
A' |
1499 |
1434 |
14.71 |
|
|
|
| 8 |
A' |
1462 |
1400 |
15.29 |
|
|
|
| 9 |
A' |
1423 |
1362 |
118.12 |
|
|
|
| 10 |
A' |
1374 |
1315 |
13.88 |
|
|
|
| 11 |
A' |
1241 |
1188 |
0.97 |
|
|
|
| 12 |
A' |
1200 |
1149 |
87.30 |
|
|
|
| 13 |
A' |
1115 |
1067 |
13.35 |
|
|
|
| 14 |
A' |
1060 |
1014 |
10.89 |
|
|
|
| 15 |
A' |
975 |
934 |
54.91 |
|
|
|
| 16 |
A' |
943 |
903 |
7.48 |
|
|
|
| 17 |
A' |
811 |
777 |
2.53 |
|
|
|
| 18 |
A' |
760 |
727 |
0.32 |
|
|
|
| 19 |
A' |
595 |
570 |
11.31 |
|
|
|
| 20 |
A' |
372 |
356 |
4.24 |
|
|
|
| 21 |
A' |
243 |
233 |
4.10 |
|
|
|
| 22 |
A" |
3251 |
3112 |
0.54 |
|
|
|
| 23 |
A" |
3163 |
3027 |
13.75 |
|
|
|
| 24 |
A" |
3121 |
2987 |
6.73 |
|
|
|
| 25 |
A" |
1474 |
1411 |
9.14 |
|
|
|
| 26 |
A" |
1457 |
1394 |
4.85 |
|
|
|
| 27 |
A" |
1212 |
1160 |
0.31 |
|
|
|
| 28 |
A" |
1129 |
1081 |
2.15 |
|
|
|
| 29 |
A" |
1100 |
1053 |
1.73 |
|
|
|
| 30 |
A" |
1049 |
1004 |
4.05 |
|
|
|
| 31 |
A" |
897 |
859 |
0.55 |
|
|
|
| 32 |
A" |
848 |
812 |
5.09 |
|
|
|
| 33 |
A" |
604 |
578 |
1.10 |
|
|
|
| 34 |
A" |
268 |
257 |
0.38 |
|
|
|
| 35 |
A" |
142 |
136 |
0.02 |
|
|
|
| 36 |
A" |
72 |
69 |
1.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26266.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 25140.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.300 |
|
|
|
| 2 |
C |
0.220 |
|
|
|
| 3 |
C |
-0.371 |
|
|
|
| 4 |
C |
-0.100 |
|
|
|
| 5 |
C |
-0.260 |
|
|
|
| 6 |
C |
-0.260 |
|
|
|
| 7 |
H |
0.132 |
|
|
|
| 8 |
H |
0.125 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.131 |
|
|
|
| 11 |
H |
0.148 |
|
|
|
| 12 |
H |
0.131 |
|
|
|
| 13 |
H |
0.148 |
|
|
|
| 14 |
H |
0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.829 |
-0.025 |
0.000 |
2.829 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.479 |
-0.902 |
0.000 |
| y |
-0.902 |
-33.938 |
0.000 |
| z |
0.000 |
0.000 |
-35.908 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.555 |
-0.902 |
0.000 |
| y |
-0.902 |
3.755 |
0.000 |
| z |
0.000 |
0.000 |
0.800 |
|
| Polar |
|---|
| 3z2-r2 | 1.601 |
|---|
| x2-y2 | -5.540 |
|---|
| xy | -0.902 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.340 |
0.130 |
0.000 |
| y |
0.130 |
10.315 |
0.000 |
| z |
0.000 |
0.000 |
7.399 |
<r2> (average value of r
2) Å
2
| <r2> |
165.538 |
| (<r2>)1/2 |
12.866 |