Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.390170 |
| Energy at 298.15K | -235.402516 |
| HF Energy | -234.260241 |
| Nuclear repulsion energy | 232.762160 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3209 |
3047 |
36.21 |
|
|
|
| 2 |
A |
3183 |
3022 |
9.05 |
|
|
|
| 3 |
A |
3169 |
3009 |
8.18 |
|
|
|
| 4 |
A |
3164 |
3004 |
26.89 |
|
|
|
| 5 |
A |
3158 |
2998 |
38.35 |
|
|
|
| 6 |
A |
3140 |
2981 |
12.35 |
|
|
|
| 7 |
A |
3132 |
2974 |
15.50 |
|
|
|
| 8 |
A |
3111 |
2954 |
4.63 |
|
|
|
| 9 |
A |
3090 |
2934 |
41.76 |
|
|
|
| 10 |
A |
3086 |
2930 |
18.69 |
|
|
|
| 11 |
A |
3086 |
2930 |
13.38 |
|
|
|
| 12 |
A |
3076 |
2920 |
12.56 |
|
|
|
| 13 |
A |
1735 |
1647 |
2.32 |
|
|
|
| 14 |
A |
1533 |
1455 |
7.52 |
|
|
|
| 15 |
A |
1527 |
1450 |
7.75 |
|
|
|
| 16 |
A |
1522 |
1445 |
5.28 |
|
|
|
| 17 |
A |
1514 |
1438 |
0.29 |
|
|
|
| 18 |
A |
1510 |
1434 |
5.72 |
|
|
|
| 19 |
A |
1502 |
1426 |
0.53 |
|
|
|
| 20 |
A |
1453 |
1379 |
8.56 |
|
|
|
| 21 |
A |
1422 |
1350 |
3.30 |
|
|
|
| 22 |
A |
1414 |
1342 |
3.15 |
|
|
|
| 23 |
A |
1386 |
1316 |
0.20 |
|
|
|
| 24 |
A |
1341 |
1273 |
0.03 |
|
|
|
| 25 |
A |
1309 |
1243 |
0.39 |
|
|
|
| 26 |
A |
1304 |
1238 |
1.24 |
|
|
|
| 27 |
A |
1267 |
1203 |
0.17 |
|
|
|
| 28 |
A |
1183 |
1123 |
0.34 |
|
|
|
| 29 |
A |
1134 |
1076 |
1.71 |
|
|
|
| 30 |
A |
1089 |
1034 |
1.77 |
|
|
|
| 31 |
A |
1077 |
1022 |
2.01 |
|
|
|
| 32 |
A |
1075 |
1020 |
0.58 |
|
|
|
| 33 |
A |
1028 |
976 |
5.44 |
|
|
|
| 34 |
A |
1020 |
968 |
0.42 |
|
|
|
| 35 |
A |
938 |
891 |
3.50 |
|
|
|
| 36 |
A |
915 |
869 |
5.90 |
|
|
|
| 37 |
A |
883 |
838 |
2.96 |
|
|
|
| 38 |
A |
759 |
720 |
1.82 |
|
|
|
| 39 |
A |
746 |
708 |
32.25 |
|
|
|
| 40 |
A |
569 |
541 |
6.16 |
|
|
|
| 41 |
A |
487 |
463 |
0.96 |
|
|
|
| 42 |
A |
368 |
349 |
0.00 |
|
|
|
| 43 |
A |
300 |
285 |
0.07 |
|
|
|
| 44 |
A |
256 |
243 |
0.00 |
|
|
|
| 45 |
A |
205 |
195 |
0.04 |
|
|
|
| 46 |
A |
136 |
129 |
0.32 |
|
|
|
| 47 |
A |
93 |
89 |
0.01 |
|
|
|
| 48 |
A |
34 |
33 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36817.3 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 34954.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.352 |
0.995 |
-0.022 |
| C2 |
1.967 |
-0.436 |
-0.174 |
| C3 |
0.779 |
-0.980 |
0.097 |
| C4 |
-0.438 |
-0.276 |
0.586 |
| C5 |
-1.500 |
-0.156 |
-0.499 |
| C6 |
-2.769 |
0.498 |
0.011 |
| H7 |
1.529 |
1.609 |
0.323 |
| H8 |
3.172 |
1.099 |
0.684 |
| H9 |
2.697 |
1.398 |
-0.971 |
| H10 |
2.739 |
-1.092 |
-0.545 |
| H11 |
0.666 |
-2.038 |
-0.085 |
| H12 |
-0.194 |
0.716 |
0.957 |
| H13 |
-0.860 |
-0.827 |
1.426 |
| H14 |
-1.089 |
0.417 |
-1.327 |
| H15 |
-1.723 |
-1.147 |
-0.889 |
| H16 |
-3.516 |
0.584 |
-0.772 |
| H17 |
-3.200 |
-0.081 |
0.823 |
| H18 |
-2.563 |
1.495 |
0.388 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4895 | 2.5265 | 3.1249 | 4.0479 | 5.1450 | 1.0832 | 1.0874 | 1.0872 | 2.1860 | 3.4705 | 2.7415 | 3.9663 | 3.7246 | 4.6842 | 5.9297 | 5.7173 | 4.9568 |
C2 | 1.4895 | | 1.3337 | 2.5272 | 3.4932 | 4.8309 | 2.1498 | 2.1325 | 2.1289 | 1.0791 | 2.0658 | 2.6973 | 3.2720 | 3.3754 | 3.8253 | 5.6089 | 5.2739 | 4.9563 | C3 | 2.5265 | 1.3337 | | 1.4888 | 2.4959 | 3.8449 | 2.7041 | 3.2236 | 3.2357 | 2.0655 | 1.0802 | 2.1356 | 2.1160 | 2.7329 | 2.6953 | 4.6531 | 4.1434 | 4.1690 | C4 | 3.1249 | 2.5272 | 1.4888 | | 1.5233 | 2.5229 | 2.7365 | 3.8642 | 3.8796 | 3.4701 | 2.1851 | 1.0863 | 1.0896 | 2.1362 | 2.1418 | 3.4726 | 2.7788 | 2.7733 | C5 | 4.0479 | 3.4932 | 2.4959 | 1.5233 | | 1.5160 | 3.6005 | 4.9803 | 4.4999 | 4.3419 | 2.8991 | 2.1414 | 2.1363 | 1.0874 | 1.0878 | 2.1648 | 2.1548 | 2.1549 | C6 | 5.1450 | 4.8309 | 3.8449 | 2.5229 | 1.5160 | | 4.4504 | 6.0096 | 5.6258 | 5.7605 | 4.2708 | 2.7523 | 2.7205 | 2.1491 | 2.1464 | 1.0852 | 1.0864 | 1.0865 | H7 | 1.0832 | 2.1498 | 2.7041 | 2.7365 | 3.6005 | 4.4504 | | 1.7577 | 1.7556 | 3.0846 | 3.7699 | 2.0413 | 3.5858 | 3.3158 | 4.4315 | 5.2632 | 5.0462 | 4.0937 | H8 | 1.0874 | 2.1325 | 3.2236 | 3.8642 | 4.9803 | 6.0096 | 1.7577 | | 1.7473 | 2.5496 | 4.0888 | 3.3988 | 4.5303 | 4.7606 | 5.6111 | 6.8642 | 6.4816 | 5.7561 | H9 | 1.0872 | 2.1289 | 3.2357 | 3.8796 | 4.4999 | 5.6258 | 1.7556 | 1.7473 | | 2.5271 | 4.0889 | 3.5407 | 4.8319 | 3.9267 | 5.1010 | 6.2690 | 6.3382 | 5.4330 | H10 | 2.1860 | 1.0791 | 2.0655 | 3.4701 | 4.3419 | 5.7605 | 3.0846 | 2.5496 | 2.5271 | | 2.3253 | 3.7588 | 4.1124 | 4.1888 | 4.4765 | 6.4802 | 6.1780 | 5.9733 | H11 | 3.4705 | 2.0658 | 1.0802 | 2.1851 | 2.8991 | 4.2708 | 3.7699 | 4.0888 | 4.0889 | 2.3253 | | 3.0674 | 2.4653 | 3.2634 | 2.6739 | 4.9835 | 4.4269 | 4.8097 | H12 | 2.7415 | 2.6973 | 2.1356 | 1.0863 | 2.1414 | 2.7523 | 2.0413 | 3.3988 | 3.5407 | 3.7588 | 3.0674 | | 1.7449 | 2.4708 | 3.0356 | 3.7475 | 3.1124 | 2.5579 | H13 | 3.9663 | 3.2720 | 2.1160 | 1.0896 | 2.1363 | 2.7205 | 3.5858 | 4.5303 | 4.8319 | 4.1124 | 2.4653 | 1.7449 | | 3.0291 | 2.4908 | 3.7246 | 2.5282 | 3.0607 | H14 | 3.7246 | 3.3754 | 2.7329 | 2.1362 | 1.0874 | 2.1491 | 3.3158 | 4.7606 | 3.9267 | 4.1888 | 3.2634 | 2.4708 | 3.0291 | | 1.7434 | 2.4953 | 3.0538 | 2.5051 | H15 | 4.6842 | 3.8253 | 2.6953 | 2.1418 | 1.0878 | 2.1464 | 4.4315 | 5.6111 | 5.1010 | 4.4765 | 2.6739 | 3.0356 | 2.4908 | 1.7434 | | 2.4943 | 2.4993 | 3.0521 | H16 | 5.9297 | 5.6089 | 4.6531 | 3.4726 | 2.1648 | 1.0852 | 5.2632 | 6.8642 | 6.2690 | 6.4802 | 4.9835 | 3.7475 | 3.7246 | 2.4953 | 2.4943 | | 1.7569 | 1.7566 | H17 | 5.7173 | 5.2739 | 4.1434 | 2.7788 | 2.1548 | 1.0864 | 5.0462 | 6.4816 | 6.3382 | 6.1780 | 4.4269 | 3.1124 | 2.5282 | 3.0538 | 2.4993 | 1.7569 | | 1.7548 | H18 | 4.9568 | 4.9563 | 4.1690 | 2.7733 | 2.1549 | 1.0865 | 4.0937 | 5.7561 | 5.4330 | 5.9733 | 4.8097 | 2.5579 | 3.0607 | 2.5051 | 3.0521 | 1.7566 | 1.7548 | |
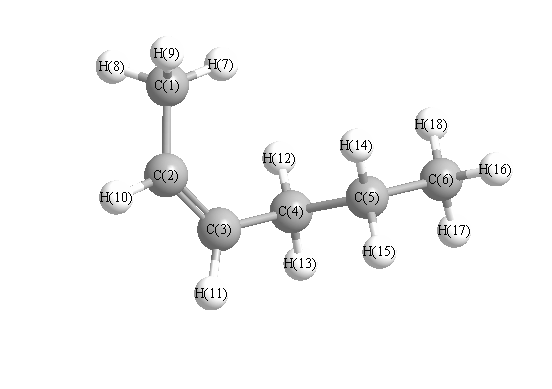 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.906 |
|
C1 |
C2 |
H10 |
115.728 |
| C2 |
C1 |
H7 |
112.398 |
|
C2 |
C1 |
H8 |
110.729 |
| C2 |
C1 |
H9 |
110.453 |
|
C2 |
C3 |
C4 |
127.031 |
| C2 |
C3 |
H11 |
117.303 |
|
C3 |
C2 |
H10 |
117.366 |
| C3 |
C4 |
C5 |
111.914 |
|
C3 |
C4 |
H12 |
111.100 |
| C3 |
C4 |
H13 |
109.333 |
|
C4 |
C3 |
H11 |
115.631 |
| C4 |
C5 |
C6 |
112.220 |
|
C4 |
C5 |
H14 |
108.672 |
| C4 |
C5 |
H15 |
109.085 |
|
C5 |
C4 |
H12 |
109.140 |
| C5 |
C4 |
H13 |
108.557 |
|
C5 |
C6 |
H16 |
111.585 |
| C5 |
C6 |
H17 |
110.702 |
|
C5 |
C6 |
H18 |
110.709 |
| C6 |
C5 |
H14 |
110.195 |
|
C6 |
C5 |
H15 |
109.949 |
| H7 |
C1 |
H8 |
108.151 |
|
H7 |
C1 |
H9 |
107.977 |
| H8 |
C1 |
H9 |
106.929 |
|
H12 |
C4 |
H13 |
106.626 |
| H14 |
C5 |
H16 |
94.359 |
|
H16 |
C6 |
H17 |
108.002 |
| H16 |
C6 |
H18 |
107.972 |
|
H17 |
C6 |
H18 |
107.726 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
