Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.781518 |
| Energy at 298.15K | -235.793775 |
| HF Energy | -235.781518 |
| Nuclear repulsion energy | 231.381629 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3167 |
3031 |
52.75 |
|
|
|
| 2 |
A |
3149 |
3014 |
7.92 |
|
|
|
| 3 |
A |
3137 |
3002 |
11.79 |
|
|
|
| 4 |
A |
3126 |
2992 |
35.84 |
|
|
|
| 5 |
A |
3121 |
2987 |
55.29 |
|
|
|
| 6 |
A |
3101 |
2968 |
10.42 |
|
|
|
| 7 |
A |
3094 |
2961 |
17.93 |
|
|
|
| 8 |
A |
3078 |
2946 |
7.24 |
|
|
|
| 9 |
A |
3050 |
2919 |
47.20 |
|
|
|
| 10 |
A |
3048 |
2918 |
12.85 |
|
|
|
| 11 |
A |
3047 |
2916 |
29.82 |
|
|
|
| 12 |
A |
3028 |
2898 |
19.10 |
|
|
|
| 13 |
A |
1754 |
1679 |
3.00 |
|
|
|
| 14 |
A |
1508 |
1443 |
6.84 |
|
|
|
| 15 |
A |
1498 |
1434 |
7.58 |
|
|
|
| 16 |
A |
1493 |
1429 |
4.42 |
|
|
|
| 17 |
A |
1491 |
1427 |
1.59 |
|
|
|
| 18 |
A |
1481 |
1418 |
5.79 |
|
|
|
| 19 |
A |
1478 |
1414 |
0.35 |
|
|
|
| 20 |
A |
1440 |
1379 |
11.21 |
|
|
|
| 21 |
A |
1410 |
1349 |
3.01 |
|
|
|
| 22 |
A |
1402 |
1342 |
3.09 |
|
|
|
| 23 |
A |
1376 |
1317 |
0.20 |
|
|
|
| 24 |
A |
1323 |
1266 |
0.03 |
|
|
|
| 25 |
A |
1294 |
1239 |
0.83 |
|
|
|
| 26 |
A |
1290 |
1234 |
2.36 |
|
|
|
| 27 |
A |
1251 |
1198 |
0.17 |
|
|
|
| 28 |
A |
1174 |
1124 |
0.47 |
|
|
|
| 29 |
A |
1118 |
1070 |
0.79 |
|
|
|
| 30 |
A |
1072 |
1026 |
2.93 |
|
|
|
| 31 |
A |
1063 |
1017 |
1.30 |
|
|
|
| 32 |
A |
1060 |
1014 |
1.63 |
|
|
|
| 33 |
A |
1020 |
976 |
1.08 |
|
|
|
| 34 |
A |
1010 |
967 |
6.87 |
|
|
|
| 35 |
A |
928 |
889 |
4.73 |
|
|
|
| 36 |
A |
904 |
865 |
6.35 |
|
|
|
| 37 |
A |
872 |
834 |
3.77 |
|
|
|
| 38 |
A |
745 |
713 |
2.49 |
|
|
|
| 39 |
A |
719 |
688 |
31.15 |
|
|
|
| 40 |
A |
568 |
543 |
7.48 |
|
|
|
| 41 |
A |
473 |
453 |
0.97 |
|
|
|
| 42 |
A |
360 |
344 |
0.00 |
|
|
|
| 43 |
A |
279 |
267 |
0.02 |
|
|
|
| 44 |
A |
240 |
230 |
0.01 |
|
|
|
| 45 |
A |
196 |
188 |
0.05 |
|
|
|
| 46 |
A |
135 |
129 |
0.28 |
|
|
|
| 47 |
A |
93 |
89 |
0.02 |
|
|
|
| 48 |
A |
55 |
53 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36359.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34799.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.474 |
0.932 |
-0.082 |
| C2 |
1.985 |
-0.474 |
-0.132 |
| C3 |
0.764 |
-0.919 |
0.140 |
| C4 |
-0.432 |
-0.126 |
0.544 |
| C5 |
-1.585 |
-0.257 |
-0.448 |
| C6 |
-2.824 |
0.502 |
-0.009 |
| H7 |
1.698 |
1.640 |
0.195 |
| H8 |
3.289 |
1.033 |
0.635 |
| H9 |
2.875 |
1.236 |
-1.050 |
| H10 |
2.727 |
-1.209 |
-0.422 |
| H11 |
0.593 |
-1.987 |
0.048 |
| H12 |
-0.182 |
0.927 |
0.668 |
| H13 |
-0.779 |
-0.474 |
1.521 |
| H14 |
-1.255 |
0.101 |
-1.423 |
| H15 |
-1.825 |
-1.313 |
-0.578 |
| H16 |
-3.633 |
0.396 |
-0.729 |
| H17 |
-3.186 |
0.139 |
0.953 |
| H18 |
-2.614 |
1.565 |
0.101 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4902 | 2.5303 | 3.1557 | 4.2449 | 5.3154 | 1.0858 | 1.0909 | 1.0907 | 2.1832 | 3.4752 | 2.7595 | 3.8899 | 4.0485 | 4.8753 | 6.1641 | 5.8080 | 5.1307 |
C2 | 1.4902 | | 1.3277 | 2.5342 | 3.5903 | 4.9082 | 2.1583 | 2.1356 | 2.1355 | 1.0840 | 2.0634 | 2.7014 | 3.2208 | 3.5346 | 3.9267 | 5.7161 | 5.3188 | 5.0368 | C3 | 2.5303 | 1.3277 | | 1.4915 | 2.5104 | 3.8618 | 2.7246 | 3.2298 | 3.2433 | 2.0627 | 1.0849 | 2.1403 | 2.1182 | 2.7500 | 2.7158 | 4.6711 | 4.1691 | 4.1940 | C4 | 3.1557 | 2.5342 | 1.4915 | | 1.5260 | 2.5332 | 2.7886 | 3.8988 | 3.9159 | 3.4772 | 2.1819 | 1.0889 | 1.0939 | 2.1445 | 2.1469 | 3.4836 | 2.7961 | 2.7960 | C5 | 4.2449 | 3.5903 | 2.5104 | 1.5260 | | 1.5173 | 3.8449 | 5.1566 | 4.7412 | 4.4161 | 2.8252 | 2.1480 | 2.1386 | 1.0906 | 1.0914 | 2.1679 | 2.1635 | 2.1633 | C6 | 5.3154 | 4.9082 | 3.8618 | 2.5332 | 1.5173 | | 4.6669 | 6.1696 | 5.8393 | 5.8233 | 4.2271 | 2.7603 | 2.7339 | 2.1502 | 2.1482 | 1.0883 | 1.0897 | 1.0895 | H7 | 1.0858 | 2.1583 | 2.7246 | 2.7886 | 3.8449 | 4.6669 | | 1.7594 | 1.7600 | 3.0913 | 3.7938 | 2.0652 | 3.5161 | 3.7016 | 4.6612 | 5.5511 | 5.1650 | 4.3139 | H8 | 1.0909 | 2.1356 | 3.2298 | 3.8988 | 5.1566 | 6.1696 | 1.7594 | | 1.7475 | 2.5415 | 4.0904 | 3.4726 | 4.4278 | 5.0747 | 5.7559 | 7.0838 | 6.5442 | 5.9517 | H9 | 1.0907 | 2.1355 | 3.2433 | 3.9159 | 4.7412 | 5.8393 | 1.7600 | 1.7475 | | 2.5289 | 4.0987 | 3.5201 | 4.7842 | 4.2988 | 5.3675 | 6.5695 | 6.4767 | 5.6183 | H10 | 2.1832 | 1.0840 | 2.0627 | 3.4772 | 4.4161 | 5.8233 | 3.0913 | 2.5415 | 2.5289 | | 2.3197 | 3.7700 | 4.0758 | 4.3099 | 4.5562 | 6.5668 | 6.2187 | 6.0420 | H11 | 3.4752 | 2.0634 | 1.0849 | 2.1819 | 2.8252 | 4.2271 | 3.7938 | 4.0904 | 4.0987 | 2.3197 | | 3.0778 | 2.5183 | 3.1525 | 2.5870 | 4.9131 | 4.4289 | 4.7861 | H12 | 2.7595 | 2.7014 | 2.1403 | 1.0889 | 2.1480 | 2.7603 | 2.0652 | 3.4726 | 3.5201 | 3.7700 | 3.0778 | | 1.7454 | 2.4916 | 3.0449 | 3.7608 | 3.1188 | 2.5785 | H13 | 3.8899 | 3.2208 | 2.1182 | 1.0939 | 2.1386 | 2.7339 | 3.5161 | 4.4278 | 4.7842 | 4.0758 | 2.5183 | 1.7454 | | 3.0380 | 2.4915 | 3.7370 | 2.5479 | 3.0898 | H14 | 4.0485 | 3.5346 | 2.7500 | 2.1445 | 1.0906 | 2.1502 | 3.7016 | 5.0747 | 4.2988 | 4.3099 | 3.1525 | 2.4916 | 3.0380 | | 1.7435 | 2.4951 | 3.0623 | 2.5130 | H15 | 4.8753 | 3.9267 | 2.7158 | 2.1469 | 1.0914 | 2.1482 | 4.6612 | 5.7559 | 5.3675 | 4.5562 | 2.5870 | 3.0449 | 2.4915 | 1.7435 | | 2.4925 | 2.5107 | 3.0609 | H16 | 6.1641 | 5.7161 | 4.6711 | 3.4836 | 2.1679 | 1.0883 | 5.5511 | 7.0838 | 6.5695 | 6.5668 | 4.9131 | 3.7608 | 3.7370 | 2.4951 | 2.4925 | | 1.7589 | 1.7583 | H17 | 5.8080 | 5.3188 | 4.1691 | 2.7961 | 2.1635 | 1.0897 | 5.1650 | 6.5442 | 6.4767 | 6.2187 | 4.4289 | 3.1188 | 2.5479 | 3.0623 | 2.5107 | 1.7589 | | 1.7572 | H18 | 5.1307 | 5.0368 | 4.1940 | 2.7960 | 2.1633 | 1.0895 | 4.3139 | 5.9517 | 5.6183 | 6.0420 | 4.7861 | 2.5785 | 3.0898 | 2.5130 | 3.0609 | 1.7583 | 1.7572 | |
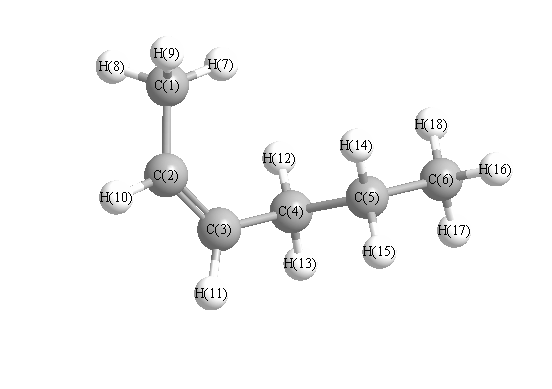 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.678 |
|
C1 |
C2 |
H10 |
115.102 |
| C2 |
C1 |
H7 |
112.888 |
|
C2 |
C1 |
H8 |
110.719 |
| C2 |
C1 |
H9 |
110.718 |
|
C2 |
C3 |
C4 |
127.941 |
| C2 |
C3 |
H11 |
117.215 |
|
C3 |
C2 |
H10 |
117.219 |
| C3 |
C4 |
C5 |
112.601 |
|
C3 |
C4 |
H12 |
111.126 |
| C3 |
C4 |
H13 |
109.060 |
|
C4 |
C3 |
H11 |
114.835 |
| C4 |
C5 |
C6 |
112.694 |
|
C4 |
C5 |
H14 |
108.952 |
| C4 |
C5 |
H15 |
109.093 |
|
C5 |
C4 |
H12 |
109.322 |
| C5 |
C4 |
H13 |
108.305 |
|
C5 |
C6 |
H16 |
111.552 |
| C5 |
C6 |
H17 |
111.114 |
|
C5 |
C6 |
H18 |
111.105 |
| C6 |
C5 |
H14 |
110.004 |
|
C6 |
C5 |
H15 |
109.791 |
| H7 |
C1 |
H8 |
107.857 |
|
H7 |
C1 |
H9 |
107.933 |
| H8 |
C1 |
H9 |
106.459 |
|
H12 |
C4 |
H13 |
106.185 |
| H14 |
C5 |
H16 |
94.081 |
|
H16 |
C6 |
H17 |
107.724 |
| H16 |
C6 |
H18 |
107.678 |
|
H17 |
C6 |
H18 |
107.481 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.342 |
|
|
|
| 2 |
C |
-0.141 |
|
|
|
| 3 |
C |
-0.163 |
|
|
|
| 4 |
C |
-0.205 |
|
|
|
| 5 |
C |
-0.197 |
|
|
|
| 6 |
C |
-0.364 |
|
|
|
| 7 |
H |
0.112 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.119 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.127 |
|
|
|
| 12 |
H |
0.115 |
|
|
|
| 13 |
H |
0.120 |
|
|
|
| 14 |
H |
0.117 |
|
|
|
| 15 |
H |
0.112 |
|
|
|
| 16 |
H |
0.119 |
|
|
|
| 17 |
H |
0.111 |
|
|
|
| 18 |
H |
0.112 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.058 |
0.219 |
-0.023 |
0.227 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.933 |
0.879 |
-0.546 |
| y |
0.879 |
-39.212 |
-0.034 |
| z |
-0.546 |
-0.034 |
-41.003 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.175 |
0.879 |
-0.546 |
| y |
0.879 |
0.755 |
-0.034 |
| z |
-0.546 |
-0.034 |
-1.930 |
|
| Polar |
|---|
| 3z2-r2 | -3.861 |
|---|
| x2-y2 | 0.280 |
|---|
| xy | 0.879 |
|---|
| xz | -0.546 |
|---|
| yz | -0.034 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.093 |
0.592 |
-0.544 |
| y |
0.592 |
10.070 |
0.045 |
| z |
-0.544 |
0.045 |
8.694 |
<r2> (average value of r
2) Å
2
| <r2> |
259.492 |
| (<r2>)1/2 |
16.109 |
