Jump to
S1C2
Vibrational Frequencies calculated at CCSD=FULL/cc-pVTZ
Geometric Data calculated at CCSD=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -624.337624 |
| Energy at 298.15K | -624.341551 |
| HF Energy | -623.352421 |
| Nuclear repulsion energy | 195.170388 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3809 |
3611 |
117.41 |
|
|
|
| 2 |
A' |
1289 |
1222 |
168.24 |
|
|
|
| 3 |
A' |
1149 |
1090 |
16.28 |
|
|
|
| 4 |
A' |
832 |
789 |
146.75 |
|
|
|
| 5 |
A' |
509 |
482 |
19.70 |
|
|
|
| 6 |
A' |
446 |
423 |
122.23 |
|
|
|
| 7 |
A' |
351 |
333 |
36.15 |
|
|
|
| 8 |
A" |
3806 |
3608 |
44.78 |
|
|
|
| 9 |
A" |
1124 |
1065 |
62.63 |
|
|
|
| 10 |
A" |
829 |
786 |
337.56 |
|
|
|
| 11 |
A" |
469 |
444 |
63.04 |
|
|
|
| 12 |
A" |
156 |
148 |
20.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7383.9 cm
-1
Scaled (by 0.9481) Zero Point Vibrational Energy (zpe) 7000.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.297 |
0.340 |
0.000 |
| O2 |
-1.031 |
0.936 |
0.000 |
| O3 |
0.297 |
-0.697 |
1.241 |
| O4 |
0.297 |
-0.697 |
-1.241 |
| H5 |
-0.626 |
-0.888 |
1.451 |
| H6 |
-0.626 |
-0.888 |
-1.451 |
Atom - Atom Distances (Å)
| |
S1 |
O2 |
O3 |
O4 |
H5 |
H6 |
| S1 | | 1.4555 | 1.6174 | 1.6174 | 2.1127 | 2.1127 |
O2 | 1.4555 | | 2.4435 | 2.4435 | 2.3653 | 2.3653 | O3 | 1.6174 | 2.4435 | | 2.4828 | 0.9656 | 2.8523 | O4 | 1.6174 | 2.4435 | 2.4828 | | 2.8523 | 0.9656 | H5 | 2.1127 | 2.3653 | 0.9656 | 2.8523 | | 2.9012 | H6 | 2.1127 | 2.3653 | 2.8523 | 0.9656 | 2.9012 | |
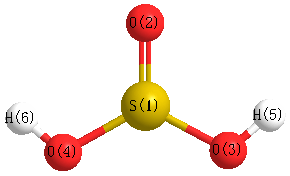 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
O3 |
H5 |
107.041 |
|
S1 |
O4 |
H6 |
107.041 |
| O2 |
S1 |
O3 |
105.223 |
|
O2 |
S1 |
O4 |
105.223 |
| O3 |
S1 |
O4 |
100.269 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
