Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVTZ
Geometric Data calculated at PBEPBEultrafine/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -624.762192 |
| Energy at 298.15K | -624.765805 |
| HF Energy | -624.762192 |
| Nuclear repulsion energy | 190.051598 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3586 |
3562 |
68.09 |
|
|
|
| 2 |
A' |
1200 |
1191 |
134.08 |
|
|
|
| 3 |
A' |
1068 |
1060 |
13.68 |
|
|
|
| 4 |
A' |
709 |
704 |
119.86 |
|
|
|
| 5 |
A' |
449 |
446 |
11.42 |
|
|
|
| 6 |
A' |
398 |
395 |
120.22 |
|
|
|
| 7 |
A' |
311 |
309 |
12.84 |
|
|
|
| 8 |
A" |
3584 |
3559 |
29.67 |
|
|
|
| 9 |
A" |
1046 |
1039 |
47.90 |
|
|
|
| 10 |
A" |
707 |
702 |
315.50 |
|
|
|
| 11 |
A" |
419 |
416 |
40.40 |
|
|
|
| 12 |
A" |
148 |
147 |
16.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6811.9 cm
-1
Scaled (by 0.9931) Zero Point Vibrational Energy (zpe) 6764.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.300 |
0.343 |
0.000 |
| O2 |
-1.040 |
0.974 |
0.000 |
| O3 |
0.300 |
-0.718 |
1.287 |
| O4 |
0.300 |
-0.718 |
-1.287 |
| H5 |
-0.643 |
-0.894 |
1.494 |
| H6 |
-0.643 |
-0.894 |
-1.494 |
Atom - Atom Distances (Å)
| |
S1 |
O2 |
O3 |
O4 |
H5 |
H6 |
| S1 | | 1.4813 | 1.6685 | 1.6685 | 2.1568 | 2.1568 |
O2 | 1.4813 | | 2.5133 | 2.5133 | 2.4244 | 2.4244 | O3 | 1.6685 | 2.5133 | | 2.5745 | 0.9813 | 2.9420 | O4 | 1.6685 | 2.5133 | 2.5745 | | 2.9420 | 0.9813 | H5 | 2.1568 | 2.4244 | 0.9813 | 2.9420 | | 2.9880 | H6 | 2.1568 | 2.4244 | 2.9420 | 0.9813 | 2.9880 | |
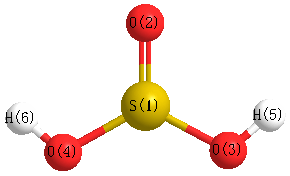 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
O3 |
H5 |
106.047 |
|
S1 |
O4 |
H6 |
106.047 |
| O2 |
S1 |
O3 |
105.709 |
|
O2 |
S1 |
O4 |
105.709 |
| O3 |
S1 |
O4 |
100.981 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.671 |
|
|
|
| 2 |
O |
-0.435 |
|
|
|
| 3 |
O |
-0.313 |
|
|
|
| 4 |
O |
-0.313 |
|
|
|
| 5 |
H |
0.195 |
|
|
|
| 6 |
H |
0.195 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.850 |
-1.233 |
0.000 |
1.498 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.961 |
5.021 |
0.000 |
| y |
5.021 |
-30.432 |
0.000 |
| z |
0.000 |
0.000 |
-28.422 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.534 |
5.021 |
0.000 |
| y |
5.021 |
-1.240 |
0.000 |
| z |
0.000 |
0.000 |
1.774 |
|
| Polar |
|---|
| 3z2-r2 | 3.549 |
|---|
| x2-y2 | 0.471 |
|---|
| xy | 5.021 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.477 |
-0.330 |
0.000 |
| y |
-0.330 |
4.584 |
0.000 |
| z |
0.000 |
0.000 |
5.213 |
<r2> (average value of r
2) Å
2
| <r2> |
81.157 |
| (<r2>)1/2 |
9.009 |
